Jump to
S1C2
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -169.731464 |
| Energy at 298.15K | |
| HF Energy | -168.986951 |
| Nuclear repulsion energy | 71.272921 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.371 |
0.000 |
| O2 |
-0.998 |
-0.530 |
0.000 |
| N3 |
1.192 |
-0.015 |
0.000 |
| H4 |
-0.329 |
1.408 |
0.000 |
| H5 |
-1.837 |
-0.076 |
0.000 |
| H6 |
1.809 |
0.784 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3451 | 1.2528 | 1.0878 | 1.8909 | 1.8554 |
O2 | 1.3451 | | 2.2499 | 2.0508 | 0.9540 | 3.0995 | N3 | 1.2528 | 2.2499 | | 2.0826 | 3.0296 | 1.0092 | H4 | 1.0878 | 2.0508 | 2.0826 | | 2.1161 | 2.2272 | H5 | 1.8909 | 0.9540 | 3.0296 | 2.1161 | | 3.7462 | H6 | 1.8554 | 3.0995 | 1.0092 | 2.2272 | 3.7462 | |
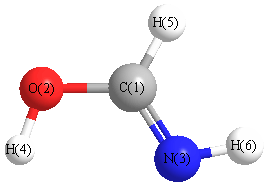 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.480 |
|
C1 |
N3 |
H6 |
109.749 |
| O2 |
C1 |
N3 |
119.960 |
|
O2 |
C1 |
H4 |
114.489 |
| N3 |
C1 |
H4 |
125.550 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -169.740077 |
| Energy at 298.15K | -169.744255 |
| HF Energy | -168.996138 |
| Nuclear repulsion energy | 71.611262 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3837 |
3837 |
64.12 |
|
|
|
| 2 |
A' |
3594 |
3594 |
11.09 |
|
|
|
| 3 |
A' |
3180 |
3180 |
20.55 |
|
|
|
| 4 |
A' |
1773 |
1773 |
263.49 |
|
|
|
| 5 |
A' |
1432 |
1432 |
23.68 |
|
|
|
| 6 |
A' |
1403 |
1403 |
0.74 |
|
|
|
| 7 |
A' |
1220 |
1220 |
101.83 |
|
|
|
| 8 |
A' |
1098 |
1098 |
183.52 |
|
|
|
| 9 |
A' |
603 |
603 |
50.05 |
|
|
|
| 10 |
A" |
1082 |
1082 |
0.10 |
|
|
|
| 11 |
A" |
855 |
855 |
33.24 |
|
|
|
| 12 |
A" |
617 |
617 |
169.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10346.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10346.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/cc-pVQZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.414 |
0.000 |
| O2 |
-1.106 |
-0.337 |
0.000 |
| N3 |
1.151 |
-0.091 |
0.000 |
| H4 |
-0.249 |
1.468 |
0.000 |
| H5 |
-0.816 |
-1.253 |
0.000 |
| H6 |
1.857 |
0.630 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3369 | 1.2566 | 1.0831 | 1.8565 | 1.8694 |
O2 | 1.3369 | | 2.2700 | 1.9983 | 0.9606 | 3.1166 | N3 | 1.2566 | 2.2700 | | 2.0951 | 2.2849 | 1.0090 | H4 | 1.0831 | 1.9983 | 2.0951 | | 2.7798 | 2.2667 | H5 | 1.8565 | 0.9606 | 2.2849 | 2.7798 | | 3.2700 | H6 | 1.8694 | 3.1166 | 1.0090 | 2.2667 | 3.2700 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.656 |
|
C1 |
N3 |
H6 |
110.730 |
| O2 |
C1 |
N3 |
122.120 |
|
O2 |
C1 |
H4 |
110.895 |
| N3 |
C1 |
H4 |
126.985 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
