Jump to
S1C2
S1C3
Energy calculated at CCSD(T)=FULL/aug-cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -1591.238886 |
| Energy at 298.15K | |
| HF Energy | -1590.166423 |
| Nuclear repulsion energy | 351.517405 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/aug-cc-pVQZ
Geometric Data calculated at CCSD(T)=FULL/aug-cc-pVQZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCSD(T)=FULL/aug-cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -1591.179911 |
| Energy at 298.15K | |
| HF Energy | -1590.110578 |
| Nuclear repulsion energy | 336.171363 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/aug-cc-pVQZ
Geometric Data calculated at CCSD(T)=FULL/aug-cc-pVQZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCSD(T)=FULL/aug-cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -1591.259369 |
| Energy at 298.15K | |
| HF Energy | -1590.105385 |
| Nuclear repulsion energy | 335.211163 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/aug-cc-pVQZ
Geometric Data calculated at CCSD(T)=FULL/aug-cc-pVQZ
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.948 |
1.269 |
| S2 |
0.000 |
0.948 |
-1.269 |
| S3 |
0.000 |
-0.948 |
1.269 |
| S4 |
0.000 |
-0.948 |
-1.269 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5387 | 1.8951 | 3.1680 |
S2 | 2.5387 | | 3.1680 | 1.8951 | S3 | 1.8951 | 3.1680 | | 2.5387 | S4 | 3.1680 | 1.8951 | 2.5387 | |
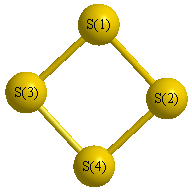 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
