Jump to
S1C2
Energy calculated at MP2/aug-cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -151.391054 |
| Energy at 298.15K | -151.393353 |
| HF Energy | -150.849516 |
| Nuclear repulsion energy | 36.933592 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/aug-cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3805 |
3614 |
18.21 |
97.24 |
0.08 |
0.15 |
| 2 |
A |
1432 |
1360 |
0.27 |
6.07 |
0.24 |
0.38 |
| 3 |
A |
933 |
886 |
0.49 |
7.87 |
0.13 |
0.23 |
| 4 |
A |
384 |
365 |
165.46 |
0.95 |
0.75 |
0.86 |
| 5 |
B |
3806 |
3614 |
66.15 |
21.64 |
0.75 |
0.86 |
| 6 |
B |
1329 |
1262 |
112.69 |
0.66 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 5843.9 cm
-1
Scaled (by 0.9497) Zero Point Vibrational Energy (zpe) 5550.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
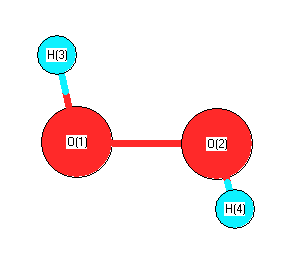
Geometric Data calculated at MP2/aug-cc-pVQZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.724 |
-0.059 |
| O2 |
0.000 |
-0.724 |
-0.059 |
| H3 |
0.791 |
0.888 |
0.468 |
| H4 |
-0.791 |
-0.888 |
0.468 |
Atom - Atom Distances (Å)
| |
O1 |
O2 |
H3 |
H4 |
| O1 | | 1.4471 | 0.9642 | 1.8705 |
O2 | 1.4471 | | 1.8705 | 0.9642 | H3 | 0.9642 | 1.8705 | | 2.3777 | H4 | 1.8705 | 0.9642 | 2.3777 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
O2 |
H4 |
99.799 |
|
O2 |
O1 |
H3 |
99.799 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/aug-cc-pVQZ
| | hartrees |
|---|
| Energy at 0K | -151.389320 |
| Energy at 298.15K | |
| HF Energy | -150.847376 |
| Nuclear repulsion energy | 36.777847 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/aug-cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3823 |
3631 |
0.00 |
|
|
|
| 2 |
Ag |
1520 |
1444 |
0.00 |
|
|
|
| 3 |
Ag |
930 |
883 |
0.00 |
|
|
|
| 4 |
Au |
303i |
288i |
250.98 |
|
|
|
| 5 |
Bu |
3832 |
3639 |
113.93 |
|
|
|
| 6 |
Bu |
1248 |
1185 |
140.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5525.2 cm
-1
Scaled (by 0.9497) Zero Point Vibrational Energy (zpe) 5247.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/aug-cc-pVQZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.729 |
0.000 |
| O2 |
0.000 |
-0.729 |
0.000 |
| H3 |
0.954 |
0.865 |
0.000 |
| H4 |
-0.954 |
-0.865 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
O2 |
H3 |
H4 |
| O1 | | 1.4581 | 0.9635 | 1.8575 |
O2 | 1.4581 | | 1.8575 | 0.9635 | H3 | 0.9635 | 1.8575 | | 2.5751 | H4 | 1.8575 | 0.9635 | 2.5751 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
O2 |
H4 |
98.101 |
|
O2 |
O1 |
H3 |
98.101 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
