return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for SiH3CCl3 ((trichloromethyl)silane)

using model chemistry: HSEh1PBE/cc-pV(T+d)Z

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3V 1A1
Energy calculated at HSEh1PBE/cc-pV(T+d)Z
 hartrees
Energy at 0K-1709.523101
Energy at 298.15K 
HF Energy-1709.523101
Nuclear repulsion energy439.794439
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/cc-pV(T+d)Z
Rotational Constants (cm-1) from geometry optimized at HSEh1PBE/cc-pV(T+d)Z
ABC
0.05607 0.05519 0.05519

See section I.F.4 to change rotational constant units
Geometric Data calculated at HSEh1PBE/cc-pV(T+d)Z

Point Group is C3v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 1.859
C2 0.000 0.000 -0.053
H3 0.000 -1.412 2.301
H4 1.223 0.706 2.301
H5 -1.223 0.706 2.301
Cl6 0.000 1.676 -0.639
Cl7 1.451 -0.838 -0.639
Cl8 -1.451 -0.838 -0.639

Atom - Atom Distances (Å)
  Si1 C2 H3 H4 H5 Cl6 Cl7 Cl8
Si11.91241.47981.47981.47983.00843.00843.0084
C21.91242.74562.74562.74561.77531.77531.7753
H31.47982.74562.44602.44604.26423.32923.3292
H41.47982.74562.44602.44603.32923.32924.2642
H51.47982.74562.44602.44603.32924.26423.3292
Cl63.00841.77534.26423.32923.32922.90252.9025
Cl73.00841.77533.32923.32924.26422.90252.9025
Cl83.00841.77533.32924.26423.32922.90252.9025

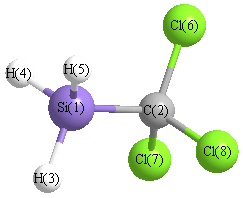
picture of (trichloromethyl)silane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Si1 C2 Cl6 109.276 Si1 C2 Cl7 109.276
Si1 C2 Cl8 109.276 C2 Si1 H3 107.389
C2 Si1 H4 107.389 C2 Si1 H5 107.389
H3 Si1 H4 111.470 H3 Si1 H5 111.470
H4 Si1 H5 111.470 Cl6 C2 Cl7 109.665
Cl6 C2 Cl8 109.665 Cl7 C2 Cl8 109.665
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/cc-pV(T+d)Z Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 Si 0.285      
2 C -0.052      
3 H -0.016      
4 H -0.016      
5 H -0.016      
6 Cl -0.062      
7 Cl -0.062      
8 Cl -0.062      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 1.381 1.381
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -58.325 0.000 0.000
y 0.000 -58.325 0.000
z 0.000 0.000 -55.865
Traceless
 xyz
x -1.230 0.000 0.000
y 0.000 -1.230 0.000
z 0.000 0.000 2.460
Polar
3z2-r24.920
x2-y20.000
xy0.000
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 270.259
(<r2>)1/2 16.440