Jump to
S1C2
Vibrational Frequencies calculated at MP4/Sadlej_pVTZ
Geometric Data calculated at MP4/Sadlej_pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/Sadlej_pVTZ
| | hartrees |
|---|
| Energy at 0K | -277.744251 |
| Energy at 298.15K | |
| HF Energy | -277.012590 |
| Nuclear repulsion energy | 126.843235 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/Sadlej_pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3093 |
3093 |
|
|
|
|
| 2 |
A |
3030 |
3030 |
|
|
|
|
| 3 |
A |
1460 |
1460 |
|
|
|
|
| 4 |
A |
1416 |
1416 |
|
|
|
|
| 5 |
A |
1275 |
1275 |
|
|
|
|
| 6 |
A |
1121 |
1121 |
|
|
|
|
| 7 |
A |
1094 |
1094 |
|
|
|
|
| 8 |
A |
866 |
866 |
|
|
|
|
| 9 |
A |
321 |
321 |
|
|
|
|
| 10 |
A |
152 |
152 |
|
|
|
|
| 11 |
B |
3106 |
3106 |
|
|
|
|
| 12 |
B |
3022 |
3022 |
|
|
|
|
| 13 |
B |
1454 |
1454 |
|
|
|
|
| 14 |
B |
1368 |
1368 |
|
|
|
|
| 15 |
B |
1235 |
1235 |
|
|
|
|
| 16 |
B |
1057 |
1057 |
|
|
|
|
| 17 |
B |
888 |
888 |
|
|
|
|
| 18 |
B |
489 |
489 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13222.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13222.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/Sadlej_pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.272 |
0.705 |
0.519 |
| C2 |
-0.272 |
-0.705 |
0.519 |
| F3 |
-0.272 |
1.422 |
-0.553 |
| F4 |
0.272 |
-1.422 |
-0.553 |
| H5 |
-0.011 |
1.220 |
1.459 |
| H6 |
1.373 |
0.698 |
0.405 |
| H7 |
0.011 |
-1.220 |
1.459 |
| H8 |
-1.373 |
-0.698 |
0.405 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5108 | 1.4003 | 2.3825 | 1.1081 | 1.1070 | 2.1574 | 2.1648 |
C2 | 1.5108 | | 2.3825 | 1.4003 | 2.1574 | 2.1648 | 1.1081 | 1.1070 | F3 | 1.4003 | 2.3825 | | 2.8963 | 2.0395 | 2.0360 | 3.3332 | 2.5745 | F4 | 2.3825 | 1.4003 | 2.8963 | | 3.3332 | 2.5745 | 2.0395 | 2.0360 | H5 | 1.1081 | 2.1574 | 2.0395 | 3.3332 | | 1.8161 | 2.4391 | 2.5778 | H6 | 1.1070 | 2.1648 | 2.0360 | 2.5745 | 1.8161 | | 2.5778 | 3.0803 | H7 | 2.1574 | 1.1081 | 3.3332 | 2.0395 | 2.4391 | 2.5778 | | 1.8161 | H8 | 2.1648 | 1.1070 | 2.5745 | 2.0360 | 2.5778 | 3.0803 | 1.8161 | |
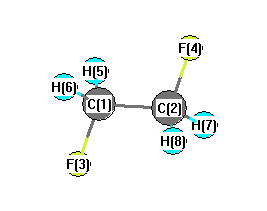 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.795 |
|
C1 |
C2 |
H7 |
109.978 |
| C1 |
C2 |
H8 |
110.627 |
|
C2 |
C1 |
F3 |
109.795 |
| C2 |
C1 |
H5 |
109.978 |
|
C2 |
C1 |
H6 |
110.627 |
| F3 |
C1 |
H5 |
108.226 |
|
F3 |
C1 |
H6 |
108.020 |
| F4 |
C2 |
H7 |
108.226 |
|
F4 |
C2 |
H8 |
108.020 |
| H5 |
C1 |
H6 |
110.139 |
|
H7 |
C2 |
H8 |
110.139 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
