Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/Sadlej_pVTZ
Geometric Data calculated at B2PLYP=FULLultrafine/Sadlej_pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/Sadlej_pVTZ
| | hartrees |
|---|
| Energy at 0K | -278.163166 |
| Energy at 298.15K | |
| HF Energy | -277.925915 |
| Nuclear repulsion energy | 127.346977 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/Sadlej_pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3122 |
3122 |
11.41 |
85.80 |
0.74 |
0.85 |
| 2 |
A |
3064 |
3064 |
35.77 |
240.73 |
0.02 |
0.03 |
| 3 |
A |
1471 |
1471 |
1.74 |
1.27 |
0.74 |
0.85 |
| 4 |
A |
1433 |
1433 |
10.37 |
1.43 |
0.21 |
0.35 |
| 5 |
A |
1297 |
1297 |
2.62 |
8.91 |
0.65 |
0.79 |
| 6 |
A |
1127 |
1127 |
22.75 |
1.83 |
0.56 |
0.72 |
| 7 |
A |
1112 |
1112 |
65.82 |
3.48 |
0.28 |
0.43 |
| 8 |
A |
878 |
878 |
29.29 |
6.89 |
0.13 |
0.24 |
| 9 |
A |
325 |
325 |
0.40 |
0.41 |
0.25 |
0.40 |
| 10 |
A |
151 |
151 |
3.82 |
0.12 |
0.75 |
0.86 |
| 11 |
B |
3135 |
3135 |
26.01 |
12.96 |
0.75 |
0.86 |
| 12 |
B |
3056 |
3056 |
8.69 |
29.96 |
0.75 |
0.86 |
| 13 |
B |
1466 |
1466 |
7.56 |
6.86 |
0.75 |
0.86 |
| 14 |
B |
1389 |
1389 |
8.05 |
0.04 |
0.75 |
0.86 |
| 15 |
B |
1256 |
1256 |
6.58 |
2.29 |
0.75 |
0.86 |
| 16 |
B |
1077 |
1077 |
54.05 |
3.65 |
0.75 |
0.86 |
| 17 |
B |
901 |
901 |
47.50 |
2.58 |
0.75 |
0.86 |
| 18 |
B |
497 |
497 |
16.16 |
0.47 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13377.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13377.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/Sadlej_pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.270 |
0.702 |
0.512 |
| C2 |
-0.270 |
-0.702 |
0.512 |
| F3 |
-0.270 |
1.426 |
-0.547 |
| F4 |
0.270 |
-1.426 |
-0.547 |
| H5 |
-0.007 |
1.208 |
1.450 |
| H6 |
1.364 |
0.697 |
0.399 |
| H7 |
0.007 |
-1.208 |
1.450 |
| H8 |
-1.364 |
-0.697 |
0.399 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5035 | 1.3922 | 2.3773 | 1.1014 | 1.1003 | 2.1442 | 2.1537 |
C2 | 1.5035 | | 2.3773 | 1.3922 | 2.1442 | 2.1537 | 1.1014 | 1.1003 | F3 | 1.3922 | 2.3773 | | 2.9034 | 2.0265 | 2.0237 | 3.3179 | 2.5696 | F4 | 2.3773 | 1.3922 | 2.9034 | | 3.3179 | 2.5696 | 2.0265 | 2.0237 | H5 | 1.1014 | 2.1442 | 2.0265 | 3.3179 | | 1.8016 | 2.4167 | 2.5649 | H6 | 1.1003 | 2.1537 | 2.0237 | 2.5696 | 1.8016 | | 2.5649 | 3.0637 | H7 | 2.1442 | 1.1014 | 3.3179 | 2.0265 | 2.4167 | 2.5649 | | 1.8016 | H8 | 2.1537 | 1.1003 | 2.5696 | 2.0237 | 2.5649 | 3.0637 | 1.8016 | |
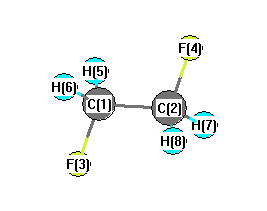 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.308 |
|
C1 |
C2 |
H7 |
109.840 |
| C1 |
C2 |
H8 |
110.658 |
|
C2 |
C1 |
F3 |
110.308 |
| C2 |
C1 |
H5 |
109.840 |
|
C2 |
C1 |
H6 |
110.658 |
| F3 |
C1 |
H5 |
108.155 |
|
F3 |
C1 |
H6 |
107.996 |
| F4 |
C2 |
H7 |
108.155 |
|
F4 |
C2 |
H8 |
107.996 |
| H5 |
C1 |
H6 |
109.829 |
|
H7 |
C2 |
H8 |
109.829 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
