return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2CH3 (Ethylbenzene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-309.916536
Energy at 298.15K-309.926827
HF Energy-308.790637
Nuclear repulsion energy340.779852
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3281 3066 16.48      
2 A' 3259 3045 2.79      
3 A' 3241 3029 9.10      
4 A' 3225 3013 35.52      
5 A' 3127 2922 28.71      
6 A' 3126 2921 24.90      
7 A' 1688 1577 2.42      
8 A' 1568 1465 6.40      
9 A' 1559 1457 5.18      
10 A' 1544 1443 1.03      
11 A' 1460 1364 1.36      
12 A' 1396 1304 3.44      
13 A' 1260 1177 0.25      
14 A' 1229 1148 0.07      
15 A' 1119 1046 7.56      
16 A' 1071 1000 1.65      
17 A' 1025 957 0.45      
18 A' 1008 942 2.26      
19 A' 918 858 0.04      
20 A' 884 826 0.37      
21 A' 793 741 0.78      
22 A' 744 695 54.32      
23 A' 599 560 8.36      
24 A' 521 487 0.38      
25 A' 483 451 3.75      
26 A' 307 287 0.25      
27 A' 135 126 0.55      
28 A" 3268 3054 29.66      
29 A" 3242 3030 8.80      
30 A" 3231 3019 22.12      
31 A" 3181 2973 10.02      
32 A" 1665 1556 0.29      
33 A" 1558 1456 7.07      
34 A" 1515 1416 6.32      
35 A" 1475 1378 0.44      
36 A" 1380 1290 0.57      
37 A" 1313 1227 0.02      
38 A" 1209 1130 0.02      
39 A" 1148 1073 2.25      
40 A" 1083 1012 2.09      
41 A" 914 854 0.00      
42 A" 850 794 0.02      
43 A" 806 753 0.40      
44 A" 635 594 0.02      
45 A" 398 372 0.00      
46 A" 365 341 0.05      
47 A" 234 218 0.01      
48 A" 40 38 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 35039.6 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 32741.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.15081 0.04933 0.04089

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.246 0.501 0.000
C2 -0.241 -0.215 1.202
C3 -0.241 -1.609 1.206
C4 -0.239 -2.310 0.000
C5 -0.241 -1.609 -1.206
C6 -0.241 -0.215 -1.202
C7 -0.183 2.004 0.000
C8 1.259 2.510 0.000
H9 -0.248 0.326 2.141
H10 -0.245 -2.146 2.145
H11 -0.241 -3.391 0.000
H12 -0.245 -2.146 -2.145
H13 -0.248 0.326 -2.141
H14 -0.706 2.391 -0.876
H15 -0.706 2.391 0.876
H16 1.789 2.148 0.880
H17 1.789 2.148 -0.880
H18 1.298 3.598 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.39902.42942.81022.42941.39901.50472.51052.14843.40633.89183.40632.14842.13362.13362.76192.76193.4608
C21.39901.39382.41532.78212.40442.52423.33471.08372.14863.39633.86403.38703.36512.66703.13183.74674.2838
C32.42941.39381.39452.41112.78213.80894.54602.14911.08202.15203.39313.86584.53264.04014.28264.75205.5616
C42.81022.41531.39451.39452.41534.31405.04703.39592.15111.08162.15113.39594.80434.80434.97574.97576.1043
C52.42942.78212.41111.39451.39383.80894.54603.86583.39312.15201.08202.14914.04014.53264.75204.28265.5616
C61.39902.40442.78212.41531.39382.52423.33473.38703.86403.39632.14861.08372.66703.36513.74673.13184.2838
C71.50472.52423.80894.31403.80892.52421.52832.72114.67155.39554.67152.72111.09111.09112.16372.16372.1756
C82.51053.33474.54605.04704.54603.33471.52833.40955.34196.08895.34193.40952.15512.15511.08841.08841.0889
H92.14841.08372.14913.39593.86583.38702.72113.40952.47174.28994.94774.28253.68452.46503.00984.07334.2046
H103.40632.14861.08202.15113.39313.86404.67155.34192.47172.48034.28974.94775.46964.73334.91665.63196.3221
H113.89183.39632.15201.08162.15203.39635.39556.08894.28992.48032.48034.28995.86665.86665.96495.96497.1568
H123.40633.86403.39312.15111.08202.14864.67155.34194.94774.28972.48032.47174.73335.46965.63194.91666.3221
H132.14843.38703.86583.39592.14911.08372.72113.40954.28254.94774.28992.47172.46503.68454.07333.00984.2046
H142.13363.36514.53264.80434.04012.66701.09112.15513.68455.46965.86664.73332.46501.75133.06012.50682.4980
H152.13362.66704.04014.80434.53263.36511.09112.15512.46504.73335.86665.46963.68451.75132.50683.06012.4980
H162.76193.13184.28264.97574.75203.74672.16371.08843.00984.91665.96495.63194.07333.06012.50681.75901.7654
H172.76193.74674.75204.97574.28263.13182.16371.08844.07335.63195.96494.91663.00982.50683.06011.75901.7654
H183.46084.28385.56166.10435.56164.28382.17561.08894.20466.32217.15686.32214.20462.49802.49801.76541.7654

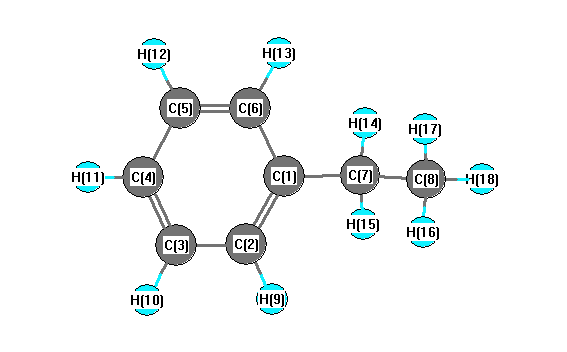
picture of Ethylbenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.890 C1 C2 H9 119.302
C1 C6 C5 120.890 C1 C6 H13 119.302
C1 C7 C8 111.730 C1 C7 H14 109.530
C1 C7 H15 109.530 C2 C1 C6 118.490
C2 C1 C7 120.715 C2 C3 C4 120.041
C2 C3 H10 119.892 C3 C2 H9 119.805
C3 C4 C5 119.645 C3 C4 H11 120.177
C4 C3 H10 120.067 C4 C5 C6 120.041
C4 C5 H12 120.067 C5 C4 H11 120.177
C5 C6 H13 119.805 C6 C1 C7 120.715
C6 C5 H12 119.892 C7 C8 H16 110.428
C7 C8 H17 110.428 C7 C8 H18 111.355
C8 C7 H14 109.589 C8 C7 H15 109.589
H14 C7 H15 106.744 H16 C8 H17 107.818
H16 C8 H18 108.352 H17 C8 H18 108.352
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability