return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-344.571832
Energy at 298.15K 
HF Energy-343.441260
Nuclear repulsion energy321.030318
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3287 3071 9.76      
2 A' 3281 3066 10.81      
3 A' 3272 3057 10.42      
4 A' 3261 3047 0.75      
5 A' 3247 3034 4.90      
6 A' 3003 2806 121.43      
7 A' 1772 1656 133.21      
8 A' 1684 1574 12.06      
9 A' 1666 1557 9.71      
10 A' 1557 1455 0.57      
11 A' 1517 1417 11.30      
12 A' 1486 1388 2.18      
13 A' 1448 1353 9.39      
14 A' 1362 1272 14.58      
15 A' 1262 1180 54.66      
16 A' 1218 1138 8.24      
17 A' 1212 1132 2.15      
18 A' 1124 1050 3.46      
19 A' 1064 994 1.20      
20 A' 1022 955 0.52      
21 A' 851 795 31.30      
22 A' 663 619 19.60      
23 A' 629 588 0.68      
24 A' 445 415 0.20      
25 A' 223 209 7.63      
26 A" 1017 950 0.08      
27 A" 936 874 0.10      
28 A" 927 866 0.15      
29 A" 899 840 0.75      
30 A" 856 800 0.07      
31 A" 741 692 69.28      
32 A" 516 483 0.01      
33 A" 442 413 4.04      
34 A" 399 372 0.10      
35 A" 230 215 6.66      
36 A" 114 107 4.45      

Unscaled Zero Point Vibrational Energy (zpe) 24314.1 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 22719.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.17479 0.05227 0.04024

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.573 0.000
C2 -1.034 -0.370 0.000
C3 -0.727 -1.726 0.000
C4 0.607 -2.143 0.000
C5 1.637 -1.203 0.000
C6 1.334 0.157 0.000
C7 -0.317 2.016 0.000
O8 -1.454 2.474 0.000
H9 0.562 2.687 0.000
H10 -2.057 -0.019 0.000
H11 -1.521 -2.460 0.000
H12 0.842 -3.199 0.000
H13 2.668 -1.529 0.000
H14 2.126 0.897 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.39932.41082.78292.41511.39751.47782.39342.18732.14023.39303.86463.39662.1509
C21.39931.39002.41602.79802.42622.49192.87513.44831.08142.14623.39433.87943.4047
C32.41081.39001.39792.42152.79183.76454.26244.59682.16371.08162.15213.40103.8753
C42.78292.41601.39791.39492.41244.26075.05634.82983.40712.15161.08172.15073.3981
C52.41512.79802.42151.39491.39313.76554.80344.03493.87913.39942.14911.08142.1553
C61.39752.42622.79182.41241.39312.48633.62502.64453.39563.87343.39222.15011.0836
C71.47782.49193.76454.26073.76552.48631.22571.10542.67784.63575.34244.63452.6875
O82.39342.87514.26245.05634.80343.62501.22572.02712.56524.93506.12015.74593.9122
H92.18733.44834.59684.82984.03492.64451.10542.02713.76565.55245.89224.71222.3770
H102.14021.08142.16373.40713.87913.39562.67782.56523.76562.49944.30284.96054.2822
H113.39302.14621.08162.15163.39943.87344.63574.93505.55242.49942.47554.29154.9569
H123.86463.39432.15211.08172.14913.39225.34246.12015.89224.30282.47552.47504.2923
H133.39663.87943.40102.15071.08142.15014.63455.74594.71224.96054.29152.47502.4852
H142.15093.40473.87533.39812.15531.08362.68753.91222.37704.28224.95694.29232.4852

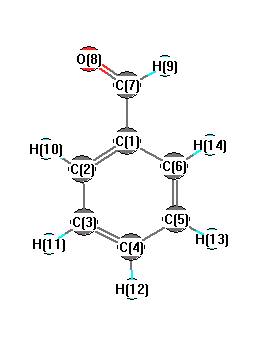
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.605 C1 C2 H10 118.695
C1 C6 C5 119.866 C1 C6 H14 119.675
C1 C7 O8 124.314 C1 C7 H9 114.951
C2 C1 C6 120.337 C2 C1 C7 119.989
C2 C3 C4 120.126 C2 C3 H11 120.018
C3 C2 H10 121.701 C3 C4 C5 120.236
C3 C4 H12 119.897 C4 C3 H11 119.856
C4 C5 C6 119.830 C4 C5 H13 120.041
C5 C4 H12 119.867 C5 C6 H14 120.458
C6 C1 C7 119.674 C6 C5 H13 120.130
O8 C7 H9 120.735
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability