return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-344.575483
Energy at 298.15K 
HF Energy-343.441404
Nuclear repulsion energy320.538527
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3277 3085 7.78      
2 A' 3269 3077 17.75      
3 A' 3258 3067 17.27      
4 A' 3246 3056 1.73      
5 A' 3234 3045 6.25      
6 A' 3001 2825 116.91      
7 A' 1806 1700 175.32      
8 A' 1701 1602 14.05      
9 A' 1683 1584 9.78      
10 A' 1568 1476 0.23      
11 A' 1526 1437 9.92      
12 A' 1458 1373 9.93      
13 A' 1373 1292 11.29      
14 A' 1334 1256 9.93      
15 A' 1255 1182 44.08      
16 A' 1219 1147 14.07      
17 A' 1202 1132 1.00      
18 A' 1124 1059 2.54      
19 A' 1066 1003 0.58      
20 A' 1026 965 0.14      
21 A' 849 800 31.59      
22 A' 663 625 21.51      
23 A' 632 595 0.79      
24 A' 445 419 0.15      
25 A' 225 212 8.07      
26 A" 1021 961 0.00      
27 A" 957 901 0.14      
28 A" 946 891 0.19      
29 A" 915 862 0.87      
30 A" 866 816 0.05      
31 A" 747 704 66.21      
32 A" 570 537 0.84      
33 A" 447 420 4.07      
34 A" 404 380 0.10      
35 A" 230 217 7.24      
36 A" 114 107 5.33      

Unscaled Zero Point Vibrational Energy (zpe) 24329.1 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 22903.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.17438 0.05204 0.04008

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.574 0.000
C2 -1.035 -0.372 0.000
C3 -0.728 -1.730 0.000
C4 0.609 -2.147 0.000
C5 1.641 -1.206 0.000
C6 1.335 0.157 0.000
C7 -0.320 2.025 0.000
O8 -1.454 2.478 0.000
H9 0.557 2.698 0.000
H10 -2.059 -0.024 0.000
H11 -1.523 -2.465 0.000
H12 0.844 -3.204 0.000
H13 2.673 -1.531 0.000
H14 2.129 0.895 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.40172.41612.78762.42031.39831.48622.39622.19602.14393.39903.87043.40212.1530
C21.40171.39222.41882.80222.42762.50132.88103.45811.08202.14933.39813.88463.4079
C32.41611.39221.40052.42622.79563.77724.27054.61072.16411.08252.15473.40713.8801
C42.78762.41881.40051.39642.41504.27385.06424.84463.40962.15541.08282.15403.4005
C52.42032.80222.42621.39641.39653.77914.81144.05103.88403.40502.15101.08242.1570
C61.39832.42762.79562.41501.39652.49553.62862.65713.39853.87813.39612.15401.0845
C71.48622.50133.77724.27383.77912.49551.22141.10572.68724.64855.35664.64782.6967
O82.39622.88104.27055.06424.81143.62861.22142.02352.57394.94396.12905.75383.9169
H92.19603.45814.61074.84464.05102.65711.10572.02353.77515.56635.90834.72832.3911
H102.14391.08202.16413.40963.88403.39852.68722.57393.77512.49984.30584.96644.2874
H113.39902.14931.08252.15543.40503.87814.64854.94395.56632.49982.47924.29864.9626
H123.87043.39812.15471.08282.15103.39615.35666.12905.90834.30582.47922.47874.2957
H133.40213.88463.40712.15401.08242.15404.64785.75384.72834.96644.29862.47872.4866
H142.15303.40793.88013.40052.15701.08452.69673.91692.39114.28744.96264.29572.4866

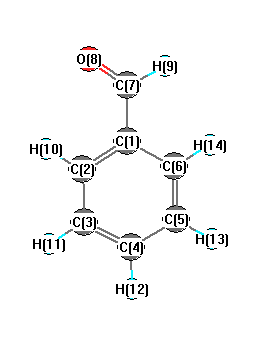
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.711 C1 C2 H10 118.784
C1 C6 C5 120.003 C1 C6 H14 119.731
C1 C7 O8 124.217 C1 C7 H9 115.049
C2 C1 C6 120.227 C2 C1 C7 119.998
C2 C3 C4 120.020 C2 C3 H11 120.054
C3 C2 H10 121.505 C3 C4 C5 120.334
C3 C4 H12 119.835 C4 C3 H11 119.926
C4 C5 C6 119.705 C4 C5 H13 120.154
C5 C4 H12 119.831 C5 C6 H14 120.266
C6 C1 C7 119.775 C6 C5 H13 120.142
O8 C7 H9 120.734
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability