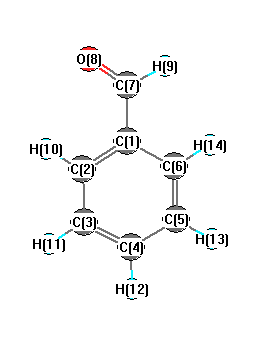
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3244 |
3092 |
8.17 |
192.65 |
0.12 |
0.21 |
| 2 |
A' |
3237 |
3085 |
12.87 |
98.47 |
0.14 |
0.24 |
| 3 |
A' |
3227 |
3076 |
12.85 |
82.22 |
0.75 |
0.86 |
| 4 |
A' |
3215 |
3064 |
1.18 |
78.50 |
0.71 |
0.83 |
| 5 |
A' |
3203 |
3052 |
4.77 |
46.87 |
0.42 |
0.60 |
| 6 |
A' |
2938 |
2801 |
128.98 |
120.28 |
0.30 |
0.46 |
| 7 |
A' |
1839 |
1752 |
237.83 |
79.77 |
0.34 |
0.51 |
| 8 |
A' |
1686 |
1607 |
27.69 |
78.16 |
0.52 |
0.69 |
| 9 |
A' |
1670 |
1591 |
10.56 |
8.56 |
0.55 |
0.71 |
| 10 |
A' |
1548 |
1476 |
0.79 |
1.14 |
0.27 |
0.43 |
| 11 |
A' |
1510 |
1439 |
13.16 |
1.07 |
0.27 |
0.42 |
| 12 |
A' |
1442 |
1374 |
9.77 |
4.16 |
0.71 |
0.83 |
| 13 |
A' |
1399 |
1334 |
6.12 |
0.96 |
0.70 |
0.82 |
| 14 |
A' |
1348 |
1285 |
15.85 |
1.64 |
0.49 |
0.66 |
| 15 |
A' |
1250 |
1191 |
61.22 |
23.56 |
0.26 |
0.41 |
| 16 |
A' |
1201 |
1145 |
16.98 |
6.66 |
0.20 |
0.33 |
| 17 |
A' |
1196 |
1140 |
2.36 |
8.38 |
0.70 |
0.82 |
| 18 |
A' |
1117 |
1065 |
5.41 |
0.72 |
0.24 |
0.39 |
| 19 |
A' |
1062 |
1012 |
1.73 |
12.71 |
0.11 |
0.20 |
| 20 |
A' |
1023 |
975 |
0.17 |
20.58 |
0.10 |
0.18 |
| 21 |
A' |
851 |
811 |
30.74 |
9.43 |
0.16 |
0.27 |
| 22 |
A' |
664 |
633 |
23.49 |
2.84 |
0.38 |
0.55 |
| 23 |
A' |
628 |
599 |
0.52 |
5.35 |
0.75 |
0.86 |
| 24 |
A' |
444 |
424 |
0.17 |
5.39 |
0.36 |
0.53 |
| 25 |
A' |
218 |
208 |
7.87 |
0.59 |
0.55 |
0.71 |
| 26 |
A" |
1042 |
993 |
0.36 |
7.16 |
0.75 |
0.86 |
| 27 |
A" |
1016 |
968 |
0.01 |
0.15 |
0.75 |
0.86 |
| 28 |
A" |
993 |
946 |
0.10 |
0.01 |
0.75 |
0.86 |
| 29 |
A" |
946 |
901 |
1.71 |
0.79 |
0.75 |
0.86 |
| 30 |
A" |
873 |
832 |
0.04 |
4.54 |
0.75 |
0.86 |
| 31 |
A" |
766 |
730 |
49.16 |
1.49 |
0.75 |
0.86 |
| 32 |
A" |
707 |
674 |
22.32 |
0.17 |
0.75 |
0.86 |
| 33 |
A" |
465 |
443 |
4.74 |
0.42 |
0.75 |
0.86 |
| 34 |
A" |
419 |
399 |
0.13 |
0.03 |
0.75 |
0.86 |
| 35 |
A" |
240 |
228 |
7.31 |
1.24 |
0.75 |
0.86 |
| 36 |
A" |
125 |
119 |
4.10 |
2.35 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24373.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 23230.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.030 |
|
|
|
| 2 |
C |
-0.175 |
|
|
|
| 3 |
C |
-0.173 |
|
|
|
| 4 |
C |
-0.157 |
|
|
|
| 5 |
C |
-0.174 |
|
|
|
| 6 |
C |
-0.199 |
|
|
|
| 7 |
C |
0.198 |
|
|
|
| 8 |
O |
-0.408 |
|
|
|
| 9 |
H |
0.131 |
|
|
|
| 10 |
H |
0.204 |
|
|
|
| 11 |
H |
0.182 |
|
|
|
| 12 |
H |
0.181 |
|
|
|
| 13 |
H |
0.180 |
|
|
|
| 14 |
H |
0.180 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.231 |
-2.408 |
0.000 |
3.283 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.441 |
5.950 |
0.000 |
| y |
5.950 |
-45.605 |
0.000 |
| z |
0.000 |
0.000 |
-48.141 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.432 |
5.950 |
0.000 |
| y |
5.950 |
0.186 |
0.000 |
| z |
0.000 |
0.000 |
-3.618 |
|
| Polar |
|---|
| 3z2-r2 | -7.235 |
|---|
| x2-y2 | 2.164 |
|---|
| xy | 5.950 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.402 |
-1.105 |
0.000 |
| y |
-1.105 |
14.709 |
0.000 |
| z |
0.000 |
0.000 |
3.887 |
<r2> (average value of r
2) Å
2
| <r2> |
260.417 |
| (<r2>)1/2 |
16.137 |