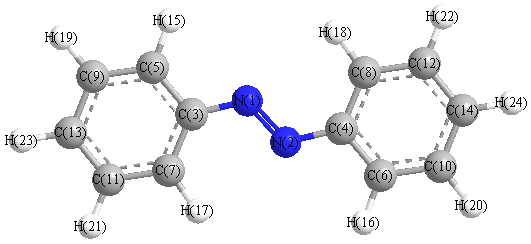
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3255 |
3103 |
0.00 |
|
|
|
| 2 |
Ag |
3244 |
3092 |
0.00 |
|
|
|
| 3 |
Ag |
3234 |
3082 |
0.00 |
|
|
|
| 4 |
Ag |
3224 |
3072 |
0.00 |
|
|
|
| 5 |
Ag |
3213 |
3062 |
0.00 |
|
|
|
| 6 |
Ag |
1689 |
1610 |
0.00 |
|
|
|
| 7 |
Ag |
1673 |
1594 |
0.00 |
|
|
|
| 8 |
Ag |
1603 |
1528 |
0.00 |
|
|
|
| 9 |
Ag |
1532 |
1460 |
0.00 |
|
|
|
| 10 |
Ag |
1508 |
1437 |
0.00 |
|
|
|
| 11 |
Ag |
1402 |
1337 |
0.00 |
|
|
|
| 12 |
Ag |
1349 |
1286 |
0.00 |
|
|
|
| 13 |
Ag |
1232 |
1174 |
0.00 |
|
|
|
| 14 |
Ag |
1193 |
1137 |
0.00 |
|
|
|
| 15 |
Ag |
1183 |
1128 |
0.00 |
|
|
|
| 16 |
Ag |
1110 |
1058 |
0.00 |
|
|
|
| 17 |
Ag |
1059 |
1009 |
0.00 |
|
|
|
| 18 |
Ag |
1023 |
975 |
0.00 |
|
|
|
| 19 |
Ag |
950 |
906 |
0.00 |
|
|
|
| 20 |
Ag |
685 |
653 |
0.00 |
|
|
|
| 21 |
Ag |
625 |
595 |
0.00 |
|
|
|
| 22 |
Ag |
309 |
294 |
0.00 |
|
|
|
| 23 |
Ag |
225 |
214 |
0.00 |
|
|
|
| 24 |
Au |
1014 |
966 |
1.03 |
|
|
|
| 25 |
Au |
993 |
946 |
0.04 |
|
|
|
| 26 |
Au |
959 |
914 |
9.62 |
|
|
|
| 27 |
Au |
868 |
827 |
0.00 |
|
|
|
| 28 |
Au |
804 |
766 |
65.68 |
|
|
|
| 29 |
Au |
709 |
675 |
69.64 |
|
|
|
| 30 |
Au |
563 |
537 |
10.27 |
|
|
|
| 31 |
Au |
419 |
399 |
0.02 |
|
|
|
| 32 |
Au |
310 |
296 |
0.69 |
|
|
|
| 33 |
Au |
65 |
62 |
1.33 |
|
|
|
| 34 |
Au |
26 |
25 |
0.04 |
|
|
|
| 35 |
Bg |
1013 |
966 |
0.00 |
|
|
|
| 36 |
Bg |
993 |
947 |
0.00 |
|
|
|
| 37 |
Bg |
954 |
909 |
0.00 |
|
|
|
| 38 |
Bg |
869 |
828 |
0.00 |
|
|
|
| 39 |
Bg |
783 |
747 |
0.00 |
|
|
|
| 40 |
Bg |
702 |
669 |
0.00 |
|
|
|
| 41 |
Bg |
491 |
468 |
0.00 |
|
|
|
| 42 |
Bg |
422 |
402 |
0.00 |
|
|
|
| 43 |
Bg |
260 |
248 |
0.00 |
|
|
|
| 44 |
Bg |
106 |
101 |
0.00 |
|
|
|
| 45 |
Bu |
3255 |
3103 |
9.69 |
|
|
|
| 46 |
Bu |
3244 |
3091 |
33.36 |
|
|
|
| 47 |
Bu |
3234 |
3082 |
40.68 |
|
|
|
| 48 |
Bu |
3224 |
3072 |
19.39 |
|
|
|
| 49 |
Bu |
3213 |
3062 |
2.29 |
|
|
|
| 50 |
Bu |
1684 |
1605 |
3.07 |
|
|
|
| 51 |
Bu |
1666 |
1588 |
5.00 |
|
|
|
| 52 |
Bu |
1544 |
1472 |
11.64 |
|
|
|
| 53 |
Bu |
1508 |
1437 |
14.23 |
|
|
|
| 54 |
Bu |
1399 |
1333 |
8.71 |
|
|
|
| 55 |
Bu |
1342 |
1279 |
1.48 |
|
|
|
| 56 |
Bu |
1287 |
1227 |
29.20 |
|
|
|
| 57 |
Bu |
1193 |
1137 |
0.18 |
|
|
|
| 58 |
Bu |
1187 |
1131 |
29.92 |
|
|
|
| 59 |
Bu |
1114 |
1062 |
14.28 |
|
|
|
| 60 |
Bu |
1059 |
1009 |
11.97 |
|
|
|
| 61 |
Bu |
1022 |
974 |
3.66 |
|
|
|
| 62 |
Bu |
850 |
810 |
0.65 |
|
|
|
| 63 |
Bu |
632 |
602 |
0.74 |
|
|
|
| 64 |
Bu |
546 |
521 |
5.10 |
|
|
|
| 65 |
Bu |
534 |
509 |
25.96 |
|
|
|
| 66 |
Bu |
84 |
80 |
1.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42327.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 40342.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.307 |
|
|
|
| 2 |
N |
-0.307 |
|
|
|
| 3 |
C |
0.244 |
|
|
|
| 4 |
C |
0.244 |
|
|
|
| 5 |
C |
-0.170 |
|
|
|
| 6 |
C |
-0.170 |
|
|
|
| 7 |
C |
-0.149 |
|
|
|
| 8 |
C |
-0.149 |
|
|
|
| 9 |
C |
-0.181 |
|
|
|
| 10 |
C |
-0.181 |
|
|
|
| 11 |
C |
-0.188 |
|
|
|
| 12 |
C |
-0.188 |
|
|
|
| 13 |
C |
-0.157 |
|
|
|
| 14 |
C |
-0.157 |
|
|
|
| 15 |
H |
0.183 |
|
|
|
| 16 |
H |
0.183 |
|
|
|
| 17 |
H |
0.197 |
|
|
|
| 18 |
H |
0.197 |
|
|
|
| 19 |
H |
0.176 |
|
|
|
| 20 |
H |
0.176 |
|
|
|
| 21 |
H |
0.176 |
|
|
|
| 22 |
H |
0.176 |
|
|
|
| 23 |
H |
0.175 |
|
|
|
| 24 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-64.834 |
5.010 |
0.000 |
| y |
5.010 |
-67.365 |
0.000 |
| z |
0.000 |
0.000 |
-85.262 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.480 |
5.010 |
0.000 |
| y |
5.010 |
7.683 |
0.000 |
| z |
0.000 |
0.000 |
-19.163 |
|
| Polar |
|---|
| 3z2-r2 | -38.326 |
|---|
| x2-y2 | 2.531 |
|---|
| xy | 5.010 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
33.409 |
10.622 |
0.000 |
| y |
10.622 |
29.209 |
0.000 |
| z |
0.000 |
0.000 |
6.717 |
<r2> (average value of r
2) Å
2
| <r2> |
1089.601 |
| (<r2>)1/2 |
33.009 |