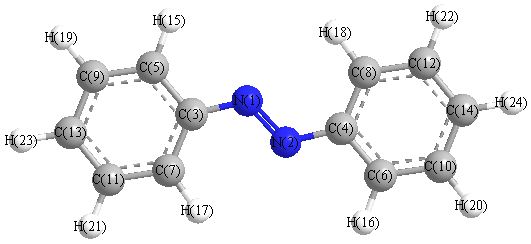
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3250 |
3101 |
0.00 |
|
|
|
| 2 |
Ag |
3239 |
3091 |
0.00 |
|
|
|
| 3 |
Ag |
3230 |
3082 |
0.00 |
|
|
|
| 4 |
Ag |
3220 |
3072 |
0.00 |
|
|
|
| 5 |
Ag |
3209 |
3062 |
0.00 |
|
|
|
| 6 |
Ag |
1685 |
1608 |
0.00 |
|
|
|
| 7 |
Ag |
1668 |
1591 |
0.00 |
|
|
|
| 8 |
Ag |
1597 |
1524 |
0.00 |
|
|
|
| 9 |
Ag |
1524 |
1454 |
0.00 |
|
|
|
| 10 |
Ag |
1501 |
1433 |
0.00 |
|
|
|
| 11 |
Ag |
1402 |
1337 |
0.00 |
|
|
|
| 12 |
Ag |
1341 |
1280 |
0.00 |
|
|
|
| 13 |
Ag |
1229 |
1173 |
0.00 |
|
|
|
| 14 |
Ag |
1186 |
1132 |
0.00 |
|
|
|
| 15 |
Ag |
1176 |
1123 |
0.00 |
|
|
|
| 16 |
Ag |
1105 |
1054 |
0.00 |
|
|
|
| 17 |
Ag |
1055 |
1007 |
0.00 |
|
|
|
| 18 |
Ag |
1021 |
974 |
0.00 |
|
|
|
| 19 |
Ag |
950 |
907 |
0.00 |
|
|
|
| 20 |
Ag |
685 |
654 |
0.00 |
|
|
|
| 21 |
Ag |
625 |
596 |
0.00 |
|
|
|
| 22 |
Ag |
309 |
295 |
0.00 |
|
|
|
| 23 |
Ag |
226 |
215 |
0.00 |
|
|
|
| 24 |
Au |
1014 |
968 |
0.79 |
|
|
|
| 25 |
Au |
996 |
951 |
0.02 |
|
|
|
| 26 |
Au |
959 |
916 |
9.13 |
|
|
|
| 27 |
Au |
867 |
828 |
0.01 |
|
|
|
| 28 |
Au |
804 |
767 |
61.82 |
|
|
|
| 29 |
Au |
708 |
676 |
65.95 |
|
|
|
| 30 |
Au |
563 |
537 |
9.44 |
|
|
|
| 31 |
Au |
418 |
399 |
0.01 |
|
|
|
| 32 |
Au |
309 |
295 |
0.76 |
|
|
|
| 33 |
Au |
65 |
62 |
1.31 |
|
|
|
| 34 |
Au |
26 |
25 |
0.03 |
|
|
|
| 35 |
Bg |
1014 |
968 |
0.00 |
|
|
|
| 36 |
Bg |
996 |
951 |
0.00 |
|
|
|
| 37 |
Bg |
954 |
911 |
0.00 |
|
|
|
| 38 |
Bg |
868 |
829 |
0.00 |
|
|
|
| 39 |
Bg |
783 |
748 |
0.00 |
|
|
|
| 40 |
Bg |
702 |
670 |
0.00 |
|
|
|
| 41 |
Bg |
491 |
469 |
0.00 |
|
|
|
| 42 |
Bg |
422 |
403 |
0.00 |
|
|
|
| 43 |
Bg |
261 |
249 |
0.00 |
|
|
|
| 44 |
Bg |
106 |
101 |
0.00 |
|
|
|
| 45 |
Bu |
3249 |
3101 |
10.76 |
|
|
|
| 46 |
Bu |
3239 |
3091 |
33.22 |
|
|
|
| 47 |
Bu |
3229 |
3082 |
35.89 |
|
|
|
| 48 |
Bu |
3220 |
3072 |
17.76 |
|
|
|
| 49 |
Bu |
3209 |
3062 |
2.71 |
|
|
|
| 50 |
Bu |
1679 |
1602 |
3.22 |
|
|
|
| 51 |
Bu |
1661 |
1585 |
4.88 |
|
|
|
| 52 |
Bu |
1537 |
1467 |
11.75 |
|
|
|
| 53 |
Bu |
1502 |
1433 |
14.29 |
|
|
|
| 54 |
Bu |
1398 |
1334 |
8.32 |
|
|
|
| 55 |
Bu |
1334 |
1273 |
1.29 |
|
|
|
| 56 |
Bu |
1287 |
1228 |
29.28 |
|
|
|
| 57 |
Bu |
1186 |
1132 |
0.27 |
|
|
|
| 58 |
Bu |
1179 |
1125 |
29.30 |
|
|
|
| 59 |
Bu |
1109 |
1058 |
14.85 |
|
|
|
| 60 |
Bu |
1054 |
1006 |
12.41 |
|
|
|
| 61 |
Bu |
1020 |
973 |
3.74 |
|
|
|
| 62 |
Bu |
850 |
811 |
0.80 |
|
|
|
| 63 |
Bu |
631 |
603 |
0.76 |
|
|
|
| 64 |
Bu |
547 |
522 |
4.77 |
|
|
|
| 65 |
Bu |
534 |
510 |
25.84 |
|
|
|
| 66 |
Bu |
85 |
81 |
1.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42248.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 40317.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.306 |
|
|
|
| 2 |
N |
-0.306 |
|
|
|
| 3 |
C |
0.218 |
|
|
|
| 4 |
C |
0.218 |
|
|
|
| 5 |
C |
-0.109 |
|
|
|
| 6 |
C |
-0.109 |
|
|
|
| 7 |
C |
-0.084 |
|
|
|
| 8 |
C |
-0.084 |
|
|
|
| 9 |
C |
-0.135 |
|
|
|
| 10 |
C |
-0.135 |
|
|
|
| 11 |
C |
-0.143 |
|
|
|
| 12 |
C |
-0.143 |
|
|
|
| 13 |
C |
-0.111 |
|
|
|
| 14 |
C |
-0.111 |
|
|
|
| 15 |
H |
0.136 |
|
|
|
| 16 |
H |
0.136 |
|
|
|
| 17 |
H |
0.147 |
|
|
|
| 18 |
H |
0.147 |
|
|
|
| 19 |
H |
0.129 |
|
|
|
| 20 |
H |
0.129 |
|
|
|
| 21 |
H |
0.130 |
|
|
|
| 22 |
H |
0.130 |
|
|
|
| 23 |
H |
0.128 |
|
|
|
| 24 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-65.089 |
4.979 |
0.000 |
| y |
4.979 |
-67.578 |
0.000 |
| z |
0.000 |
0.000 |
-85.068 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.234 |
4.979 |
0.000 |
| y |
4.979 |
7.500 |
0.000 |
| z |
0.000 |
0.000 |
-18.734 |
|
| Polar |
|---|
| 3z2-r2 | -37.469 |
|---|
| x2-y2 | 2.489 |
|---|
| xy | 4.979 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
33.853 |
10.834 |
0.000 |
| y |
10.834 |
29.536 |
0.000 |
| z |
0.000 |
0.000 |
6.814 |
<r2> (average value of r
2) Å
2
| <r2> |
1088.520 |
| (<r2>)1/2 |
32.993 |