return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C12H10N2 (azobenzene)

using model chemistry: TPSSh/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1AG
Energy calculated at TPSSh/6-31G**
 hartrees
Energy at 0K-572.819137
Energy at 298.15K-572.830436
HF Energy-572.819137
Nuclear repulsion energy749.261396
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3224 3091 0.00      
2 Ag 3213 3080 0.00      
3 Ag 3202 3070 0.00      
4 Ag 3191 3060 0.00      
5 Ag 3181 3049 0.00      
6 Ag 1650 1582 0.00      
7 Ag 1637 1569 0.00      
8 Ag 1539 1476 0.00      
9 Ag 1510 1447 0.00      
10 Ag 1477 1416 0.00      
11 Ag 1381 1324 0.00      
12 Ag 1341 1286 0.00      
13 Ag 1212 1162 0.00      
14 Ag 1185 1137 0.00      
15 Ag 1163 1115 0.00      
16 Ag 1098 1052 0.00      
17 Ag 1043 1000 0.00      
18 Ag 1011 969 0.00      
19 Ag 934 895 0.00      
20 Ag 673 645 0.00      
21 Ag 615 589 0.00      
22 Ag 302 289 0.00      
23 Ag 219 210 0.00      
24 Au 1005 963 0.63      
25 Au 988 947 0.02      
26 Au 951 912 8.44      
27 Au 862 826 0.01      
28 Au 795 762 65.20      
29 Au 700 671 57.20      
30 Au 554 531 8.01      
31 Au 412 395 0.01      
32 Au 304 291 0.83      
33 Au 63 61 1.31      
34 Au 27 25 0.05      
35 Bg 1004 963 0.00      
36 Bg 988 947 0.00      
37 Bg 946 907 0.00      
38 Bg 863 827 0.00      
39 Bg 776 744 0.00      
40 Bg 693 665 0.00      
41 Bg 483 463 0.00      
42 Bg 416 399 0.00      
43 Bg 255 244 0.00      
44 Bg 107 103 0.00      
45 Bu 3224 3091 15.86      
46 Bu 3212 3080 54.81      
47 Bu 3202 3070 57.61      
48 Bu 3191 3060 25.07      
49 Bu 3181 3049 3.20      
50 Bu 1648 1580 4.05      
51 Bu 1632 1564 4.46      
52 Bu 1524 1461 9.40      
53 Bu 1492 1430 11.36      
54 Bu 1378 1321 7.93      
55 Bu 1335 1280 2.44      
56 Bu 1259 1207 19.89      
57 Bu 1185 1136 0.16      
58 Bu 1176 1128 35.36      
59 Bu 1101 1056 12.41      
60 Bu 1043 1000 10.32      
61 Bu 1011 969 3.03      
62 Bu 834 800 0.64      
63 Bu 622 596 0.87      
64 Bu 535 513 7.62      
65 Bu 523 502 23.74      
66 Bu 79 76 1.78      

Unscaled Zero Point Vibrational Energy (zpe) 41787.9 cm-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 40062.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at TPSSh/6-31G**
ABC
0.09083 0.00976 0.00881

See section I.F.4 to change rotational constant units
Geometric Data calculated at TPSSh/6-31G**

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -0.010 0.633 0.000
N2 0.010 -0.633 0.000
C3 1.275 1.235 0.000
C4 -1.275 -1.235 0.000
C5 1.280 2.637 0.000
C6 -1.280 -2.637 0.000
C7 2.489 0.525 0.000
C8 -2.489 -0.525 0.000
C9 2.489 3.333 0.000
C10 -2.489 -3.333 0.000
C11 3.691 1.225 0.000
C12 -3.691 -1.225 0.000
C13 3.696 2.628 0.000
C14 -3.696 -2.628 0.000
H15 0.326 3.155 0.000
H16 -0.326 -3.155 0.000
H17 2.459 -0.558 0.000
H18 -2.459 0.558 0.000
H19 2.492 4.418 0.000
H20 -2.492 -4.418 0.000
H21 4.632 0.682 0.000
H22 -4.632 -0.682 0.000
H23 4.639 3.166 0.000
H24 -4.639 -3.166 0.000

Atom - Atom Distances (Å)
  N1 N2 C3 C4 C5 C6 C7 C8 C9 C10 C11 C12 C13 C14 H15 H16 H17 H18 H19 H20 H21 H22 H23 H24
N11.26671.41832.25602.38343.50872.50162.73703.67874.67723.74794.12384.20844.92162.54433.80172.74102.44994.53715.62844.64184.80555.29425.9888
N21.26672.25601.41833.50872.38342.73702.50164.67723.67874.12383.74794.92164.20843.80172.54432.44992.74105.62844.53714.80554.64185.98885.2942
C31.41832.25603.54931.40264.63911.40694.15532.42415.91852.41655.54152.79346.29492.14194.67282.14893.79393.40836.79303.40256.20963.87957.3716
C42.25601.41833.54934.63911.40264.15531.40695.91852.42415.54152.41656.29492.79344.67282.14193.79392.14896.79303.40836.20963.40257.37163.8795
C52.38343.50871.40264.63915.86342.43384.92071.39497.06052.79416.29542.41597.24451.08556.01133.40624.27782.15418.00083.88046.78013.40078.2895
C63.50872.38344.63911.40265.86344.92072.43387.06051.39496.29542.79417.24452.41596.01131.08554.27783.40628.00082.15416.78013.88048.28953.4007
C72.50162.73701.40694.15532.43384.92075.08862.80716.29871.39056.42362.42396.94273.40544.63411.08434.94813.89297.01822.14807.22293.40498.0278
C82.73702.50164.15531.40694.92072.43385.08866.29872.80716.42361.39056.94272.42394.63413.40544.94811.08437.01823.89297.22292.14808.02783.4049
C93.67874.67722.42415.91851.39497.06052.80716.29878.31952.42587.67931.39728.58982.17077.07243.89115.67271.08589.21363.40808.17502.15659.6461
C104.67723.67875.91852.42417.06051.39496.29872.80718.31957.67932.42588.58981.39727.07242.17075.67273.89119.21361.08588.17503.40809.64612.1565
C113.74794.12382.41655.54152.79416.29541.39056.42362.42587.67937.77811.40268.33143.87925.94352.16806.18563.41088.37131.08638.53862.15989.4168
C124.12383.74795.54152.41656.29542.79416.42361.39057.67932.42587.77818.33141.40265.94353.87926.18562.16808.37133.41088.53861.08639.41682.1598
C134.20844.92162.79346.29492.41597.24452.42396.94271.39728.58981.40268.33149.06983.41097.04413.41806.49312.15779.37752.15918.96141.086110.1510
C144.92164.20846.29492.79347.24452.41596.94272.42398.58981.39728.33141.40269.06987.04413.41096.49313.41809.37752.15778.96142.159110.15101.0861
H152.54433.80172.14194.67281.08556.01133.40544.63412.17077.07243.87925.94353.41097.04416.34414.28243.80762.50738.08094.96556.26944.31358.0380
H163.80172.54434.67282.14196.01131.08554.63413.40547.07242.17075.94353.87927.04413.41096.34413.80764.28248.08092.50736.26944.96558.03804.3135
H172.74102.44992.14893.79393.40624.27781.08434.94813.89115.67272.16806.18563.41806.49314.28243.80765.04244.97686.27742.50247.09144.31567.5617
H182.44992.74103.79392.14894.27783.40624.94811.08435.67273.89116.18562.16806.49313.41803.80764.28245.04246.27744.97687.09142.50247.56174.3156
H194.53715.62843.40836.79302.15418.00083.89297.01821.08589.21363.41088.37132.15779.37752.50738.08094.97686.277410.14524.30568.76142.486210.4102
H205.62844.53716.79303.40838.00082.15417.01823.89299.21361.08588.37133.41089.37752.15778.08092.50736.27744.976810.14528.76144.305610.41022.4862
H214.64184.80553.40256.20963.88046.78012.14807.22293.40808.17501.08638.53862.15918.96144.96556.26942.50247.09144.30568.76149.36352.483510.0380
H224.80554.64186.20963.40256.78013.88047.22292.14808.17503.40808.53861.08638.96142.15916.26944.96557.09142.50248.76144.30569.363510.03802.4835
H235.29425.98883.87957.37163.40078.28953.40498.02782.15659.64612.15989.41681.086110.15104.31358.03804.31567.56172.486210.41022.483510.038011.2331
H245.98885.29427.37163.87958.28953.40078.02783.40499.64612.15659.41682.159810.15101.08618.03804.31357.56174.315610.41022.486210.03802.483511.2331

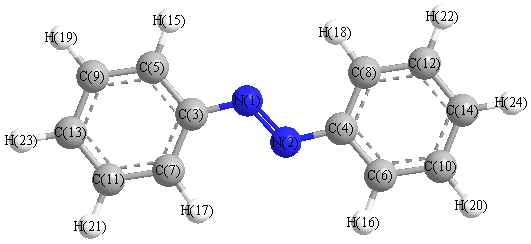
picture of azobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 114.215 N1 C3 C5 115.326
N1 C3 C7 124.616 N2 N1 C3 114.215
N2 C4 C6 115.326 N2 C4 C8 124.616
C3 C5 C9 120.112 C3 C5 H15 118.265
C3 C7 C11 119.500 C3 C7 H17 118.650
C4 C6 C10 120.112 C4 C6 H16 118.265
C4 C8 C12 119.500 C4 C8 H18 118.650
C5 C3 C7 120.057 C5 C9 C13 119.822
C5 C9 H19 120.014 C6 C4 C8 120.057
C6 C10 C14 119.822 C6 C10 H20 120.014
C7 C11 C13 120.417 C7 C11 H21 119.786
C8 C12 C14 120.417 C8 C12 H22 119.786
C9 C5 H15 121.623 C9 C13 C11 120.091
C9 C13 H23 120.027 C10 C6 H16 121.623
C10 C14 C12 120.091 C10 C14 H24 120.027
C11 C7 H17 121.850 C11 C13 H23 119.882
C12 C8 H18 121.850 C12 C14 H24 119.882
C13 C9 H19 120.163 C13 C11 H21 119.798
C14 C10 H20 120.163 C14 C12 H22 119.798
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 N -0.314      
2 N -0.314      
3 C 0.235      
4 C 0.235      
5 C -0.109      
6 C -0.109      
7 C -0.086      
8 C -0.086      
9 C -0.129      
10 C -0.129      
11 C -0.138      
12 C -0.138      
13 C -0.105      
14 C -0.105      
15 H 0.132      
16 H 0.132      
17 H 0.143      
18 H 0.143      
19 H 0.124      
20 H 0.124      
21 H 0.125      
22 H 0.125      
23 H 0.123      
24 H 0.123      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 0.000 0.000
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 35.136 11.607 0.000
y 11.607 30.415 0.000
z 0.000 0.000 6.831


<r2> (average value of r2) Å2
<r2> 1096.697
(<r2>)1/2 33.116