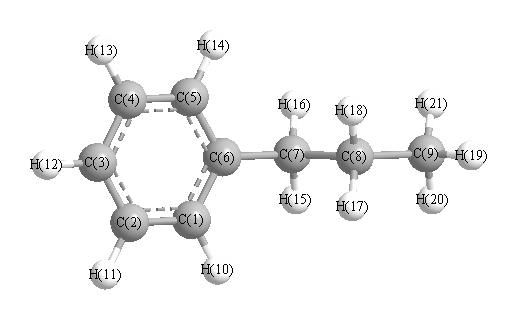
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3235 |
3083 |
18.46 |
|
|
|
| 2 |
A' |
3213 |
3062 |
4.64 |
|
|
|
| 3 |
A' |
3197 |
3047 |
10.14 |
|
|
|
| 4 |
A' |
3149 |
3001 |
32.12 |
|
|
|
| 5 |
A' |
3069 |
2925 |
57.65 |
|
|
|
| 6 |
A' |
3064 |
2921 |
22.25 |
|
|
|
| 7 |
A' |
3056 |
2913 |
7.38 |
|
|
|
| 8 |
A' |
1692 |
1613 |
6.22 |
|
|
|
| 9 |
A' |
1556 |
1483 |
13.45 |
|
|
|
| 10 |
A' |
1535 |
1463 |
6.71 |
|
|
|
| 11 |
A' |
1520 |
1448 |
0.29 |
|
|
|
| 12 |
A' |
1510 |
1439 |
0.40 |
|
|
|
| 13 |
A' |
1435 |
1368 |
2.18 |
|
|
|
| 14 |
A' |
1403 |
1337 |
1.24 |
|
|
|
| 15 |
A' |
1324 |
1262 |
1.30 |
|
|
|
| 16 |
A' |
1246 |
1188 |
0.60 |
|
|
|
| 17 |
A' |
1216 |
1159 |
0.13 |
|
|
|
| 18 |
A' |
1130 |
1077 |
9.01 |
|
|
|
| 19 |
A' |
1076 |
1025 |
0.96 |
|
|
|
| 20 |
A' |
1069 |
1019 |
2.03 |
|
|
|
| 21 |
A' |
1022 |
974 |
0.31 |
|
|
|
| 22 |
A' |
1001 |
954 |
0.04 |
|
|
|
| 23 |
A' |
933 |
889 |
0.08 |
|
|
|
| 24 |
A' |
914 |
871 |
2.57 |
|
|
|
| 25 |
A' |
834 |
795 |
0.50 |
|
|
|
| 26 |
A' |
768 |
732 |
30.52 |
|
|
|
| 27 |
A' |
718 |
685 |
29.91 |
|
|
|
| 28 |
A' |
598 |
570 |
4.28 |
|
|
|
| 29 |
A' |
510 |
486 |
6.60 |
|
|
|
| 30 |
A' |
312 |
297 |
0.04 |
|
|
|
| 31 |
A' |
279 |
266 |
0.61 |
|
|
|
| 32 |
A' |
104 |
99 |
0.27 |
|
|
|
| 33 |
A" |
3222 |
3071 |
34.62 |
|
|
|
| 34 |
A" |
3198 |
3048 |
6.15 |
|
|
|
| 35 |
A" |
3143 |
2995 |
49.65 |
|
|
|
| 36 |
A" |
3115 |
2969 |
14.08 |
|
|
|
| 37 |
A" |
3093 |
2948 |
0.95 |
|
|
|
| 38 |
A" |
1670 |
1592 |
1.04 |
|
|
|
| 39 |
A" |
1525 |
1454 |
6.77 |
|
|
|
| 40 |
A" |
1509 |
1438 |
7.34 |
|
|
|
| 41 |
A" |
1391 |
1325 |
0.13 |
|
|
|
| 42 |
A" |
1369 |
1305 |
0.47 |
|
|
|
| 43 |
A" |
1335 |
1272 |
0.21 |
|
|
|
| 44 |
A" |
1263 |
1204 |
0.27 |
|
|
|
| 45 |
A" |
1193 |
1137 |
0.03 |
|
|
|
| 46 |
A" |
1137 |
1084 |
4.97 |
|
|
|
| 47 |
A" |
1084 |
1033 |
0.81 |
|
|
|
| 48 |
A" |
973 |
928 |
0.01 |
|
|
|
| 49 |
A" |
883 |
842 |
0.91 |
|
|
|
| 50 |
A" |
863 |
823 |
0.04 |
|
|
|
| 51 |
A" |
750 |
715 |
1.80 |
|
|
|
| 52 |
A" |
635 |
605 |
0.03 |
|
|
|
| 53 |
A" |
416 |
396 |
0.00 |
|
|
|
| 54 |
A" |
344 |
328 |
0.05 |
|
|
|
| 55 |
A" |
245 |
234 |
0.01 |
|
|
|
| 56 |
A" |
90 |
86 |
0.01 |
|
|
|
| 57 |
A" |
33 |
31 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41081.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 39154.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.206 |
|
|
|
| 2 |
C |
-0.169 |
|
|
|
| 3 |
C |
-0.172 |
|
|
|
| 4 |
C |
-0.169 |
|
|
|
| 5 |
C |
-0.206 |
|
|
|
| 6 |
C |
0.133 |
|
|
|
| 7 |
C |
-0.399 |
|
|
|
| 8 |
C |
-0.312 |
|
|
|
| 9 |
C |
-0.521 |
|
|
|
| 10 |
H |
0.161 |
|
|
|
| 11 |
H |
0.167 |
|
|
|
| 12 |
H |
0.166 |
|
|
|
| 13 |
H |
0.167 |
|
|
|
| 14 |
H |
0.161 |
|
|
|
| 15 |
H |
0.172 |
|
|
|
| 16 |
H |
0.172 |
|
|
|
| 17 |
H |
0.171 |
|
|
|
| 18 |
H |
0.171 |
|
|
|
| 19 |
H |
0.175 |
|
|
|
| 20 |
H |
0.170 |
|
|
|
| 21 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.125 |
0.302 |
0.000 |
0.326 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-58.186 |
-0.113 |
0.000 |
| y |
-0.113 |
-51.273 |
0.000 |
| z |
0.000 |
0.000 |
-50.577 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.261 |
-0.113 |
0.000 |
| y |
-0.113 |
3.109 |
0.000 |
| z |
0.000 |
0.000 |
4.153 |
|
| Polar |
|---|
| 3z2-r2 | 8.305 |
|---|
| x2-y2 | -6.913 |
|---|
| xy | -0.113 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.745 |
-1.001 |
0.000 |
| y |
-1.001 |
17.859 |
0.000 |
| z |
0.000 |
0.000 |
13.981 |
<r2> (average value of r
2) Å
2
| <r2> |
437.196 |
| (<r2>)1/2 |
20.909 |