return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-1149.604656
Energy at 298.15K-1149.608691
HF Energy-1148.508339
Nuclear repulsion energy449.428631
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3296 3079 0.00      
2 Ag 1664 1554 0.00      
3 Ag 1225 1145 0.00      
4 Ag 1146 1071 0.00      
5 Ag 773 723 0.00      
6 Ag 336 314 0.00      
7 Au 915 855 0.00      
8 Au 408 381 0.00      
9 B1g 823 769 0.00      
10 B1u 3281 3065 1.00      
11 B1u 1542 1441 96.16      
12 B1u 1147 1071 92.88      
13 B1u 1032 965 51.73      
14 B1u 569 531 23.20      
15 B2g 887 828 0.00      
16 B2g 497 464 0.00      
17 B2g 297 277 0.00      
18 B2u 3294 3078 1.52      
19 B2u 1483 1386 1.61      
20 B2u 1424 1331 3.97      
21 B2u 1148 1072 4.33      
22 B2u 225 210 0.92      
23 B3g 3282 3067 0.00      
24 B3g 1648 1540 0.00      
25 B3g 1337 1249 0.00      
26 B3g 642 600 0.00      
27 B3g 361 337 0.00      
28 B3u 798 745 57.50      
29 B3u 477 446 10.88      
30 B3u 102 96 0.56      

Unscaled Zero Point Vibrational Energy (zpe) 18027.9 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 16845.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.18961 0.02228 0.01994

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.385
C2 0.000 0.000 -1.385
C3 0.000 1.211 0.696
C4 0.000 -1.211 0.696
C5 0.000 -1.211 -0.696
C6 0.000 1.211 -0.696
Cl7 0.000 0.000 3.121
Cl8 0.000 0.000 -3.121
H9 0.000 2.143 1.242
H10 0.000 -2.143 1.242
H11 0.000 -2.143 -1.242
H12 0.000 2.143 -1.242

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.77031.39331.39332.40792.40791.73574.50602.14832.14833.39043.3904
C22.77032.40792.40791.39331.39334.50601.73573.39043.39042.14832.1483
C31.39332.40792.42192.79351.39222.71034.00451.08043.39853.87392.1506
C41.39332.40792.42191.39222.79352.71034.00453.39851.08042.15063.8739
C52.40791.39332.79351.39222.42194.00452.71033.87392.15061.08043.3985
C62.40791.39331.39222.79352.42194.00452.71032.15063.87393.39851.0804
Cl71.73574.50602.71032.71034.00454.00456.24182.85052.85054.86084.8608
Cl84.50601.73574.00454.00452.71032.71036.24184.86084.86082.85052.8505
H92.14833.39041.08043.39853.87392.15062.85054.86084.28694.95442.4835
H102.14833.39043.39851.08042.15063.87392.85054.86084.28692.48354.9544
H113.39042.14833.87392.15061.08043.39854.86082.85054.95442.48354.2869
H123.39042.14832.15063.87393.39851.08044.86082.85052.48354.95444.2869

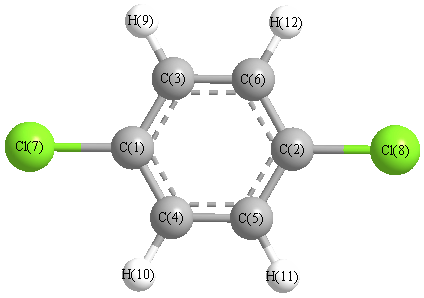
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.640 C1 C3 H9 120.026
C1 C4 C5 119.640 C1 C4 H10 120.026
C2 C5 C4 119.640 C2 C5 H11 120.026
C2 C6 C3 119.640 C2 C6 H12 120.026
C3 C1 C4 120.719 C3 C1 Cl7 119.640
C3 C6 H12 120.334 C4 C1 Cl7 119.640
C4 C5 H11 120.334 C5 C2 C6 120.719
C5 C2 Cl8 119.640 C5 C4 H10 120.334
C6 C2 Cl8 119.640 C6 C3 H9 120.334
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability