Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -5218.431122 |
| Energy at 298.15K | -5218.441356 |
| HF Energy | -5217.854091 |
| Nuclear repulsion energy | 418.345338 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3198 |
2988 |
0.00 |
|
|
|
| 2 |
Ag |
1543 |
1442 |
0.00 |
|
|
|
| 3 |
Ag |
1337 |
1249 |
0.00 |
|
|
|
| 4 |
Ag |
1110 |
1037 |
0.00 |
|
|
|
| 5 |
Ag |
708 |
662 |
0.00 |
|
|
|
| 6 |
Ag |
197 |
184 |
0.00 |
|
|
|
| 7 |
Au |
3294 |
3078 |
0.41 |
|
|
|
| 8 |
Au |
1139 |
1065 |
3.47 |
|
|
|
| 9 |
Au |
771 |
721 |
3.32 |
|
|
|
| 10 |
Au |
109 |
102 |
4.50 |
|
|
|
| 11 |
Bg |
3275 |
3060 |
0.00 |
|
|
|
| 12 |
Bg |
1330 |
1243 |
0.00 |
|
|
|
| 13 |
Bg |
973 |
909 |
0.00 |
|
|
|
| 14 |
Bu |
3205 |
2995 |
5.91 |
|
|
|
| 15 |
Bu |
1537 |
1436 |
5.92 |
|
|
|
| 16 |
Bu |
1249 |
1167 |
58.99 |
|
|
|
| 17 |
Bu |
631 |
590 |
53.69 |
|
|
|
| 18 |
Bu |
184 |
172 |
6.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12895.1 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 12049.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.493 |
0.569 |
0.000 |
| C2 |
-0.493 |
-0.569 |
0.000 |
| Br3 |
-0.493 |
2.257 |
0.000 |
| Br4 |
0.493 |
-2.257 |
0.000 |
| H5 |
1.114 |
0.566 |
0.889 |
| H6 |
1.114 |
0.566 |
-0.889 |
| H7 |
-1.114 |
-0.566 |
0.889 |
| H8 |
-1.114 |
-0.566 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5062 | 1.9557 | 2.8261 | 1.0842 | 1.0842 | 2.1592 | 2.1592 |
C2 | 1.5062 | | 2.8261 | 1.9557 | 2.1592 | 2.1592 | 1.0842 | 1.0842 | Br3 | 1.9557 | 2.8261 | | 4.6211 | 2.4966 | 2.4966 | 3.0245 | 3.0245 | Br4 | 2.8261 | 1.9557 | 4.6211 | | 3.0245 | 3.0245 | 2.4966 | 2.4966 | H5 | 1.0842 | 2.1592 | 2.4966 | 3.0245 | | 1.7786 | 2.4987 | 3.0670 | H6 | 1.0842 | 2.1592 | 2.4966 | 3.0245 | 1.7786 | | 3.0670 | 2.4987 | H7 | 2.1592 | 1.0842 | 3.0245 | 2.4966 | 2.4987 | 3.0670 | | 1.7786 | H8 | 2.1592 | 1.0842 | 3.0245 | 2.4966 | 3.0670 | 2.4987 | 1.7786 | |
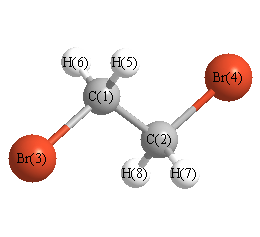 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.750 |
|
C1 |
C2 |
H7 |
111.900 |
| C1 |
C2 |
H8 |
111.900 |
|
C2 |
C1 |
Br3 |
108.750 |
| C2 |
C1 |
H5 |
111.900 |
|
C2 |
C1 |
H6 |
111.900 |
| Br3 |
C1 |
H5 |
106.905 |
|
Br3 |
C1 |
H6 |
106.905 |
| Br4 |
C2 |
H7 |
106.905 |
|
Br4 |
C2 |
H8 |
106.905 |
| H5 |
C1 |
H6 |
110.217 |
|
H7 |
C2 |
H8 |
110.217 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -5218.429102 |
| Energy at 298.15K | |
| HF Energy | -5217.850610 |
| Nuclear repulsion energy | 458.051799 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3258 |
3044 |
0.08 |
87.91 |
0.74 |
0.85 |
| 2 |
A |
3180 |
2972 |
13.50 |
171.24 |
0.02 |
0.04 |
| 3 |
A |
1524 |
1424 |
0.00 |
5.66 |
0.72 |
0.84 |
| 4 |
A |
1357 |
1268 |
24.47 |
3.01 |
0.63 |
0.77 |
| 5 |
A |
1233 |
1152 |
2.30 |
14.36 |
0.75 |
0.86 |
| 6 |
A |
1070 |
999 |
0.16 |
3.73 |
0.74 |
0.85 |
| 7 |
A |
945 |
883 |
5.53 |
5.85 |
0.47 |
0.64 |
| 8 |
A |
587 |
548 |
7.93 |
16.89 |
0.08 |
0.16 |
| 9 |
A |
234 |
219 |
0.94 |
0.91 |
0.33 |
0.50 |
| 10 |
A |
95 |
89 |
0.18 |
0.79 |
0.71 |
0.83 |
| 11 |
B |
3268 |
3054 |
0.96 |
28.87 |
0.75 |
0.86 |
| 12 |
B |
3175 |
2967 |
1.93 |
44.91 |
0.75 |
0.86 |
| 13 |
B |
1516 |
1416 |
8.75 |
16.39 |
0.75 |
0.86 |
| 14 |
B |
1326 |
1239 |
56.22 |
0.57 |
0.75 |
0.86 |
| 15 |
B |
1158 |
1082 |
2.54 |
5.22 |
0.75 |
0.86 |
| 16 |
B |
872 |
815 |
13.42 |
0.92 |
0.75 |
0.86 |
| 17 |
B |
620 |
579 |
8.84 |
9.04 |
0.75 |
0.86 |
| 18 |
B |
366 |
342 |
5.87 |
2.35 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12891.5 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 12045.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.307 |
0.689 |
1.214 |
| C2 |
-0.307 |
-0.689 |
1.214 |
| Br3 |
-0.307 |
1.745 |
-0.301 |
| Br4 |
0.307 |
-1.745 |
-0.301 |
| H5 |
0.008 |
1.231 |
2.108 |
| H6 |
1.388 |
0.645 |
1.148 |
| H7 |
-0.008 |
-1.231 |
2.108 |
| H8 |
-1.388 |
-0.645 |
1.148 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5075 | 1.9465 | 2.8672 | 1.0871 | 1.0848 | 2.1408 | 2.1574 |
C2 | 1.5075 | | 2.8672 | 1.9465 | 2.1408 | 2.1574 | 1.0871 | 1.0848 | Br3 | 1.9465 | 2.8672 | | 3.5444 | 2.4832 | 2.4869 | 3.8408 | 2.9971 | Br4 | 2.8672 | 1.9465 | 3.5444 | | 3.8408 | 2.9971 | 2.4832 | 2.4869 | H5 | 1.0871 | 2.1408 | 2.4832 | 3.8408 | | 1.7806 | 2.4622 | 2.5277 | H6 | 1.0848 | 2.1574 | 2.4869 | 2.9971 | 1.7806 | | 2.5277 | 3.0614 | H7 | 2.1408 | 1.0871 | 3.8408 | 2.4832 | 2.4622 | 2.5277 | | 1.7806 | H8 | 2.1574 | 1.0848 | 2.9971 | 2.4869 | 2.5277 | 3.0614 | 1.7806 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
111.589 |
|
C1 |
C2 |
H7 |
110.136 |
| C1 |
C2 |
H8 |
111.619 |
|
C2 |
C1 |
Br3 |
111.589 |
| C2 |
C1 |
H5 |
110.136 |
|
C2 |
C1 |
H6 |
111.619 |
| Br3 |
C1 |
H5 |
106.410 |
|
Br3 |
C1 |
H6 |
106.780 |
| Br4 |
C2 |
H7 |
106.410 |
|
Br4 |
C2 |
H8 |
106.780 |
| H5 |
C1 |
H6 |
110.131 |
|
H7 |
C2 |
H8 |
110.131 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
