Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -5221.277227 |
| Energy at 298.15K | |
| HF Energy | -5221.277227 |
| Nuclear repulsion energy | 418.513900 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3152 |
3004 |
0.00 |
138.72 |
0.07 |
0.12 |
| 2 |
Ag |
1514 |
1443 |
0.00 |
14.65 |
0.74 |
0.85 |
| 3 |
Ag |
1306 |
1245 |
0.00 |
30.60 |
0.46 |
0.63 |
| 4 |
Ag |
1096 |
1045 |
0.00 |
10.93 |
0.70 |
0.82 |
| 5 |
Ag |
687 |
654 |
0.00 |
79.74 |
0.28 |
0.44 |
| 6 |
Ag |
194 |
185 |
0.00 |
4.04 |
0.30 |
0.46 |
| 7 |
Au |
3240 |
3088 |
0.31 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1115 |
1063 |
3.37 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
762 |
726 |
5.04 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
105 |
100 |
4.47 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3218 |
3067 |
0.00 |
89.69 |
0.75 |
0.86 |
| 12 |
Bg |
1305 |
1244 |
0.00 |
7.38 |
0.75 |
0.86 |
| 13 |
Bg |
949 |
905 |
0.00 |
7.25 |
0.75 |
0.86 |
| 14 |
Bu |
3159 |
3011 |
5.44 |
0.00 |
0.07 |
0.00 |
| 15 |
Bu |
1506 |
1435 |
8.19 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1225 |
1168 |
54.08 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
618 |
589 |
71.59 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
181 |
173 |
7.29 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 12665.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12071.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.493 |
0.569 |
0.000 |
| C2 |
-0.493 |
-0.569 |
0.000 |
| Br3 |
-0.493 |
2.255 |
0.000 |
| Br4 |
0.493 |
-2.255 |
0.000 |
| H5 |
1.117 |
0.571 |
0.893 |
| H6 |
1.117 |
0.571 |
-0.893 |
| H7 |
-1.117 |
-0.571 |
0.893 |
| H8 |
-1.117 |
-0.571 |
-0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5059 | 1.9537 | 2.8244 | 1.0895 | 1.0895 | 2.1657 | 2.1657 |
C2 | 1.5059 | | 2.8244 | 1.9537 | 2.1657 | 2.1657 | 1.0895 | 1.0895 | Br3 | 1.9537 | 2.8244 | | 4.6174 | 2.4960 | 2.4960 | 3.0288 | 3.0288 | Br4 | 2.8244 | 1.9537 | 4.6174 | | 3.0288 | 3.0288 | 2.4960 | 2.4960 | H5 | 1.0895 | 2.1657 | 2.4960 | 3.0288 | | 1.7858 | 2.5095 | 3.0801 | H6 | 1.0895 | 2.1657 | 2.4960 | 3.0288 | 1.7858 | | 3.0801 | 2.5095 | H7 | 2.1657 | 1.0895 | 3.0288 | 2.4960 | 2.5095 | 3.0801 | | 1.7858 | H8 | 2.1657 | 1.0895 | 3.0288 | 2.4960 | 3.0801 | 2.5095 | 1.7858 | |
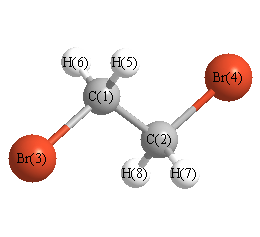 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.763 |
|
C1 |
C2 |
H7 |
112.116 |
| C1 |
C2 |
H8 |
112.116 |
|
C2 |
C1 |
Br3 |
108.763 |
| C2 |
C1 |
H5 |
112.116 |
|
C2 |
C1 |
H6 |
112.116 |
| Br3 |
C1 |
H5 |
106.735 |
|
Br3 |
C1 |
H6 |
106.735 |
| Br4 |
C2 |
H7 |
106.735 |
|
Br4 |
C2 |
H8 |
106.735 |
| H5 |
C1 |
H6 |
110.075 |
|
H7 |
C2 |
H8 |
110.075 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.359 |
|
|
|
| 2 |
C |
-0.359 |
|
|
|
| 3 |
Br |
-0.120 |
|
|
|
| 4 |
Br |
-0.120 |
|
|
|
| 5 |
H |
0.239 |
|
|
|
| 6 |
H |
0.239 |
|
|
|
| 7 |
H |
0.239 |
|
|
|
| 8 |
H |
0.239 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.095 |
1.213 |
0.000 |
| y |
1.213 |
-54.962 |
0.000 |
| z |
0.000 |
0.000 |
-48.850 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.811 |
1.213 |
0.000 |
| y |
1.213 |
-6.489 |
0.000 |
| z |
0.000 |
0.000 |
2.678 |
|
| Polar |
|---|
| 3z2-r2 | 5.356 |
|---|
| x2-y2 | 6.867 |
|---|
| xy | 1.213 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.768 |
-1.798 |
0.000 |
| y |
-1.798 |
11.981 |
0.000 |
| z |
0.000 |
0.000 |
5.888 |
<r2> (average value of r
2) Å
2
| <r2> |
421.026 |
| (<r2>)1/2 |
20.519 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -5221.274824 |
| Energy at 298.15K | |
| HF Energy | -5221.274824 |
| Nuclear repulsion energy | 457.271169 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3199 |
3049 |
0.02 |
97.37 |
0.75 |
0.86 |
| 2 |
A |
3133 |
2986 |
13.78 |
201.79 |
0.02 |
0.04 |
| 3 |
A |
1494 |
1424 |
0.44 |
5.76 |
0.69 |
0.82 |
| 4 |
A |
1331 |
1269 |
24.43 |
4.18 |
0.63 |
0.78 |
| 5 |
A |
1213 |
1156 |
2.04 |
16.22 |
0.75 |
0.86 |
| 6 |
A |
1052 |
1003 |
0.57 |
3.72 |
0.69 |
0.81 |
| 7 |
A |
924 |
881 |
7.49 |
6.54 |
0.46 |
0.63 |
| 8 |
A |
579 |
551 |
9.20 |
17.10 |
0.08 |
0.15 |
| 9 |
A |
230 |
219 |
1.07 |
0.93 |
0.30 |
0.46 |
| 10 |
A |
88 |
84 |
0.18 |
0.77 |
0.72 |
0.84 |
| 11 |
B |
3212 |
3062 |
0.98 |
30.61 |
0.75 |
0.86 |
| 12 |
B |
3126 |
2979 |
1.73 |
54.30 |
0.75 |
0.86 |
| 13 |
B |
1484 |
1415 |
12.33 |
17.25 |
0.75 |
0.86 |
| 14 |
B |
1296 |
1235 |
56.75 |
0.56 |
0.75 |
0.86 |
| 15 |
B |
1138 |
1085 |
2.29 |
5.54 |
0.75 |
0.86 |
| 16 |
B |
854 |
814 |
18.71 |
0.95 |
0.75 |
0.86 |
| 17 |
B |
610 |
581 |
11.96 |
8.89 |
0.75 |
0.86 |
| 18 |
B |
362 |
345 |
6.97 |
2.39 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12662.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12069.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.303 |
0.690 |
1.209 |
| C2 |
-0.303 |
-0.690 |
1.209 |
| Br3 |
-0.303 |
1.753 |
-0.300 |
| Br4 |
0.303 |
-1.753 |
-0.300 |
| H5 |
-0.001 |
1.232 |
2.107 |
| H6 |
1.391 |
0.655 |
1.149 |
| H7 |
0.001 |
-1.232 |
2.107 |
| H8 |
-1.391 |
-0.655 |
1.149 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5066 | 1.9432 | 2.8714 | 1.0928 | 1.0901 | 2.1431 | 2.1634 |
C2 | 1.5066 | | 2.8714 | 1.9432 | 2.1431 | 2.1634 | 1.0928 | 1.0901 | Br3 | 1.9432 | 2.8714 | | 3.5585 | 2.4818 | 2.4851 | 3.8474 | 3.0133 | Br4 | 2.8714 | 1.9432 | 3.5585 | | 3.8474 | 3.0133 | 2.4818 | 2.4851 | H5 | 1.0928 | 2.1431 | 2.4818 | 3.8474 | | 1.7861 | 2.4647 | 2.5323 | H6 | 1.0901 | 2.1634 | 2.4851 | 3.0133 | 1.7861 | | 2.5323 | 3.0746 | H7 | 2.1431 | 1.0928 | 3.8474 | 2.4818 | 2.4647 | 2.5323 | | 1.7861 | H8 | 2.1634 | 1.0901 | 3.0133 | 2.4851 | 2.5323 | 3.0746 | 1.7861 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.060 |
|
C1 |
C2 |
H7 |
110.047 |
| C1 |
C2 |
H8 |
111.844 |
|
C2 |
C1 |
Br3 |
112.060 |
| C2 |
C1 |
H5 |
110.047 |
|
C2 |
C1 |
H6 |
111.844 |
| Br3 |
C1 |
H5 |
106.259 |
|
Br3 |
C1 |
H6 |
106.615 |
| Br4 |
C2 |
H7 |
106.259 |
|
Br4 |
C2 |
H8 |
106.615 |
| H5 |
C1 |
H6 |
109.812 |
|
H7 |
C2 |
H8 |
109.812 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.362 |
|
|
|
| 2 |
C |
-0.362 |
|
|
|
| 3 |
Br |
-0.105 |
|
|
|
| 4 |
Br |
-0.105 |
|
|
|
| 5 |
H |
0.228 |
|
|
|
| 6 |
H |
0.239 |
|
|
|
| 7 |
H |
0.228 |
|
|
|
| 8 |
H |
0.239 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.675 |
2.675 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.575 |
0.857 |
0.000 |
| y |
0.857 |
-52.912 |
0.000 |
| z |
0.000 |
0.000 |
-44.069 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.085 |
0.857 |
0.000 |
| y |
0.857 |
-6.590 |
0.000 |
| z |
0.000 |
0.000 |
6.674 |
|
| Polar |
|---|
| 3z2-r2 | 13.349 |
|---|
| x2-y2 | 4.337 |
|---|
| xy | 0.857 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.172 |
-0.715 |
0.000 |
| y |
-0.715 |
9.025 |
0.000 |
| z |
0.000 |
0.000 |
8.110 |
<r2> (average value of r
2) Å
2
| <r2> |
301.847 |
| (<r2>)1/2 |
17.374 |
