Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -5221.239538 |
| Energy at 298.15K | -5221.249695 |
| HF Energy | -5221.239538 |
| Nuclear repulsion energy | 418.324255 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3140 |
2997 |
0.00 |
|
|
|
| 2 |
Ag |
1498 |
1430 |
0.00 |
|
|
|
| 3 |
Ag |
1293 |
1233 |
0.00 |
|
|
|
| 4 |
Ag |
1092 |
1042 |
0.00 |
|
|
|
| 5 |
Ag |
681 |
650 |
0.00 |
|
|
|
| 6 |
Ag |
192 |
183 |
0.00 |
|
|
|
| 7 |
Au |
3231 |
3084 |
0.28 |
|
|
|
| 8 |
Au |
1108 |
1057 |
3.23 |
|
|
|
| 9 |
Au |
758 |
723 |
4.89 |
|
|
|
| 10 |
Au |
105 |
100 |
4.43 |
|
|
|
| 11 |
Bg |
3210 |
3063 |
0.00 |
|
|
|
| 12 |
Bg |
1294 |
1235 |
0.00 |
|
|
|
| 13 |
Bg |
943 |
899 |
0.00 |
|
|
|
| 14 |
Bu |
3148 |
3004 |
5.41 |
|
|
|
| 15 |
Bu |
1489 |
1421 |
7.84 |
|
|
|
| 16 |
Bu |
1212 |
1157 |
52.22 |
|
|
|
| 17 |
Bu |
610 |
582 |
71.79 |
|
|
|
| 18 |
Bu |
180 |
171 |
7.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12591.4 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12016.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.493 |
0.568 |
0.000 |
| C2 |
-0.493 |
-0.568 |
0.000 |
| Br3 |
-0.493 |
2.257 |
0.000 |
| Br4 |
0.493 |
-2.257 |
0.000 |
| H5 |
1.117 |
0.571 |
0.893 |
| H6 |
1.117 |
0.571 |
-0.893 |
| H7 |
-1.117 |
-0.571 |
0.893 |
| H8 |
-1.117 |
-0.571 |
-0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5048 | 1.9553 | 2.8252 | 1.0891 | 1.0891 | 2.1648 | 2.1648 |
C2 | 1.5048 | | 2.8252 | 1.9553 | 2.1648 | 2.1648 | 1.0891 | 1.0891 | Br3 | 1.9553 | 2.8252 | | 4.6201 | 2.4962 | 2.4962 | 3.0301 | 3.0301 | Br4 | 2.8252 | 1.9553 | 4.6201 | | 3.0301 | 3.0301 | 2.4962 | 2.4962 | H5 | 1.0891 | 2.1648 | 2.4962 | 3.0301 | | 1.7853 | 2.5085 | 3.0790 | H6 | 1.0891 | 2.1648 | 2.4962 | 3.0301 | 1.7853 | | 3.0790 | 2.5085 | H7 | 2.1648 | 1.0891 | 3.0301 | 2.4962 | 2.5085 | 3.0790 | | 1.7853 | H8 | 2.1648 | 1.0891 | 3.0301 | 2.4962 | 3.0790 | 2.5085 | 1.7853 | |
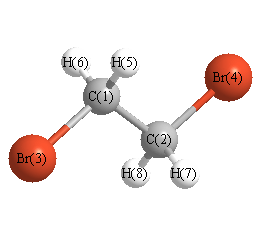 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.777 |
|
C1 |
C2 |
H7 |
112.151 |
| C1 |
C2 |
H8 |
112.151 |
|
C2 |
C1 |
Br3 |
108.777 |
| C2 |
C1 |
H5 |
112.151 |
|
C2 |
C1 |
H6 |
112.151 |
| Br3 |
C1 |
H5 |
106.675 |
|
Br3 |
C1 |
H6 |
106.675 |
| Br4 |
C2 |
H7 |
106.675 |
|
Br4 |
C2 |
H8 |
106.675 |
| H5 |
C1 |
H6 |
110.102 |
|
H7 |
C2 |
H8 |
110.102 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.267 |
|
|
|
| 2 |
C |
-0.267 |
|
|
|
| 3 |
Br |
-0.128 |
|
|
|
| 4 |
Br |
-0.128 |
|
|
|
| 5 |
H |
0.197 |
|
|
|
| 6 |
H |
0.197 |
|
|
|
| 7 |
H |
0.197 |
|
|
|
| 8 |
H |
0.197 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.106 |
1.217 |
0.000 |
| y |
1.217 |
-54.952 |
0.000 |
| z |
0.000 |
0.000 |
-48.885 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.813 |
1.217 |
0.000 |
| y |
1.217 |
-6.456 |
0.000 |
| z |
0.000 |
0.000 |
2.644 |
|
| Polar |
|---|
| 3z2-r2 | 5.287 |
|---|
| x2-y2 | 6.846 |
|---|
| xy | 1.217 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.822 |
-1.808 |
0.000 |
| y |
-1.808 |
12.106 |
0.000 |
| z |
0.000 |
0.000 |
5.935 |
<r2> (average value of r
2) Å
2
| <r2> |
421.444 |
| (<r2>)1/2 |
20.529 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -5221.237112 |
| Energy at 298.15K | |
| HF Energy | -5221.237112 |
| Nuclear repulsion energy | 457.169460 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3191 |
3045 |
0.03 |
98.15 |
0.75 |
0.86 |
| 2 |
A |
3122 |
2979 |
13.62 |
209.76 |
0.02 |
0.03 |
| 3 |
A |
1475 |
1408 |
0.46 |
5.63 |
0.69 |
0.81 |
| 4 |
A |
1318 |
1258 |
23.55 |
3.96 |
0.62 |
0.77 |
| 5 |
A |
1203 |
1148 |
1.97 |
16.03 |
0.75 |
0.86 |
| 6 |
A |
1047 |
999 |
0.61 |
3.71 |
0.68 |
0.81 |
| 7 |
A |
917 |
875 |
7.32 |
6.55 |
0.46 |
0.63 |
| 8 |
A |
575 |
548 |
9.22 |
17.26 |
0.08 |
0.15 |
| 9 |
A |
228 |
217 |
1.10 |
0.96 |
0.30 |
0.46 |
| 10 |
A |
88 |
84 |
0.18 |
0.79 |
0.72 |
0.84 |
| 11 |
B |
3204 |
3058 |
0.86 |
31.80 |
0.75 |
0.86 |
| 12 |
B |
3115 |
2972 |
1.82 |
54.12 |
0.75 |
0.86 |
| 13 |
B |
1465 |
1398 |
12.17 |
17.14 |
0.75 |
0.86 |
| 14 |
B |
1282 |
1223 |
55.80 |
0.64 |
0.75 |
0.86 |
| 15 |
B |
1130 |
1078 |
2.17 |
5.43 |
0.75 |
0.86 |
| 16 |
B |
848 |
809 |
18.36 |
0.98 |
0.75 |
0.86 |
| 17 |
B |
605 |
577 |
11.97 |
9.01 |
0.75 |
0.86 |
| 18 |
B |
359 |
343 |
7.15 |
2.39 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12584.4 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12009.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.304 |
0.689 |
1.210 |
| C2 |
-0.304 |
-0.689 |
1.210 |
| Br3 |
-0.304 |
1.753 |
-0.300 |
| Br4 |
0.304 |
-1.753 |
-0.300 |
| H5 |
0.002 |
1.233 |
2.107 |
| H6 |
1.391 |
0.652 |
1.146 |
| H7 |
-0.002 |
-1.233 |
2.107 |
| H8 |
-1.391 |
-0.652 |
1.146 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5054 | 1.9452 | 2.8711 | 1.0922 | 1.0896 | 2.1428 | 2.1623 |
C2 | 1.5054 | | 2.8711 | 1.9452 | 2.1428 | 2.1623 | 1.0922 | 1.0896 | Br3 | 1.9452 | 2.8711 | | 3.5589 | 2.4819 | 2.4854 | 3.8479 | 3.0101 | Br4 | 2.8711 | 1.9452 | 3.5589 | | 3.8479 | 3.0101 | 2.4819 | 2.4854 | H5 | 1.0922 | 2.1428 | 2.4819 | 3.8479 | | 1.7864 | 2.4663 | 2.5337 | H6 | 1.0896 | 2.1623 | 2.4854 | 3.0101 | 1.7864 | | 2.5337 | 3.0730 | H7 | 2.1428 | 1.0922 | 3.8479 | 2.4819 | 2.4663 | 2.5337 | | 1.7864 | H8 | 2.1623 | 1.0896 | 3.0101 | 2.4854 | 2.5337 | 3.0730 | 1.7864 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
111.993 |
|
C1 |
C2 |
H7 |
110.147 |
| C1 |
C2 |
H8 |
111.873 |
|
C2 |
C1 |
Br3 |
111.993 |
| C2 |
C1 |
H5 |
110.147 |
|
C2 |
C1 |
H6 |
111.873 |
| Br3 |
C1 |
H5 |
106.168 |
|
Br3 |
C1 |
H6 |
106.525 |
| Br4 |
C2 |
H7 |
106.168 |
|
Br4 |
C2 |
H8 |
106.525 |
| H5 |
C1 |
H6 |
109.922 |
|
H7 |
C2 |
H8 |
109.922 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.269 |
|
|
|
| 2 |
C |
-0.269 |
|
|
|
| 3 |
Br |
-0.113 |
|
|
|
| 4 |
Br |
-0.113 |
|
|
|
| 5 |
H |
0.185 |
|
|
|
| 6 |
H |
0.197 |
|
|
|
| 7 |
H |
0.185 |
|
|
|
| 8 |
H |
0.197 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.668 |
2.668 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.595 |
0.866 |
0.000 |
| y |
0.866 |
-52.910 |
0.000 |
| z |
0.000 |
0.000 |
-44.106 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.087 |
0.866 |
0.000 |
| y |
0.866 |
-6.559 |
0.000 |
| z |
0.000 |
0.000 |
6.646 |
|
| Polar |
|---|
| 3z2-r2 | 13.291 |
|---|
| x2-y2 | 4.315 |
|---|
| xy | 0.866 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.222 |
-0.718 |
0.000 |
| y |
-0.718 |
9.107 |
0.000 |
| z |
0.000 |
0.000 |
8.179 |
<r2> (average value of r
2) Å
2
| <r2> |
301.909 |
| (<r2>)1/2 |
17.376 |
