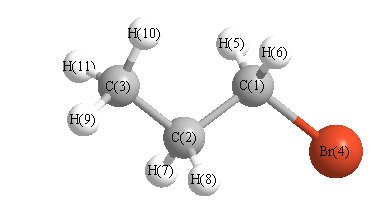
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3155 |
3011 |
22.83 |
|
|
|
| 2 |
A' |
3120 |
2978 |
16.10 |
|
|
|
| 3 |
A' |
3081 |
2940 |
10.38 |
|
|
|
| 4 |
A' |
3064 |
2924 |
21.80 |
|
|
|
| 5 |
A' |
1517 |
1448 |
6.04 |
|
|
|
| 6 |
A' |
1502 |
1434 |
1.23 |
|
|
|
| 7 |
A' |
1493 |
1425 |
1.77 |
|
|
|
| 8 |
A' |
1424 |
1359 |
2.07 |
|
|
|
| 9 |
A' |
1371 |
1309 |
6.55 |
|
|
|
| 10 |
A' |
1256 |
1199 |
38.92 |
|
|
|
| 11 |
A' |
1129 |
1077 |
1.73 |
|
|
|
| 12 |
A' |
1062 |
1014 |
1.48 |
|
|
|
| 13 |
A' |
917 |
876 |
10.63 |
|
|
|
| 14 |
A' |
665 |
635 |
23.30 |
|
|
|
| 15 |
A' |
312 |
298 |
1.81 |
|
|
|
| 16 |
A' |
213 |
203 |
1.50 |
|
|
|
| 17 |
A" |
3191 |
3046 |
10.16 |
|
|
|
| 18 |
A" |
3146 |
3003 |
32.13 |
|
|
|
| 19 |
A" |
3121 |
2979 |
0.18 |
|
|
|
| 20 |
A" |
1509 |
1440 |
8.77 |
|
|
|
| 21 |
A" |
1323 |
1263 |
0.01 |
|
|
|
| 22 |
A" |
1246 |
1189 |
0.23 |
|
|
|
| 23 |
A" |
1064 |
1015 |
2.22 |
|
|
|
| 24 |
A" |
856 |
817 |
0.06 |
|
|
|
| 25 |
A" |
749 |
714 |
4.93 |
|
|
|
| 26 |
A" |
241 |
230 |
0.03 |
|
|
|
| 27 |
A" |
119 |
113 |
0.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20923.8 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 19967.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.261 |
|
|
|
| 2 |
C |
-0.242 |
|
|
|
| 3 |
C |
-0.402 |
|
|
|
| 4 |
Br |
-0.152 |
|
|
|
| 5 |
H |
0.173 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
H |
0.149 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
| 9 |
H |
0.143 |
|
|
|
| 10 |
H |
0.135 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.317 |
1.818 |
0.000 |
2.245 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.219 |
0.115 |
0.000 |
| y |
0.115 |
-37.329 |
0.000 |
| z |
0.000 |
0.000 |
-38.209 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.450 |
0.115 |
0.000 |
| y |
0.115 |
0.884 |
0.000 |
| z |
0.000 |
0.000 |
-0.435 |
|
| Polar |
|---|
| 3z2-r2 | -0.869 |
|---|
| x2-y2 | -0.890 |
|---|
| xy | 0.115 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.846 |
1.716 |
0.000 |
| y |
1.716 |
8.133 |
0.000 |
| z |
0.000 |
0.000 |
6.021 |
<r2> (average value of r
2) Å
2
| <r2> |
205.984 |
| (<r2>)1/2 |
14.352 |