Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -158.250179 |
| Energy at 298.15K | -158.260831 |
| HF Energy | -158.250179 |
| Nuclear repulsion energy | 130.659969 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
127 |
121 |
0.01 |
|
|
|
|
Au |
223 |
212 |
0.00 |
|
|
|
|
Bu |
257 |
245 |
0.06 |
|
|
|
|
Bg |
263 |
251 |
0.00 |
|
|
|
|
Ag |
428 |
408 |
0.00 |
|
|
|
|
Au |
743 |
708 |
4.84 |
|
|
|
|
Bg |
819 |
780 |
0.00 |
|
|
|
|
Ag |
861 |
821 |
0.00 |
|
|
|
|
Au |
972 |
926 |
1.60 |
|
|
|
|
Bu |
990 |
944 |
8.41 |
|
|
|
|
Bu |
1051 |
1002 |
0.17 |
|
|
|
|
Ag |
1098 |
1047 |
0.00 |
|
|
|
|
Ag |
1188 |
1132 |
0.00 |
|
|
|
|
Bg |
1223 |
1166 |
0.00 |
|
|
|
|
Au |
1306 |
1244 |
0.06 |
|
|
|
|
Bu |
1334 |
1272 |
2.82 |
|
|
|
|
Bg |
1349 |
1286 |
0.00 |
|
|
|
|
Ag |
1420 |
1353 |
0.00 |
|
|
|
|
Ag |
1438 |
1371 |
0.00 |
|
|
|
|
Bu |
1439 |
1371 |
6.44 |
|
|
|
|
Ag |
1514 |
1443 |
0.00 |
|
|
|
|
Bu |
1518 |
1447 |
1.79 |
|
|
|
|
Bg |
1526 |
1454 |
0.00 |
|
|
|
|
Au |
1527 |
1456 |
15.48 |
|
|
|
|
Ag |
1533 |
1461 |
0.00 |
|
|
|
|
Bu |
1539 |
1467 |
8.00 |
|
|
|
|
Ag |
3049 |
2906 |
0.00 |
|
|
|
|
Bu |
3057 |
2913 |
43.32 |
|
|
|
|
Bu |
3065 |
2922 |
76.29 |
|
|
|
|
Ag |
3066 |
2922 |
0.00 |
|
|
|
|
Bg |
3078 |
2934 |
0.00 |
|
|
|
|
Au |
3101 |
2955 |
24.89 |
|
|
|
|
Bg |
3139 |
2992 |
0.00 |
|
|
|
|
Au |
3143 |
2995 |
100.64 |
|
|
|
|
Ag |
3147 |
3000 |
0.00 |
|
|
|
|
Bu |
3148 |
3000 |
69.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29339.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 27963.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.420 |
0.637 |
0.000 |
| C2 |
0.420 |
-0.637 |
0.000 |
| C3 |
0.420 |
1.908 |
0.000 |
| C4 |
-0.420 |
-1.908 |
0.000 |
| H5 |
-1.082 |
0.632 |
0.878 |
| H6 |
-1.082 |
0.632 |
-0.878 |
| H7 |
1.082 |
-0.632 |
0.878 |
| H8 |
1.082 |
-0.632 |
-0.878 |
| H9 |
-0.207 |
2.806 |
0.000 |
| H10 |
1.067 |
1.953 |
0.884 |
| H11 |
1.067 |
1.953 |
-0.884 |
| H12 |
0.207 |
-2.806 |
0.000 |
| H13 |
-1.067 |
-1.953 |
0.884 |
| H14 |
-1.067 |
-1.953 |
-0.884 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5258 | 1.5237 | 2.5444 | 1.0992 | 1.0992 | 2.1531 | 2.1531 | 2.1797 | 2.1743 | 2.1743 | 3.4993 | 2.8121 | 2.8121 |
C2 | 1.5258 | | 2.5444 | 1.5237 | 2.1531 | 2.1531 | 1.0992 | 1.0992 | 3.4993 | 2.8121 | 2.8121 | 2.1797 | 2.1743 | 2.1743 | C3 | 1.5237 | 2.5444 | | 3.9067 | 2.1575 | 2.1575 | 2.7670 | 2.7670 | 1.0955 | 1.0966 | 1.0966 | 4.7185 | 4.2308 | 4.2308 | C4 | 2.5444 | 1.5237 | 3.9067 | | 2.7670 | 2.7670 | 2.1575 | 2.1575 | 4.7185 | 4.2308 | 4.2308 | 1.0955 | 1.0966 | 1.0966 | H5 | 1.0992 | 2.1531 | 2.1575 | 2.7670 | | 1.7552 | 2.5059 | 3.0594 | 2.5028 | 2.5231 | 3.0773 | 3.7747 | 2.5849 | 3.1281 | H6 | 1.0992 | 2.1531 | 2.1575 | 2.7670 | 1.7552 | | 3.0594 | 2.5059 | 2.5028 | 3.0773 | 2.5231 | 3.7747 | 3.1281 | 2.5849 | H7 | 2.1531 | 1.0992 | 2.7670 | 2.1575 | 2.5059 | 3.0594 | | 1.7552 | 3.7747 | 2.5849 | 3.1281 | 2.5028 | 2.5231 | 3.0773 | H8 | 2.1531 | 1.0992 | 2.7670 | 2.1575 | 3.0594 | 2.5059 | 1.7552 | | 3.7747 | 3.1281 | 2.5849 | 2.5028 | 3.0773 | 2.5231 | H9 | 2.1797 | 3.4993 | 1.0955 | 4.7185 | 2.5028 | 2.5028 | 3.7747 | 3.7747 | | 1.7698 | 1.7698 | 5.6272 | 4.9165 | 4.9165 | H10 | 2.1743 | 2.8121 | 1.0966 | 4.2308 | 2.5231 | 3.0773 | 2.5849 | 3.1281 | 1.7698 | | 1.7683 | 4.9165 | 4.4515 | 4.7899 | H11 | 2.1743 | 2.8121 | 1.0966 | 4.2308 | 3.0773 | 2.5231 | 3.1281 | 2.5849 | 1.7698 | 1.7683 | | 4.9165 | 4.7899 | 4.4515 | H12 | 3.4993 | 2.1797 | 4.7185 | 1.0955 | 3.7747 | 3.7747 | 2.5028 | 2.5028 | 5.6272 | 4.9165 | 4.9165 | | 1.7698 | 1.7698 | H13 | 2.8121 | 2.1743 | 4.2308 | 1.0966 | 2.5849 | 3.1281 | 2.5231 | 3.0773 | 4.9165 | 4.4515 | 4.7899 | 1.7698 | | 1.7683 | H14 | 2.8121 | 2.1743 | 4.2308 | 1.0966 | 3.1281 | 2.5849 | 3.0773 | 2.5231 | 4.9165 | 4.7899 | 4.4515 | 1.7698 | 1.7683 | |
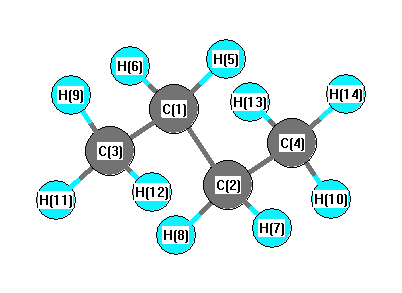 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.099 |
|
C1 |
C2 |
H7 |
109.137 |
| C1 |
C2 |
H8 |
109.137 |
|
C1 |
C3 |
H9 |
111.614 |
| C1 |
C3 |
H11 |
111.107 |
|
C1 |
C3 |
H12 |
30.883 |
| C2 |
C1 |
C3 |
113.099 |
|
C2 |
C1 |
H5 |
109.137 |
| C2 |
C1 |
H6 |
109.137 |
|
C2 |
C4 |
H10 |
17.236 |
| C2 |
C4 |
H13 |
111.107 |
|
C2 |
C4 |
H14 |
111.107 |
| C3 |
C1 |
H5 |
109.629 |
|
C3 |
C1 |
H6 |
109.629 |
| C4 |
C2 |
H7 |
109.629 |
|
C4 |
C2 |
H8 |
109.629 |
| H5 |
C1 |
H6 |
105.960 |
|
H7 |
C2 |
H8 |
105.960 |
| H9 |
C3 |
H11 |
107.681 |
|
H9 |
C3 |
H12 |
142.497 |
| H10 |
C4 |
H13 |
94.408 |
|
H10 |
C4 |
H14 |
114.445 |
| H11 |
C3 |
H12 |
93.907 |
|
H13 |
C4 |
H14 |
107.459 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.310 |
|
|
|
| 2 |
C |
-0.310 |
|
|
|
| 3 |
C |
-0.520 |
|
|
|
| 4 |
C |
-0.520 |
|
|
|
| 5 |
H |
0.161 |
|
|
|
| 6 |
H |
0.161 |
|
|
|
| 7 |
H |
0.161 |
|
|
|
| 8 |
H |
0.161 |
|
|
|
| 9 |
H |
0.171 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
H |
0.168 |
|
|
|
| 12 |
H |
0.171 |
|
|
|
| 13 |
H |
0.168 |
|
|
|
| 14 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.153 |
-0.323 |
0.000 |
| y |
-0.323 |
-28.505 |
0.000 |
| z |
0.000 |
0.000 |
-27.574 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.114 |
-0.323 |
0.000 |
| y |
-0.323 |
-0.642 |
0.000 |
| z |
0.000 |
0.000 |
0.755 |
|
| Polar |
|---|
| 3z2-r2 | 1.511 |
|---|
| x2-y2 | 0.352 |
|---|
| xy | -0.323 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.199 |
0.154 |
0.000 |
| y |
0.154 |
7.311 |
0.000 |
| z |
0.000 |
0.000 |
6.039 |
<r2> (average value of r
2) Å
2
| <r2> |
118.444 |
| (<r2>)1/2 |
10.883 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -158.248999 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3155 |
3007 |
33.47 |
|
|
|
| 2 |
A |
3142 |
2995 |
2.42 |
|
|
|
| 3 |
A |
3093 |
2948 |
25.91 |
|
|
|
| 4 |
A |
3069 |
2925 |
23.75 |
|
|
|
| 5 |
A |
3054 |
2911 |
21.87 |
|
|
|
| 6 |
A |
1540 |
1468 |
1.56 |
|
|
|
| 7 |
A |
1533 |
1461 |
5.04 |
|
|
|
| 8 |
A |
1513 |
1442 |
0.00 |
|
|
|
| 9 |
A |
1439 |
1371 |
4.44 |
|
|
|
| 10 |
A |
1401 |
1335 |
0.32 |
|
|
|
| 11 |
A |
1330 |
1268 |
0.37 |
|
|
|
| 12 |
A |
1210 |
1153 |
0.03 |
|
|
|
| 13 |
A |
1117 |
1065 |
0.03 |
|
|
|
| 14 |
A |
1007 |
960 |
0.22 |
|
|
|
| 15 |
A |
852 |
813 |
0.00 |
|
|
|
| 16 |
A |
806 |
768 |
2.21 |
|
|
|
| 17 |
A |
324 |
309 |
0.05 |
|
|
|
| 18 |
A |
272 |
259 |
0.01 |
|
|
|
| 19 |
A |
112 |
106 |
0.01 |
|
|
|
| 20 |
B |
3149 |
3002 |
48.82 |
|
|
|
| 21 |
B |
3143 |
2996 |
76.43 |
|
|
|
| 22 |
B |
3098 |
2953 |
29.13 |
|
|
|
| 23 |
B |
3067 |
2923 |
23.17 |
|
|
|
| 24 |
B |
3053 |
2910 |
28.83 |
|
|
|
| 25 |
B |
1533 |
1461 |
10.20 |
|
|
|
| 26 |
B |
1525 |
1454 |
10.51 |
|
|
|
| 27 |
B |
1509 |
1438 |
0.49 |
|
|
|
| 28 |
B |
1441 |
1373 |
7.93 |
|
|
|
| 29 |
B |
1385 |
1320 |
2.26 |
|
|
|
| 30 |
B |
1304 |
1243 |
0.54 |
|
|
|
| 31 |
B |
1168 |
1114 |
3.34 |
|
|
|
| 32 |
B |
994 |
947 |
1.45 |
|
|
|
| 33 |
B |
977 |
931 |
3.59 |
|
|
|
| 34 |
B |
754 |
718 |
5.26 |
|
|
|
| 35 |
B |
434 |
414 |
0.61 |
|
|
|
| 36 |
B |
216 |
206 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29358.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 27981.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
