Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -158.275836 |
| Energy at 298.15K | -158.286472 |
| HF Energy | -158.275836 |
| Nuclear repulsion energy | 130.715677 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
128 |
122 |
0.00 |
|
|
|
|
Au |
223 |
213 |
0.00 |
|
|
|
|
Bu |
257 |
246 |
0.06 |
|
|
|
|
Bg |
263 |
251 |
0.00 |
|
|
|
|
Ag |
428 |
408 |
0.00 |
|
|
|
|
Au |
739 |
706 |
4.69 |
|
|
|
|
Bg |
814 |
777 |
0.00 |
|
|
|
|
Ag |
857 |
818 |
0.00 |
|
|
|
|
Au |
966 |
921 |
1.55 |
|
|
|
|
Bu |
984 |
939 |
8.05 |
|
|
|
|
Bu |
1045 |
998 |
0.12 |
|
|
|
|
Ag |
1093 |
1043 |
0.00 |
|
|
|
|
Ag |
1180 |
1126 |
0.00 |
|
|
|
|
Bg |
1214 |
1158 |
0.00 |
|
|
|
|
Au |
1296 |
1237 |
0.05 |
|
|
|
|
Bu |
1323 |
1263 |
3.01 |
|
|
|
|
Bg |
1340 |
1278 |
0.00 |
|
|
|
|
Ag |
1408 |
1343 |
0.00 |
|
|
|
|
Bu |
1424 |
1359 |
5.65 |
|
|
|
|
Ag |
1424 |
1359 |
0.00 |
|
|
|
|
Ag |
1497 |
1429 |
0.00 |
|
|
|
|
Bu |
1501 |
1432 |
1.65 |
|
|
|
|
Bg |
1508 |
1439 |
0.00 |
|
|
|
|
Au |
1510 |
1441 |
14.22 |
|
|
|
|
Ag |
1516 |
1447 |
0.00 |
|
|
|
|
Bu |
1521 |
1452 |
7.01 |
|
|
|
|
Ag |
3042 |
2903 |
0.00 |
|
|
|
|
Bu |
3050 |
2911 |
39.96 |
|
|
|
|
Bu |
3057 |
2918 |
80.37 |
|
|
|
|
Ag |
3058 |
2918 |
0.00 |
|
|
|
|
Bg |
3074 |
2933 |
0.00 |
|
|
|
|
Au |
3096 |
2955 |
25.52 |
|
|
|
|
Bg |
3135 |
2992 |
0.00 |
|
|
|
|
Au |
3138 |
2994 |
98.76 |
|
|
|
|
Ag |
3143 |
3000 |
0.00 |
|
|
|
|
Bu |
3144 |
3000 |
68.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29196.8 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 27862.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.420 |
0.637 |
0.000 |
| C2 |
0.420 |
-0.637 |
0.000 |
| C3 |
0.420 |
1.907 |
0.000 |
| C4 |
-0.420 |
-1.907 |
0.000 |
| H5 |
-1.082 |
0.631 |
0.877 |
| H6 |
-1.082 |
0.631 |
-0.877 |
| H7 |
1.082 |
-0.631 |
0.877 |
| H8 |
1.082 |
-0.631 |
-0.877 |
| H9 |
-0.206 |
2.805 |
0.000 |
| H10 |
1.067 |
1.953 |
0.883 |
| H11 |
1.067 |
1.953 |
-0.883 |
| H12 |
0.206 |
-2.805 |
0.000 |
| H13 |
-1.067 |
-1.953 |
0.883 |
| H14 |
-1.067 |
-1.953 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5254 | 1.5231 | 2.5441 | 1.0984 | 1.0984 | 2.1520 | 2.1520 | 2.1792 | 2.1732 | 2.1732 | 3.4984 | 2.8116 | 2.8116 |
C2 | 1.5254 | | 2.5441 | 1.5231 | 2.1520 | 2.1520 | 1.0984 | 1.0984 | 3.4984 | 2.8116 | 2.8116 | 2.1792 | 2.1732 | 2.1732 | C3 | 1.5231 | 2.5441 | | 3.9062 | 2.1569 | 2.1569 | 2.7661 | 2.7661 | 1.0943 | 1.0956 | 1.0956 | 4.7176 | 4.2301 | 4.2301 | C4 | 2.5441 | 1.5231 | 3.9062 | | 2.7661 | 2.7661 | 2.1569 | 2.1569 | 4.7176 | 4.2301 | 4.2301 | 1.0943 | 1.0956 | 1.0956 | H5 | 1.0984 | 2.1520 | 2.1569 | 2.7661 | | 1.7532 | 2.5050 | 3.0575 | 2.5026 | 2.5227 | 3.0758 | 3.7730 | 2.5843 | 3.1265 | H6 | 1.0984 | 2.1520 | 2.1569 | 2.7661 | 1.7532 | | 3.0575 | 2.5050 | 2.5026 | 3.0758 | 2.5227 | 3.7730 | 3.1265 | 2.5843 | H7 | 2.1520 | 1.0984 | 2.7661 | 2.1569 | 2.5050 | 3.0575 | | 1.7532 | 3.7730 | 2.5843 | 3.1265 | 2.5026 | 2.5227 | 3.0758 | H8 | 2.1520 | 1.0984 | 2.7661 | 2.1569 | 3.0575 | 2.5050 | 1.7532 | | 3.7730 | 3.1265 | 2.5843 | 2.5026 | 3.0758 | 2.5227 | H9 | 2.1792 | 3.4984 | 1.0943 | 4.7176 | 2.5026 | 2.5026 | 3.7730 | 3.7730 | | 1.7678 | 1.7678 | 5.6257 | 4.9157 | 4.9157 | H10 | 2.1732 | 2.8116 | 1.0956 | 4.2301 | 2.5227 | 3.0758 | 2.5843 | 3.1265 | 1.7678 | | 1.7661 | 4.9157 | 4.4509 | 4.7885 | H11 | 2.1732 | 2.8116 | 1.0956 | 4.2301 | 3.0758 | 2.5227 | 3.1265 | 2.5843 | 1.7678 | 1.7661 | | 4.9157 | 4.7885 | 4.4509 | H12 | 3.4984 | 2.1792 | 4.7176 | 1.0943 | 3.7730 | 3.7730 | 2.5026 | 2.5026 | 5.6257 | 4.9157 | 4.9157 | | 1.7678 | 1.7678 | H13 | 2.8116 | 2.1732 | 4.2301 | 1.0956 | 2.5843 | 3.1265 | 2.5227 | 3.0758 | 4.9157 | 4.4509 | 4.7885 | 1.7678 | | 1.7661 | H14 | 2.8116 | 2.1732 | 4.2301 | 1.0956 | 3.1265 | 2.5843 | 3.0758 | 2.5227 | 4.9157 | 4.7885 | 4.4509 | 1.7678 | 1.7661 | |
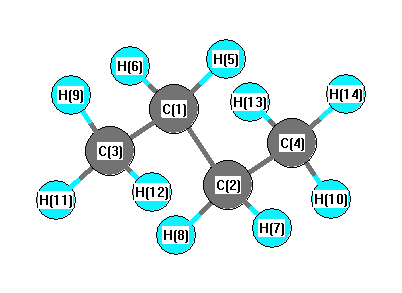 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.135 |
|
C1 |
C2 |
H7 |
109.121 |
| C1 |
C2 |
H8 |
109.121 |
|
C1 |
C3 |
H9 |
111.678 |
| C1 |
C3 |
H11 |
111.121 |
|
C1 |
C3 |
H12 |
30.857 |
| C2 |
C1 |
C3 |
113.135 |
|
C2 |
C1 |
H5 |
109.121 |
| C2 |
C1 |
H6 |
109.121 |
|
C2 |
C4 |
H10 |
17.223 |
| C2 |
C4 |
H13 |
111.121 |
|
C2 |
C4 |
H14 |
111.121 |
| C3 |
C1 |
H5 |
109.660 |
|
C3 |
C1 |
H6 |
109.660 |
| C4 |
C2 |
H7 |
109.660 |
|
C4 |
C2 |
H8 |
109.660 |
| H5 |
C1 |
H6 |
105.885 |
|
H7 |
C2 |
H8 |
105.885 |
| H9 |
C3 |
H11 |
107.655 |
|
H9 |
C3 |
H12 |
142.535 |
| H10 |
C4 |
H13 |
94.435 |
|
H10 |
C4 |
H14 |
114.446 |
| H11 |
C3 |
H12 |
93.927 |
|
H13 |
C4 |
H14 |
107.412 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.230 |
|
|
|
| 2 |
C |
-0.230 |
|
|
|
| 3 |
C |
-0.390 |
|
|
|
| 4 |
C |
-0.390 |
|
|
|
| 5 |
H |
0.120 |
|
|
|
| 6 |
H |
0.120 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.127 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
| 12 |
H |
0.127 |
|
|
|
| 13 |
H |
0.127 |
|
|
|
| 14 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.142 |
-0.283 |
0.000 |
| y |
-0.283 |
-28.424 |
0.000 |
| z |
0.000 |
0.000 |
-27.620 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.119 |
-0.283 |
0.000 |
| y |
-0.283 |
-0.543 |
0.000 |
| z |
0.000 |
0.000 |
0.663 |
|
| Polar |
|---|
| 3z2-r2 | 1.325 |
|---|
| x2-y2 | 0.283 |
|---|
| xy | -0.283 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.309 |
0.163 |
0.000 |
| y |
0.163 |
7.469 |
0.000 |
| z |
0.000 |
0.000 |
6.135 |
<r2> (average value of r
2) Å
2
| <r2> |
118.385 |
| (<r2>)1/2 |
10.880 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -158.274673 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3150 |
3006 |
31.62 |
|
|
|
| 2 |
A |
3139 |
2996 |
3.80 |
|
|
|
| 3 |
A |
3090 |
2949 |
25.67 |
|
|
|
| 4 |
A |
3062 |
2922 |
24.50 |
|
|
|
| 5 |
A |
3048 |
2909 |
21.95 |
|
|
|
| 6 |
A |
1522 |
1452 |
1.29 |
|
|
|
| 7 |
A |
1515 |
1446 |
4.75 |
|
|
|
| 8 |
A |
1496 |
1427 |
0.01 |
|
|
|
| 9 |
A |
1423 |
1358 |
4.05 |
|
|
|
| 10 |
A |
1390 |
1326 |
0.30 |
|
|
|
| 11 |
A |
1320 |
1260 |
0.33 |
|
|
|
| 12 |
A |
1201 |
1146 |
0.03 |
|
|
|
| 13 |
A |
1111 |
1060 |
0.04 |
|
|
|
| 14 |
A |
999 |
954 |
0.20 |
|
|
|
| 15 |
A |
848 |
810 |
0.00 |
|
|
|
| 16 |
A |
801 |
764 |
2.10 |
|
|
|
| 17 |
A |
325 |
310 |
0.04 |
|
|
|
| 18 |
A |
269 |
256 |
0.01 |
|
|
|
| 19 |
A |
112 |
107 |
0.01 |
|
|
|
| 20 |
B |
3146 |
3002 |
49.19 |
|
|
|
| 21 |
B |
3140 |
2997 |
74.89 |
|
|
|
| 22 |
B |
3095 |
2954 |
27.82 |
|
|
|
| 23 |
B |
3060 |
2921 |
23.59 |
|
|
|
| 24 |
B |
3048 |
2909 |
29.48 |
|
|
|
| 25 |
B |
1515 |
1446 |
8.88 |
|
|
|
| 26 |
B |
1507 |
1438 |
9.89 |
|
|
|
| 27 |
B |
1491 |
1423 |
0.49 |
|
|
|
| 28 |
B |
1426 |
1361 |
7.79 |
|
|
|
| 29 |
B |
1372 |
1309 |
2.11 |
|
|
|
| 30 |
B |
1295 |
1235 |
0.57 |
|
|
|
| 31 |
B |
1161 |
1108 |
3.31 |
|
|
|
| 32 |
B |
988 |
943 |
1.51 |
|
|
|
| 33 |
B |
970 |
926 |
3.36 |
|
|
|
| 34 |
B |
750 |
716 |
5.09 |
|
|
|
| 35 |
B |
434 |
414 |
0.60 |
|
|
|
| 36 |
B |
216 |
206 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29217.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 27882.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
