return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-156.706348
Energy at 298.15K-156.714374
HF Energy-156.118632
Nuclear repulsion energy117.492737
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3340 3121 16.06      
2 A 3242 3029 3.88      
3 A 3232 3020 33.42      
4 A 3227 3015 42.70      
5 A 3224 3013 4.87      
6 A 3177 2968 11.20      
7 A 3130 2925 25.31      
8 A 3119 2915 24.83      
9 A 1743 1628 3.81      
10 A 1568 1465 3.44      
11 A 1561 1459 4.84      
12 A 1542 1441 2.58      
13 A 1504 1405 1.11      
14 A 1464 1368 1.13      
15 A 1389 1298 2.69      
16 A 1348 1260 1.45      
17 A 1332 1244 0.10      
18 A 1237 1155 0.20      
19 A 1130 1056 3.09      
20 A 1073 1002 1.98      
21 A 1046 978 13.41      
22 A 1013 946 2.13      
23 A 934 873 33.75      
24 A 892 833 1.33      
25 A 813 760 2.29      
26 A 663 619 7.79      
27 A 439 411 0.62      
28 A 325 303 0.42      
29 A 248 231 0.03      
30 A 108 101 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 24530.0 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 22920.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.74659 0.13996 0.13706

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.706 -0.252 -0.300
C2 0.541 0.524 0.310
C3 -0.714 -0.291 0.358
C4 -1.843 0.016 -0.288
H5 -0.674 -1.201 0.948
H6 1.476 -0.544 -1.323
H7 2.615 0.346 -0.312
H8 1.908 -1.158 0.269
H9 0.811 0.839 1.321
H10 0.359 1.434 -0.264
H11 -1.919 0.912 -0.888
H12 -2.718 -0.614 -0.232

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.52762.50833.55942.85071.08801.08831.08882.14942.15833.85314.4396
C21.52761.49712.50972.20452.16362.17302.16861.09241.09092.76393.4942
C32.50831.49711.33631.08542.77173.45522.76342.12782.12422.11012.1139
C43.55942.50971.33632.09153.52124.47063.97003.21022.61881.08131.0796
H52.85072.20451.08542.09153.19563.84752.67102.54993.07873.06382.4320
H61.08802.16362.77173.52123.19561.76421.75963.05662.50563.71964.3338
H71.08832.17303.45524.47063.84751.76421.76082.48312.50514.60605.4196
H81.08882.16862.76343.97002.67101.75961.76082.51003.06644.50284.6851
H92.14941.09242.12783.21022.54993.05662.48312.51001.75263.51274.1197
H102.15831.09092.12422.61883.07872.50562.50513.06641.75262.41913.6960
H113.85312.76392.11011.08133.06383.71964.60604.50283.51272.41911.8430
H124.43963.49422.11391.07962.43204.33385.41964.68514.11973.69601.8430

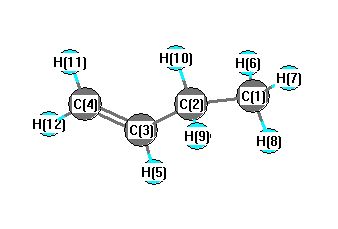
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.047 C1 C2 H9 109.121
C1 C2 H10 109.906 C2 C1 H6 110.498
C2 C1 H7 111.228 C2 C1 H8 110.849
C2 C3 C4 124.588 C2 C3 H5 116.307
C3 C2 H9 109.517 C3 C2 H10 109.322
C3 C4 H11 121.222 C3 C4 H12 121.725
C4 C3 H5 119.094 H6 C1 H7 108.319
H6 C1 H8 107.872 H7 C1 H8 107.955
H9 C2 H10 106.785 H11 C4 H12 117.053
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability