return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: MP3=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3=FULL/6-31G**
 hartrees
Energy at 0K-156.749570
Energy at 298.15K-156.757602
HF Energy-156.118864
Nuclear repulsion energy117.401069
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3339 3339 19.79      
2 A 3249 3249 4.33      
3 A 3235 3235 32.25      
4 A 3228 3228 33.41      
5 A 3222 3222 30.32      
6 A 3177 3177 11.80      
7 A 3137 3137 25.61      
8 A 3127 3127 28.30      
9 A 1779 1779 4.06      
10 A 1574 1574 3.44      
11 A 1567 1567 4.42      
12 A 1552 1552 2.14      
13 A 1515 1515 1.19      
14 A 1476 1476 1.02      
15 A 1403 1403 3.12      
16 A 1360 1360 1.75      
17 A 1339 1339 0.10      
18 A 1243 1243 0.23      
19 A 1135 1135 3.25      
20 A 1075 1075 1.89      
21 A 1049 1049 12.26      
22 A 1019 1019 1.46      
23 A 947 947 33.73      
24 A 892 892 1.19      
25 A 815 815 1.90      
26 A 663 663 7.54      
27 A 440 440 0.67      
28 A 325 325 0.40      
29 A 244 244 0.03      
30 A 107 107 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 24616.5 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 24616.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3=FULL/6-31G**
ABC
0.75333 0.13922 0.13619

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.715 -0.246 -0.298
C2 0.540 0.520 0.313
C3 -0.718 -0.300 0.347
C4 -1.848 0.021 -0.283
H5 -0.673 -1.220 0.918
H6 1.494 -0.529 -1.325
H7 2.620 0.358 -0.297
H8 1.918 -1.157 0.263
H9 0.801 0.824 1.328
H10 0.359 1.434 -0.253
H11 -1.931 0.927 -0.865
H12 -2.723 -0.608 -0.235

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.52992.51793.57292.85151.08761.08761.08852.15072.15933.87174.4537
C21.52991.50192.51122.20562.16632.17312.17061.09171.08992.76743.4960
C32.51791.50191.33261.08422.78253.46232.77342.12952.12712.10842.1107
C43.57292.51121.33262.08893.54364.48013.98333.20312.62071.08061.0789
H52.85152.20561.08422.08893.19483.84802.67352.55333.07873.06182.4313
H61.08762.16632.78253.54363.19481.76351.75903.05782.50813.75004.3569
H71.08762.17313.46234.48013.84801.76351.76032.48282.50384.62085.4299
H81.08852.17062.77343.98332.67351.75901.76032.51113.06684.51984.7004
H92.15071.09172.12953.20312.55333.05782.48282.51111.75103.50514.1133
H102.15931.08992.12712.62073.07872.50812.50383.06681.75102.42383.6973
H113.87172.76742.10841.08063.06183.75004.62084.51983.50512.42381.8389
H124.45373.49602.11071.07892.43134.35695.42994.70044.11333.69731.8389

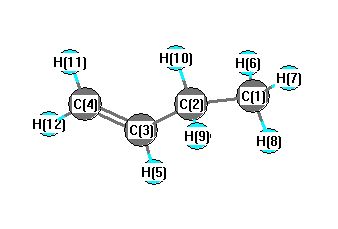
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.295 C1 C2 H9 109.100
C1 C2 H10 109.879 C2 C1 H6 110.570
C2 C1 H7 111.115 C2 C1 H8 110.866
C2 C3 C4 124.627 C2 C3 H5 116.107
C3 C2 H9 109.362 C3 C2 H10 109.283
C3 C4 H11 121.429 C3 C4 H12 121.806
C4 C3 H5 119.258 H6 C1 H7 108.328
H6 C1 H8 107.863 H7 C1 H8 107.981
H9 C2 H10 106.767 H11 C4 H12 116.765
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability