Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -155.783401 |
| Energy at 298.15K | -155.788745 |
| HF Energy | -155.783401 |
| Nuclear repulsion energy | 104.883692 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3278 |
3124 |
6.02 |
|
|
|
| 2 |
A |
3195 |
3045 |
0.69 |
|
|
|
| 3 |
A |
3182 |
3033 |
28.36 |
|
|
|
| 4 |
A |
1722 |
1641 |
1.49 |
|
|
|
| 5 |
A |
1484 |
1415 |
8.99 |
|
|
|
| 6 |
A |
1356 |
1293 |
0.07 |
|
|
|
| 7 |
A |
1080 |
1029 |
0.02 |
|
|
|
| 8 |
A |
1027 |
978 |
1.54 |
|
|
|
| 9 |
A |
943 |
898 |
5.03 |
|
|
|
| 10 |
A |
903 |
861 |
0.75 |
|
|
|
| 11 |
A |
763 |
727 |
3.71 |
|
|
|
| 12 |
A |
273 |
260 |
0.00 |
|
|
|
| 13 |
A |
168 |
160 |
0.12 |
|
|
|
| 14 |
B |
3276 |
3123 |
17.86 |
|
|
|
| 15 |
B |
3190 |
3040 |
6.30 |
|
|
|
| 16 |
B |
3172 |
3023 |
9.86 |
|
|
|
| 17 |
B |
1736 |
1654 |
2.33 |
|
|
|
| 18 |
B |
1452 |
1384 |
1.89 |
|
|
|
| 19 |
B |
1323 |
1261 |
0.24 |
|
|
|
| 20 |
B |
1112 |
1060 |
6.75 |
|
|
|
| 21 |
B |
1041 |
992 |
28.99 |
|
|
|
| 22 |
B |
943 |
899 |
80.66 |
|
|
|
| 23 |
B |
618 |
589 |
6.16 |
|
|
|
| 24 |
B |
474 |
451 |
11.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18853.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 17969.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.099 |
0.726 |
0.554 |
| C2 |
-0.099 |
-0.726 |
0.554 |
| C3 |
-0.099 |
1.533 |
-0.493 |
| C4 |
0.099 |
-1.533 |
-0.493 |
| H5 |
0.414 |
1.165 |
1.501 |
| H6 |
-0.414 |
-1.165 |
1.501 |
| H7 |
0.087 |
2.601 |
-0.429 |
| H8 |
-0.467 |
1.153 |
-1.443 |
| H9 |
-0.087 |
-2.601 |
-0.429 |
| H10 |
0.467 |
-1.153 |
-1.443 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4655 | 1.3364 | 2.4898 | 1.0904 | 2.1764 | 2.1171 | 2.1190 | 3.4743 | 2.7666 |
C2 | 1.4655 | | 2.4898 | 1.3364 | 2.1764 | 1.0904 | 3.4743 | 2.7666 | 2.1171 | 2.1190 | C3 | 1.3364 | 2.4898 | | 3.0722 | 2.0914 | 3.3697 | 1.0861 | 1.0874 | 4.1344 | 2.9046 | C4 | 2.4898 | 1.3364 | 3.0722 | | 3.3697 | 2.0914 | 4.1344 | 2.9046 | 1.0861 | 1.0874 | H5 | 1.0904 | 2.1764 | 2.0914 | 3.3697 | | 2.4731 | 2.4277 | 3.0729 | 4.2616 | 3.7475 | H6 | 2.1764 | 1.0904 | 3.3697 | 2.0914 | 2.4731 | | 4.2616 | 3.7475 | 2.4277 | 3.0729 | H7 | 2.1171 | 3.4743 | 1.0861 | 4.1344 | 2.4277 | 4.2616 | | 1.8526 | 5.2049 | 3.9069 | H8 | 2.1190 | 2.7666 | 1.0874 | 2.9046 | 3.0729 | 3.7475 | 1.8526 | | 3.9069 | 2.4878 | H9 | 3.4743 | 2.1171 | 4.1344 | 1.0861 | 4.2616 | 2.4277 | 5.2049 | 3.9069 | | 1.8526 | H10 | 2.7666 | 2.1190 | 2.9046 | 1.0874 | 3.7475 | 3.0729 | 3.9069 | 2.4878 | 1.8526 | |
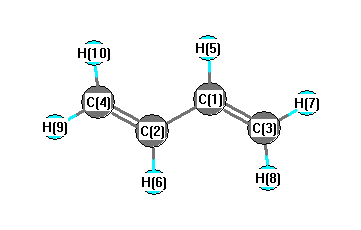 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
125.328 |
|
C1 |
C2 |
H6 |
115.978 |
| C1 |
C3 |
H7 |
121.490 |
|
C1 |
C3 |
H8 |
121.566 |
| C2 |
C1 |
C3 |
125.328 |
|
C2 |
C1 |
H5 |
115.978 |
| C2 |
C4 |
H9 |
121.490 |
|
C2 |
C4 |
H10 |
121.566 |
| C3 |
C1 |
H5 |
118.682 |
|
C4 |
C2 |
H6 |
118.682 |
| H7 |
C3 |
H8 |
116.938 |
|
H9 |
C4 |
H10 |
116.938 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.128 |
|
|
|
| 2 |
C |
-0.128 |
|
|
|
| 3 |
C |
-0.383 |
|
|
|
| 4 |
C |
-0.383 |
|
|
|
| 5 |
H |
0.167 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
| 7 |
H |
0.172 |
|
|
|
| 8 |
H |
0.172 |
|
|
|
| 9 |
H |
0.172 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.114 |
0.114 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.517 |
0.712 |
0.000 |
| y |
0.712 |
-22.611 |
0.000 |
| z |
0.000 |
0.000 |
-22.549 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.937 |
0.712 |
0.000 |
| y |
0.712 |
2.422 |
0.000 |
| z |
0.000 |
0.000 |
2.515 |
|
| Polar |
|---|
| 3z2-r2 | 5.030 |
|---|
| x2-y2 | -4.906 |
|---|
| xy | 0.712 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.737 |
0.035 |
0.000 |
| y |
0.035 |
8.851 |
0.000 |
| z |
0.000 |
0.000 |
6.573 |
<r2> (average value of r
2) Å
2
| <r2> |
85.225 |
| (<r2>)1/2 |
9.232 |
