Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -155.805959 |
| Energy at 298.15K | -155.811301 |
| HF Energy | -155.805959 |
| Nuclear repulsion energy | 104.948309 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3273 |
3124 |
5.68 |
|
|
|
| 2 |
A |
3186 |
3041 |
3.09 |
|
|
|
| 3 |
A |
3174 |
3029 |
24.74 |
|
|
|
| 4 |
A |
1716 |
1637 |
1.54 |
|
|
|
| 5 |
A |
1473 |
1405 |
9.32 |
|
|
|
| 6 |
A |
1347 |
1286 |
0.09 |
|
|
|
| 7 |
A |
1074 |
1025 |
0.01 |
|
|
|
| 8 |
A |
1024 |
977 |
1.45 |
|
|
|
| 9 |
A |
944 |
900 |
4.70 |
|
|
|
| 10 |
A |
901 |
859 |
0.63 |
|
|
|
| 11 |
A |
763 |
728 |
3.58 |
|
|
|
| 12 |
A |
273 |
261 |
0.00 |
|
|
|
| 13 |
A |
169 |
162 |
0.14 |
|
|
|
| 14 |
B |
3272 |
3122 |
17.26 |
|
|
|
| 15 |
B |
3180 |
3034 |
6.08 |
|
|
|
| 16 |
B |
3166 |
3022 |
9.54 |
|
|
|
| 17 |
B |
1730 |
1650 |
2.36 |
|
|
|
| 18 |
B |
1439 |
1373 |
2.00 |
|
|
|
| 19 |
B |
1313 |
1253 |
0.26 |
|
|
|
| 20 |
B |
1106 |
1055 |
6.46 |
|
|
|
| 21 |
B |
1039 |
991 |
27.09 |
|
|
|
| 22 |
B |
944 |
901 |
76.65 |
|
|
|
| 23 |
B |
618 |
590 |
6.04 |
|
|
|
| 24 |
B |
474 |
452 |
11.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18798.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 17939.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.099 |
0.726 |
0.554 |
| C2 |
-0.099 |
-0.726 |
0.554 |
| C3 |
-0.099 |
1.531 |
-0.493 |
| C4 |
0.099 |
-1.531 |
-0.493 |
| H5 |
0.413 |
1.165 |
1.501 |
| H6 |
-0.413 |
-1.165 |
1.501 |
| H7 |
0.088 |
2.599 |
-0.430 |
| H8 |
-0.467 |
1.149 |
-1.441 |
| H9 |
-0.088 |
-2.599 |
-0.430 |
| H10 |
0.467 |
-1.149 |
-1.441 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4648 | 1.3360 | 2.4882 | 1.0898 | 2.1758 | 2.1157 | 2.1168 | 3.4719 | 2.7627 |
C2 | 1.4648 | | 2.4882 | 1.3360 | 2.1758 | 1.0898 | 3.4719 | 2.7627 | 2.1157 | 2.1168 | C3 | 1.3360 | 2.4882 | | 3.0691 | 2.0905 | 3.3684 | 1.0852 | 1.0867 | 4.1304 | 2.8990 | C4 | 2.4882 | 1.3360 | 3.0691 | | 3.3684 | 2.0905 | 4.1304 | 2.8990 | 1.0852 | 1.0867 | H5 | 1.0898 | 2.1758 | 2.0905 | 3.3684 | | 2.4731 | 2.4261 | 3.0706 | 4.2597 | 3.7435 | H6 | 2.1758 | 1.0898 | 3.3684 | 2.0905 | 2.4731 | | 4.2597 | 3.7435 | 2.4261 | 3.0706 | H7 | 2.1157 | 3.4719 | 1.0852 | 4.1304 | 2.4261 | 4.2597 | | 1.8525 | 5.2000 | 3.9001 | H8 | 2.1168 | 2.7627 | 1.0867 | 2.8990 | 3.0706 | 3.7435 | 1.8525 | | 3.9001 | 2.4806 | H9 | 3.4719 | 2.1157 | 4.1304 | 1.0852 | 4.2597 | 2.4261 | 5.2000 | 3.9001 | | 1.8525 | H10 | 2.7627 | 2.1168 | 2.8990 | 1.0867 | 3.7435 | 3.0706 | 3.9001 | 2.4806 | 1.8525 | |
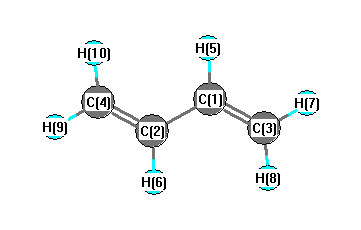 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
125.284 |
|
C1 |
C2 |
H6 |
116.022 |
| C1 |
C3 |
H7 |
121.466 |
|
C1 |
C3 |
H8 |
121.462 |
| C2 |
C1 |
C3 |
125.284 |
|
C2 |
C1 |
H5 |
116.022 |
| C2 |
C4 |
H9 |
121.466 |
|
C2 |
C4 |
H10 |
121.462 |
| C3 |
C1 |
H5 |
118.682 |
|
C4 |
C2 |
H6 |
118.682 |
| H7 |
C3 |
H8 |
117.066 |
|
H9 |
C4 |
H10 |
117.066 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.092 |
|
|
|
| 2 |
C |
-0.092 |
|
|
|
| 3 |
C |
-0.285 |
|
|
|
| 4 |
C |
-0.285 |
|
|
|
| 5 |
H |
0.122 |
|
|
|
| 6 |
H |
0.122 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.128 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.113 |
0.113 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.431 |
0.696 |
0.000 |
| y |
0.696 |
-22.668 |
0.000 |
| z |
0.000 |
0.000 |
-22.618 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.788 |
0.696 |
0.000 |
| y |
0.696 |
2.356 |
0.000 |
| z |
0.000 |
0.000 |
2.432 |
|
| Polar |
|---|
| 3z2-r2 | 4.863 |
|---|
| x2-y2 | -4.763 |
|---|
| xy | 0.696 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.810 |
0.037 |
0.000 |
| y |
0.037 |
8.936 |
0.000 |
| z |
0.000 |
0.000 |
6.654 |
<r2> (average value of r
2) Å
2
| <r2> |
85.120 |
| (<r2>)1/2 |
9.226 |
