Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -191.706764 |
| Energy at 298.15K | |
| HF Energy | -191.706764 |
| Nuclear repulsion energy | 103.131818 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3273 |
3123 |
4.88 |
64.99 |
0.54 |
0.70 |
| 2 |
A' |
3220 |
3073 |
1.54 |
95.29 |
0.31 |
0.48 |
| 3 |
A' |
3173 |
3028 |
4.65 |
69.57 |
0.10 |
0.19 |
| 4 |
A' |
2913 |
2780 |
99.77 |
86.64 |
0.28 |
0.44 |
| 5 |
A' |
1838 |
1754 |
219.22 |
59.17 |
0.39 |
0.56 |
| 6 |
A' |
1714 |
1636 |
2.77 |
28.21 |
0.20 |
0.34 |
| 7 |
A' |
1464 |
1397 |
12.11 |
6.40 |
0.54 |
0.70 |
| 8 |
A' |
1401 |
1337 |
6.93 |
21.43 |
0.45 |
0.62 |
| 9 |
A' |
1301 |
1241 |
4.09 |
6.74 |
0.34 |
0.51 |
| 10 |
A' |
1181 |
1127 |
35.51 |
11.98 |
0.63 |
0.77 |
| 11 |
A' |
930 |
887 |
19.68 |
1.26 |
0.12 |
0.22 |
| 12 |
A' |
576 |
550 |
6.33 |
5.24 |
0.35 |
0.51 |
| 13 |
A' |
318 |
303 |
9.48 |
0.90 |
0.36 |
0.53 |
| 14 |
A" |
1046 |
998 |
7.69 |
2.70 |
0.75 |
0.86 |
| 15 |
A" |
1029 |
982 |
8.29 |
3.88 |
0.75 |
0.86 |
| 16 |
A" |
999 |
953 |
41.71 |
0.42 |
0.75 |
0.86 |
| 17 |
A" |
613 |
585 |
8.90 |
4.44 |
0.75 |
0.86 |
| 18 |
A" |
174 |
166 |
3.97 |
1.61 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13580.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12959.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.743 |
0.000 |
| C2 |
0.000 |
0.720 |
0.000 |
| C3 |
1.212 |
1.279 |
0.000 |
| O4 |
-1.216 |
-1.319 |
0.000 |
| H5 |
0.811 |
-1.305 |
0.000 |
| H6 |
-0.915 |
1.307 |
0.000 |
| H7 |
1.356 |
2.354 |
0.000 |
| H8 |
2.109 |
0.663 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4703 | 2.4384 | 1.2111 | 1.1140 | 2.1875 | 3.4444 | 2.6615 |
C2 | 1.4703 | | 1.3350 | 2.3740 | 2.1807 | 1.0869 | 2.1239 | 2.1100 | C3 | 2.4384 | 1.3350 | | 3.5562 | 2.6144 | 2.1270 | 1.0851 | 1.0882 | O4 | 1.2111 | 2.3740 | 3.5562 | | 2.0273 | 2.6434 | 4.4845 | 3.8711 | H5 | 1.1140 | 2.1807 | 2.6144 | 2.0273 | | 3.1302 | 3.6993 | 2.3570 | H6 | 2.1875 | 1.0869 | 2.1270 | 2.6434 | 3.1302 | | 2.5005 | 3.0917 | H7 | 3.4444 | 2.1239 | 1.0851 | 4.4845 | 3.6993 | 2.5005 | | 1.8518 | H8 | 2.6615 | 2.1100 | 1.0882 | 3.8711 | 2.3570 | 3.0917 | 1.8518 | |
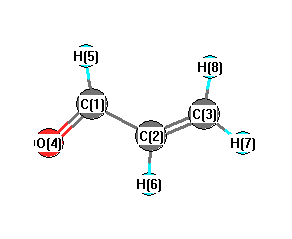 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.656 |
|
C1 |
C2 |
H6 |
116.815 |
| C2 |
C1 |
O4 |
124.303 |
|
C2 |
C1 |
H5 |
114.388 |
| C2 |
C3 |
H7 |
122.380 |
|
C2 |
C3 |
H8 |
120.750 |
| C3 |
C2 |
H6 |
122.529 |
|
O4 |
C1 |
H5 |
121.308 |
| H7 |
C3 |
H8 |
116.870 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.266 |
|
|
|
| 2 |
C |
-0.115 |
|
|
|
| 3 |
C |
-0.273 |
|
|
|
| 4 |
O |
-0.413 |
|
|
|
| 5 |
H |
0.087 |
|
|
|
| 6 |
H |
0.150 |
|
|
|
| 7 |
H |
0.153 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.403 |
2.072 |
0.000 |
3.173 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.888 |
-2.506 |
0.000 |
| y |
-2.506 |
-23.490 |
0.000 |
| z |
0.000 |
0.000 |
-24.370 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.043 |
-2.506 |
0.000 |
| y |
-2.506 |
0.639 |
0.000 |
| z |
0.000 |
0.000 |
-0.681 |
|
| Polar |
|---|
| 3z2-r2 | -1.363 |
|---|
| x2-y2 | -0.397 |
|---|
| xy | -2.506 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.709 |
1.676 |
0.000 |
| y |
1.676 |
6.147 |
0.000 |
| z |
0.000 |
0.000 |
2.131 |
<r2> (average value of r
2) Å
2
| <r2> |
83.050 |
| (<r2>)1/2 |
9.113 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -191.704081 |
| Energy at 298.15K | |
| HF Energy | -191.704081 |
| Nuclear repulsion energy | 104.780924 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3285 |
3134 |
1.69 |
53.64 |
0.64 |
0.78 |
| 2 |
A' |
3206 |
3060 |
9.83 |
117.91 |
0.23 |
0.38 |
| 3 |
A' |
3182 |
3036 |
4.44 |
51.04 |
0.14 |
0.24 |
| 4 |
A' |
2946 |
2811 |
143.74 |
145.62 |
0.32 |
0.48 |
| 5 |
A' |
1835 |
1751 |
105.60 |
13.19 |
0.49 |
0.66 |
| 6 |
A' |
1704 |
1626 |
66.66 |
44.55 |
0.22 |
0.37 |
| 7 |
A' |
1446 |
1380 |
26.46 |
9.88 |
0.71 |
0.83 |
| 8 |
A' |
1431 |
1365 |
18.81 |
7.80 |
0.25 |
0.40 |
| 9 |
A' |
1318 |
1258 |
2.30 |
15.97 |
0.44 |
0.61 |
| 10 |
A' |
1070 |
1021 |
5.01 |
4.89 |
0.73 |
0.84 |
| 11 |
A' |
954 |
911 |
60.87 |
4.77 |
0.21 |
0.34 |
| 12 |
A' |
687 |
655 |
11.19 |
0.67 |
0.74 |
0.85 |
| 13 |
A' |
288 |
275 |
7.38 |
3.07 |
0.47 |
0.64 |
| 14 |
A" |
1046 |
998 |
5.46 |
9.52 |
0.75 |
0.86 |
| 15 |
A" |
1034 |
987 |
28.98 |
0.46 |
0.75 |
0.86 |
| 16 |
A" |
1010 |
964 |
15.10 |
0.16 |
0.75 |
0.86 |
| 17 |
A" |
566 |
540 |
7.87 |
6.51 |
0.75 |
0.86 |
| 18 |
A" |
183 |
175 |
7.49 |
0.88 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13594.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12973.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.888 |
-0.284 |
0.000 |
| C2 |
0.000 |
0.900 |
0.000 |
| C3 |
1.327 |
0.760 |
0.000 |
| O4 |
-0.494 |
-1.431 |
0.000 |
| H5 |
-1.976 |
-0.062 |
0.000 |
| H6 |
-0.472 |
1.880 |
0.000 |
| H7 |
1.998 |
1.613 |
0.000 |
| H8 |
1.761 |
-0.236 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4799 | 2.4492 | 1.2125 | 1.1107 | 2.2036 | 3.4540 | 2.6495 |
C2 | 1.4799 | | 1.3348 | 2.3823 | 2.1977 | 1.0877 | 2.1218 | 2.0956 | C3 | 2.4492 | 1.3348 | | 2.8491 | 3.4044 | 2.1192 | 1.0850 | 1.0867 | O4 | 1.2125 | 2.3823 | 2.8491 | | 2.0179 | 3.3107 | 3.9339 | 2.5518 | H5 | 1.1107 | 2.1977 | 3.4044 | 2.0179 | | 2.4562 | 4.3130 | 3.7414 | H6 | 2.2036 | 1.0877 | 2.1192 | 3.3107 | 2.4562 | | 2.4846 | 3.0762 | H7 | 3.4540 | 2.1218 | 1.0850 | 3.9339 | 4.3130 | 2.4846 | | 1.8643 | H8 | 2.6495 | 2.0956 | 1.0867 | 2.5518 | 3.7414 | 3.0762 | 1.8643 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.870 |
|
C1 |
C2 |
H6 |
117.420 |
| C2 |
C1 |
O4 |
124.159 |
|
C2 |
C1 |
H5 |
115.321 |
| C2 |
C3 |
H7 |
122.194 |
|
C2 |
C3 |
H8 |
119.521 |
| C3 |
C2 |
H6 |
121.710 |
|
O4 |
C1 |
H5 |
120.520 |
| H7 |
C3 |
H8 |
118.285 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.259 |
|
|
|
| 2 |
C |
-0.150 |
|
|
|
| 3 |
C |
-0.240 |
|
|
|
| 4 |
O |
-0.407 |
|
|
|
| 5 |
H |
0.093 |
|
|
|
| 6 |
H |
0.132 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
| 8 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.274 |
2.624 |
0.000 |
2.638 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.847 |
-0.641 |
0.000 |
| y |
-0.641 |
-25.551 |
0.000 |
| z |
0.000 |
0.000 |
-24.345 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.101 |
-0.641 |
0.000 |
| y |
-0.641 |
-3.455 |
0.000 |
| z |
0.000 |
0.000 |
-1.646 |
|
| Polar |
|---|
| 3z2-r2 | -3.292 |
|---|
| x2-y2 | 5.704 |
|---|
| xy | -0.641 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.883 |
0.501 |
0.000 |
| y |
0.501 |
5.524 |
0.000 |
| z |
0.000 |
0.000 |
2.127 |
<r2> (average value of r
2) Å
2
| <r2> |
74.379 |
| (<r2>)1/2 |
8.624 |
