Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -3109.938223 |
| Energy at 298.15K | |
| HF Energy | -3109.938223 |
| Nuclear repulsion energy | 289.235288 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3144 |
3000 |
6.83 |
34.85 |
0.07 |
0.12 |
| 2 |
A' |
3130 |
2987 |
2.91 |
120.41 |
0.07 |
0.13 |
| 3 |
A' |
1500 |
1431 |
1.26 |
14.90 |
0.74 |
0.85 |
| 4 |
A' |
1493 |
1425 |
6.90 |
2.14 |
0.73 |
0.85 |
| 5 |
A' |
1331 |
1270 |
2.56 |
13.98 |
0.59 |
0.74 |
| 6 |
A' |
1235 |
1179 |
39.63 |
2.29 |
0.43 |
0.60 |
| 7 |
A' |
1090 |
1040 |
1.45 |
7.69 |
0.75 |
0.85 |
| 8 |
A' |
761 |
726 |
33.46 |
47.28 |
0.30 |
0.46 |
| 9 |
A' |
659 |
629 |
55.67 |
12.15 |
0.25 |
0.40 |
| 10 |
A' |
253 |
241 |
0.97 |
4.45 |
0.34 |
0.50 |
| 11 |
A' |
194 |
185 |
7.89 |
0.14 |
0.28 |
0.44 |
| 12 |
A" |
3224 |
3077 |
0.99 |
12.43 |
0.75 |
0.86 |
| 13 |
A" |
3196 |
3050 |
0.60 |
85.93 |
0.75 |
0.86 |
| 14 |
A" |
1300 |
1241 |
0.08 |
9.07 |
0.75 |
0.86 |
| 15 |
A" |
1137 |
1085 |
2.42 |
0.32 |
0.75 |
0.86 |
| 16 |
A" |
980 |
935 |
0.09 |
7.94 |
0.75 |
0.86 |
| 17 |
A" |
769 |
734 |
4.33 |
0.03 |
0.75 |
0.86 |
| 18 |
A" |
114 |
108 |
5.87 |
0.03 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12753.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12171.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.653 |
0.000 |
| C2 |
1.222 |
-0.231 |
0.000 |
| Br3 |
-1.589 |
-0.479 |
0.000 |
| Cl4 |
2.699 |
0.789 |
0.000 |
| H5 |
-0.056 |
1.274 |
0.893 |
| H6 |
-0.056 |
1.274 |
-0.893 |
| H7 |
1.256 |
-0.862 |
0.889 |
| H8 |
1.256 |
-0.862 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5087 | 1.9511 | 2.7020 | 1.0892 | 1.0892 | 2.1595 | 2.1595 |
C2 | 1.5087 | | 2.8216 | 1.7948 | 2.1675 | 2.1675 | 1.0905 | 1.0905 | Br3 | 1.9511 | 2.8216 | | 4.4709 | 2.4945 | 2.4945 | 3.0050 | 3.0050 | Cl4 | 2.7020 | 1.7948 | 4.4709 | | 2.9360 | 2.9360 | 2.3656 | 2.3656 | H5 | 1.0892 | 2.1675 | 2.4945 | 2.9360 | | 1.7861 | 2.5066 | 3.0758 | H6 | 1.0892 | 2.1675 | 2.4945 | 2.9360 | 1.7861 | | 3.0758 | 2.5066 | H7 | 2.1595 | 1.0905 | 3.0050 | 2.3656 | 2.5066 | 3.0758 | | 1.7787 | H8 | 2.1595 | 1.0905 | 3.0050 | 2.3656 | 3.0758 | 2.5066 | 1.7787 | |
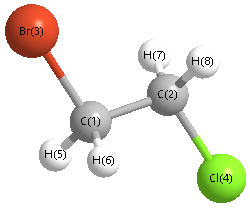 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.448 |
|
C1 |
C2 |
H7 |
111.355 |
| C1 |
C2 |
H8 |
111.355 |
|
C2 |
C1 |
Br3 |
108.611 |
| C2 |
C1 |
Cl4 |
38.783 |
|
C2 |
C1 |
H6 |
112.082 |
| Br3 |
C1 |
H5 |
106.813 |
|
Br3 |
C1 |
H6 |
106.813 |
| Cl4 |
C2 |
H7 |
107.626 |
|
Cl4 |
C2 |
H8 |
107.626 |
| H5 |
C1 |
H6 |
110.148 |
|
H7 |
C2 |
H8 |
109.288 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.263 |
|
|
|
| 2 |
C |
-0.355 |
|
|
|
| 3 |
Br |
-0.124 |
|
|
|
| 4 |
Cl |
-0.059 |
|
|
|
| 5 |
H |
0.198 |
|
|
|
| 6 |
H |
0.198 |
|
|
|
| 7 |
H |
0.203 |
|
|
|
| 8 |
H |
0.203 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.202 |
-0.053 |
0.000 |
0.209 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.982 |
-2.215 |
0.000 |
| y |
-2.215 |
-42.825 |
0.000 |
| z |
0.000 |
0.000 |
-43.122 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.009 |
-2.215 |
0.000 |
| y |
-2.215 |
3.728 |
0.000 |
| z |
0.000 |
0.000 |
3.281 |
|
| Polar |
|---|
| 3z2-r2 | 6.563 |
|---|
| x2-y2 | -7.158 |
|---|
| xy | -2.215 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.000 |
1.894 |
0.000 |
| y |
1.894 |
6.297 |
0.000 |
| z |
0.000 |
0.000 |
5.164 |
<r2> (average value of r
2) Å
2
| <r2> |
281.974 |
| (<r2>)1/2 |
16.792 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -3109.935618 |
| Energy at 298.15K | |
| HF Energy | -3109.935618 |
| Nuclear repulsion energy | 306.048356 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3202 |
3056 |
1.11 |
46.11 |
0.75 |
0.86 |
| 2 |
A |
3177 |
3032 |
1.43 |
84.12 |
0.75 |
0.86 |
| 3 |
A |
3124 |
2982 |
9.59 |
140.71 |
0.05 |
0.10 |
| 4 |
A |
3104 |
2962 |
9.65 |
113.58 |
0.19 |
0.32 |
| 5 |
A |
1479 |
1411 |
2.10 |
7.26 |
0.69 |
0.82 |
| 6 |
A |
1470 |
1403 |
11.63 |
16.25 |
0.75 |
0.86 |
| 7 |
A |
1348 |
1287 |
24.22 |
4.33 |
0.75 |
0.86 |
| 8 |
A |
1300 |
1241 |
45.35 |
1.44 |
0.52 |
0.69 |
| 9 |
A |
1224 |
1168 |
3.20 |
16.95 |
0.75 |
0.85 |
| 10 |
A |
1157 |
1104 |
1.57 |
6.03 |
0.73 |
0.85 |
| 11 |
A |
1057 |
1009 |
1.94 |
3.78 |
0.72 |
0.84 |
| 12 |
A |
945 |
901 |
9.32 |
6.36 |
0.43 |
0.60 |
| 13 |
A |
876 |
836 |
22.30 |
1.62 |
0.72 |
0.84 |
| 14 |
A |
695 |
664 |
19.16 |
9.18 |
0.40 |
0.57 |
| 15 |
A |
590 |
563 |
11.65 |
14.52 |
0.20 |
0.34 |
| 16 |
A |
392 |
374 |
8.11 |
2.36 |
0.71 |
0.83 |
| 17 |
A |
244 |
233 |
1.28 |
1.01 |
0.33 |
0.50 |
| 18 |
A |
102 |
98 |
0.56 |
1.22 |
0.73 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 12743.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12160.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.005 |
1.114 |
-0.379 |
| C2 |
1.279 |
0.903 |
0.398 |
| Br3 |
-1.342 |
-0.227 |
0.036 |
| Cl4 |
2.145 |
-0.581 |
-0.087 |
| H5 |
-0.433 |
2.078 |
-0.115 |
| H6 |
0.171 |
1.066 |
-1.455 |
| H7 |
1.957 |
1.742 |
0.215 |
| H8 |
1.079 |
0.830 |
1.468 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5074 | 1.9453 | 2.7461 | 1.0916 | 1.0897 | 2.1353 | 2.1560 |
C2 | 1.5074 | | 2.8770 | 1.7861 | 2.1393 | 2.1654 | 1.0937 | 1.0910 | Br3 | 1.9453 | 2.8770 | | 3.5070 | 2.4822 | 2.4865 | 3.8459 | 3.0050 | Cl4 | 2.7461 | 1.7861 | 3.5070 | | 3.7046 | 2.9123 | 2.3504 | 2.3552 | H5 | 1.0916 | 2.1393 | 2.4822 | 3.7046 | | 1.7851 | 2.4369 | 2.5207 | H6 | 1.0897 | 2.1654 | 2.4865 | 2.9123 | 1.7851 | | 2.5375 | 3.0703 | H7 | 2.1353 | 1.0937 | 3.8459 | 2.3504 | 2.4369 | 2.5375 | | 1.7808 | H8 | 2.1560 | 1.0910 | 3.0050 | 2.3552 | 2.5207 | 3.0703 | 1.7808 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.703 |
|
C1 |
C2 |
H7 |
109.320 |
| C1 |
C2 |
H8 |
111.131 |
|
C2 |
C1 |
Br3 |
112.253 |
| C2 |
C1 |
Cl4 |
36.872 |
|
C2 |
C1 |
H6 |
111.967 |
| Br3 |
C1 |
H5 |
106.209 |
|
Br3 |
C1 |
H6 |
106.602 |
| Cl4 |
C2 |
H7 |
106.934 |
|
Cl4 |
C2 |
H8 |
107.412 |
| H5 |
C1 |
H6 |
109.842 |
|
H7 |
C2 |
H8 |
109.199 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.268 |
|
|
|
| 2 |
C |
-0.350 |
|
|
|
| 3 |
Br |
-0.111 |
|
|
|
| 4 |
Cl |
-0.047 |
|
|
|
| 5 |
H |
0.187 |
|
|
|
| 6 |
H |
0.198 |
|
|
|
| 7 |
H |
0.190 |
|
|
|
| 8 |
H |
0.202 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.305 |
2.771 |
0.124 |
2.790 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.138 |
2.881 |
0.458 |
| y |
2.881 |
-40.015 |
-0.131 |
| z |
0.458 |
-0.131 |
-42.716 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.772 |
2.881 |
0.458 |
| y |
2.881 |
4.912 |
-0.131 |
| z |
0.458 |
-0.131 |
0.860 |
|
| Polar |
|---|
| 3z2-r2 | 1.720 |
|---|
| x2-y2 | -7.123 |
|---|
| xy | 2.881 |
|---|
| xz | 0.458 |
|---|
| yz | -0.131 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.071 |
0.197 |
-0.282 |
| y |
0.197 |
7.200 |
0.082 |
| z |
-0.282 |
0.082 |
5.350 |
<r2> (average value of r
2) Å
2
| <r2> |
218.693 |
| (<r2>)1/2 |
14.788 |
