Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -998.588678 |
| Energy at 298.15K | -998.593533 |
| HF Energy | -998.588678 |
| Nuclear repulsion energy | 202.109105 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3184 |
3034 |
0.58 |
|
|
|
| 2 |
A |
3124 |
2978 |
20.17 |
|
|
|
| 3 |
A |
1497 |
1426 |
0.57 |
|
|
|
| 4 |
A |
1373 |
1309 |
23.53 |
|
|
|
| 5 |
A |
1259 |
1200 |
1.08 |
|
|
|
| 6 |
A |
1075 |
1025 |
0.93 |
|
|
|
| 7 |
A |
977 |
931 |
14.00 |
|
|
|
| 8 |
A |
688 |
656 |
19.40 |
|
|
|
| 9 |
A |
264 |
251 |
0.93 |
|
|
|
| 10 |
A |
118 |
112 |
0.99 |
|
|
|
| 11 |
B |
3196 |
3046 |
4.13 |
|
|
|
| 12 |
B |
3117 |
2970 |
3.33 |
|
|
|
| 13 |
B |
1492 |
1422 |
14.11 |
|
|
|
| 14 |
B |
1346 |
1283 |
39.20 |
|
|
|
| 15 |
B |
1191 |
1135 |
1.32 |
|
|
|
| 16 |
B |
914 |
871 |
23.12 |
|
|
|
| 17 |
B |
713 |
679 |
25.39 |
|
|
|
| 18 |
B |
417 |
397 |
8.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12971.5 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12363.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.293 |
0.696 |
0.889 |
| C2 |
-0.293 |
-0.696 |
0.889 |
| Cl3 |
-0.293 |
1.695 |
-0.469 |
| Cl4 |
0.293 |
-1.695 |
-0.469 |
| H5 |
-0.000 |
1.205 |
1.812 |
| H6 |
1.382 |
0.665 |
0.829 |
| H7 |
0.000 |
-1.205 |
1.812 |
| H8 |
-1.382 |
-0.665 |
0.829 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5103 | 1.7846 | 2.7497 | 1.0938 | 1.0917 | 2.1331 | 2.1594 |
C2 | 1.5103 | | 2.7497 | 1.7846 | 2.1331 | 2.1594 | 1.0938 | 1.0917 | Cl3 | 1.7846 | 2.7497 | | 3.4398 | 2.3516 | 2.3563 | 3.7008 | 2.9056 | Cl4 | 2.7497 | 1.7846 | 3.4398 | | 3.7008 | 2.9056 | 2.3516 | 2.3563 | H5 | 1.0938 | 2.1331 | 2.3516 | 3.7008 | | 1.7801 | 2.4091 | 2.5244 | H6 | 1.0917 | 2.1594 | 2.3563 | 2.9056 | 1.7801 | | 2.5244 | 3.0683 | H7 | 2.1331 | 1.0938 | 3.7008 | 2.3516 | 2.4091 | 2.5244 | | 1.7801 | H8 | 2.1594 | 1.0917 | 2.9056 | 2.3563 | 2.5244 | 3.0683 | 1.7801 | |
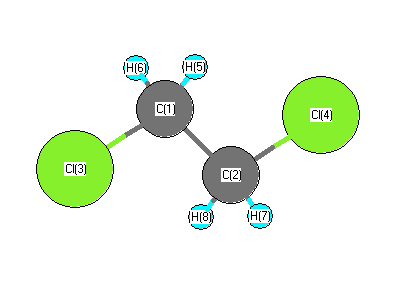 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.866 |
|
C1 |
C2 |
H7 |
108.944 |
| C1 |
C2 |
H8 |
111.159 |
|
C2 |
C1 |
Cl3 |
112.866 |
| C2 |
C1 |
H5 |
108.944 |
|
C2 |
C1 |
H6 |
111.159 |
| Cl3 |
C1 |
H5 |
107.108 |
|
Cl3 |
C1 |
H6 |
107.556 |
| Cl4 |
C2 |
H7 |
107.108 |
|
Cl4 |
C2 |
H8 |
107.556 |
| H5 |
C1 |
H6 |
109.071 |
|
H7 |
C2 |
H8 |
109.071 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.434 |
|
|
|
| 2 |
C |
-0.434 |
|
|
|
| 3 |
Cl |
-0.042 |
|
|
|
| 4 |
Cl |
-0.042 |
|
|
|
| 5 |
H |
0.232 |
|
|
|
| 6 |
H |
0.244 |
|
|
|
| 7 |
H |
0.232 |
|
|
|
| 8 |
H |
0.244 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.914 |
2.914 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.223 |
1.230 |
0.000 |
| y |
1.230 |
-42.185 |
0.000 |
| z |
0.000 |
0.000 |
-35.100 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.419 |
1.230 |
0.000 |
| y |
1.230 |
-6.023 |
0.000 |
| z |
0.000 |
0.000 |
4.604 |
|
| Polar |
|---|
| 3z2-r2 | 9.209 |
|---|
| x2-y2 | 4.961 |
|---|
| xy | 1.230 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.664 |
-0.583 |
0.000 |
| y |
-0.583 |
6.631 |
0.000 |
| z |
0.000 |
0.000 |
6.255 |
<r2> (average value of r
2) Å
2
| <r2> |
163.774 |
| (<r2>)1/2 |
12.797 |
