Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -998.634720 |
| Energy at 298.15K | -998.639561 |
| HF Energy | -998.634720 |
| Nuclear repulsion energy | 202.038317 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3032 |
0.65 |
|
|
|
| 2 |
A |
3115 |
2973 |
19.83 |
|
|
|
| 3 |
A |
1480 |
1412 |
0.70 |
|
|
|
| 4 |
A |
1360 |
1298 |
22.63 |
|
|
|
| 5 |
A |
1248 |
1191 |
1.01 |
|
|
|
| 6 |
A |
1069 |
1020 |
1.11 |
|
|
|
| 7 |
A |
969 |
925 |
13.65 |
|
|
|
| 8 |
A |
684 |
653 |
19.62 |
|
|
|
| 9 |
A |
264 |
252 |
0.96 |
|
|
|
| 10 |
A |
117 |
112 |
0.99 |
|
|
|
| 11 |
B |
3190 |
3044 |
3.95 |
|
|
|
| 12 |
B |
3108 |
2966 |
3.46 |
|
|
|
| 13 |
B |
1475 |
1408 |
13.61 |
|
|
|
| 14 |
B |
1331 |
1270 |
38.68 |
|
|
|
| 15 |
B |
1179 |
1125 |
1.15 |
|
|
|
| 16 |
B |
907 |
865 |
22.72 |
|
|
|
| 17 |
B |
708 |
675 |
25.81 |
|
|
|
| 18 |
B |
416 |
397 |
8.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12897.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 12307.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.293 |
0.695 |
0.890 |
| C2 |
-0.293 |
-0.695 |
0.890 |
| Cl3 |
-0.293 |
1.696 |
-0.469 |
| Cl4 |
0.293 |
-1.696 |
-0.469 |
| H5 |
-0.000 |
1.205 |
1.811 |
| H6 |
1.382 |
0.665 |
0.828 |
| H7 |
0.000 |
-1.205 |
1.811 |
| H8 |
-1.382 |
-0.665 |
0.828 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5092 | 1.7862 | 2.7504 | 1.0931 | 1.0910 | 2.1321 | 2.1586 |
C2 | 1.5092 | | 2.7504 | 1.7862 | 2.1321 | 2.1586 | 1.0931 | 1.0910 | Cl3 | 1.7862 | 2.7504 | | 3.4418 | 2.3514 | 2.3560 | 3.7014 | 2.9059 | Cl4 | 2.7504 | 1.7862 | 3.4418 | | 3.7014 | 2.9059 | 2.3514 | 2.3560 | H5 | 1.0931 | 2.1321 | 2.3514 | 3.7014 | | 1.7797 | 2.4093 | 2.5242 | H6 | 1.0910 | 2.1586 | 2.3560 | 2.9059 | 1.7797 | | 2.5242 | 3.0671 | H7 | 2.1321 | 1.0931 | 3.7014 | 2.3514 | 2.4093 | 2.5242 | | 1.7797 | H8 | 2.1586 | 1.0910 | 2.9059 | 2.3560 | 2.5242 | 3.0671 | 1.7797 | |
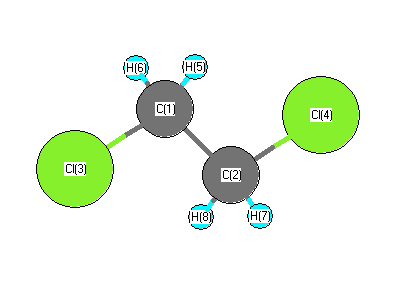 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.882 |
|
C1 |
C2 |
H7 |
108.981 |
| C1 |
C2 |
H8 |
111.212 |
|
C2 |
C1 |
Cl3 |
112.882 |
| C2 |
C1 |
H5 |
108.981 |
|
C2 |
C1 |
H6 |
111.212 |
| Cl3 |
C1 |
H5 |
107.023 |
|
Cl3 |
C1 |
H6 |
107.464 |
| Cl4 |
C2 |
H7 |
107.023 |
|
Cl4 |
C2 |
H8 |
107.464 |
| H5 |
C1 |
H6 |
109.137 |
|
H7 |
C2 |
H8 |
109.137 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.348 |
|
|
|
| 2 |
C |
-0.348 |
|
|
|
| 3 |
Cl |
-0.047 |
|
|
|
| 4 |
Cl |
-0.047 |
|
|
|
| 5 |
H |
0.192 |
|
|
|
| 6 |
H |
0.203 |
|
|
|
| 7 |
H |
0.192 |
|
|
|
| 8 |
H |
0.203 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.908 |
2.908 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.216 |
1.242 |
0.000 |
| y |
1.242 |
-42.150 |
0.000 |
| z |
0.000 |
0.000 |
-35.111 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.414 |
1.242 |
0.000 |
| y |
1.242 |
-5.986 |
0.000 |
| z |
0.000 |
0.000 |
4.572 |
|
| Polar |
|---|
| 3z2-r2 | 9.144 |
|---|
| x2-y2 | 4.933 |
|---|
| xy | 1.242 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.711 |
-0.581 |
0.000 |
| y |
-0.581 |
6.703 |
0.000 |
| z |
0.000 |
0.000 |
6.313 |
<r2> (average value of r
2) Å
2
| <r2> |
163.884 |
| (<r2>)1/2 |
12.802 |
