return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2ClCH2OH (2-Chloroethanol)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-613.630888
Energy at 298.15K-613.636865
HF Energy-612.989046
Nuclear repulsion energy159.211797
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3916 3687 25.82      
2 A' 3174 2988 17.16      
3 A' 3082 2901 37.32      
4 A' 1592 1498 2.26      
5 A' 1552 1461 4.82      
6 A' 1507 1419 3.59      
7 A' 1358 1279 1.33      
8 A' 1267 1193 69.28      
9 A' 1113 1048 78.85      
10 A' 1061 999 9.96      
11 A' 811 763 65.69      
12 A' 400 377 2.64      
13 A' 255 240 9.83      
14 A" 3243 3053 10.86      
15 A" 3127 2944 45.84      
16 A" 1343 1264 0.05      
17 A" 1247 1174 1.12      
18 A" 1111 1046 3.31      
19 A" 823 775 0.07      
20 A" 235 221 131.89      
21 A" 135 127 15.32      

Unscaled Zero Point Vibrational Energy (zpe) 16176.0 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 15228.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.98649 0.08168 0.07764

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.967 -0.565 0.000
C2 0.000 0.604 0.000
Cl3 -1.690 0.016 0.000
O4 2.266 0.015 0.000
H5 0.797 -1.184 0.886
H6 0.797 -1.184 -0.886
H7 0.146 1.216 0.886
H8 0.146 1.216 -0.886
H9 2.911 -0.699 0.000

Atom - Atom Distances (Å)
  C1 C2 Cl3 O4 H5 H6 H7 H8 H9
C11.51702.72021.42251.09481.09482.15162.15161.9484
C21.51701.78932.34182.14902.14901.08661.08663.1894
Cl32.72021.78933.95652.90012.90012.36502.36504.6564
O41.42252.34183.95652.09382.09382.59322.59320.9620
H51.09482.14902.90012.09381.77262.48653.05372.3434
H61.09482.14902.90012.09381.77263.05372.48652.3434
H72.15161.08662.36502.59322.48653.05371.77273.4783
H82.15161.08662.36502.59323.05372.48651.77273.4783
H91.94843.18944.65640.96202.34342.34343.47833.4783

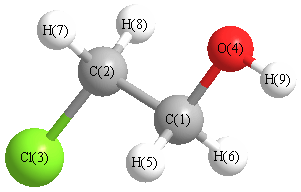
picture of 2-Chloroethanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 Cl3 110.445 C1 C2 H7 110.364
C1 C2 H8 110.364 C1 O4 H9 108.030
C2 C1 O4 105.574 C2 C1 H5 109.668
C2 C1 H6 109.668 Cl3 C2 H7 108.146
Cl3 C2 H8 108.146 O4 C1 H5 111.899
O4 C1 H6 111.899 H5 C1 H6 108.103
H7 C2 H8 109.313
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability