return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H8N2 (Ethylenediamine)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-189.945674
Energy at 298.15K-189.956030
HF Energy-189.286444
Nuclear repulsion energy131.973673
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3568 3334 0.00      
2 Ag 3114 2909 0.00      
3 Ag 1704 1592 0.00      
4 Ag 1541 1440 0.00      
5 Ag 1445 1351 0.00      
6 Ag 1161 1085 0.00      
7 Ag 1110 1037 0.00      
8 Ag 885 827 0.00      
9 Ag 474 442 0.00      
10 Au 3671 3430 0.11      
11 Au 3189 2979 65.45      
12 Au 1433 1339 2.06      
13 Au 1098 1026 0.60      
14 Au 775 724 0.17      
15 Au 290 271 122.08      
16 Au 159 149 0.20      
17 Bg 3671 3430 0.00      
18 Bg 3165 2957 0.00      
19 Bg 1410 1318 0.00      
20 Bg 1343 1255 0.00      
21 Bg 969 905 0.00      
22 Bg 325 304 0.00      
23 Bu 3568 3334 0.40      
24 Bu 3121 2916 77.52      
25 Bu 1702 1590 47.33      
26 Bu 1558 1456 0.28      
27 Bu 1365 1275 27.79      
28 Bu 1127 1054 7.12      
29 Bu 923 862 405.92      
30 Bu 279 261 20.41      

Unscaled Zero Point Vibrational Energy (zpe) 25070.6 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 23426.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.88327 0.12550 0.11875

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.397 0.653 0.000
C2 -0.397 -0.653 0.000
N3 -0.397 1.879 0.000
N4 0.397 -1.879 0.000
H5 1.008 -1.882 0.809
H6 1.008 -1.882 -0.809
H7 -1.008 1.882 0.809
H8 -1.008 1.882 -0.809
H9 -1.050 -0.674 -0.875
H10 -1.050 -0.674 0.875
H11 1.050 0.674 -0.875
H12 1.050 0.674 0.875

Atom - Atom Distances (Å)
  C1 C2 N3 N4 H5 H6 H7 H8 H9 H10 H11 H12
C11.52901.46052.53222.73022.73022.03452.03452.14982.14981.09201.0920
C21.52902.53221.46052.03452.03452.73022.73021.09201.09202.14982.1498
N31.46052.53223.84094.09534.09531.01401.01402.77642.77642.07672.0767
N42.53221.46053.84091.01401.01404.09534.09532.07672.07672.77642.7764
H52.73022.03454.09531.01401.61824.26964.56602.92112.38773.06072.5566
H62.73022.03454.09531.01401.61824.56604.26962.38772.92112.55663.0607
H72.03452.73021.01404.09534.26964.56601.61823.06072.55662.92112.3877
H82.03452.73021.01404.09534.56604.26961.61822.55663.06072.38772.9211
H92.14981.09202.77642.07672.92112.38773.06072.55661.74992.49553.0480
H102.14981.09202.77642.07672.38772.92112.55663.06071.74993.04802.4955
H111.09202.14982.07672.77643.06072.55662.92112.38772.49553.04801.7499
H121.09202.14982.07672.77642.55663.06072.38772.92113.04802.49551.7499

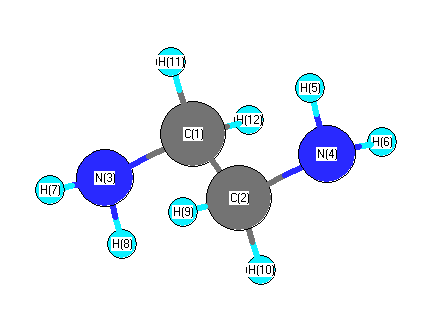
picture of Ethylenediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N4 115.756 C1 C2 H9 109.070
C1 C2 H10 109.070 C1 N3 H7 109.273
C1 N3 H8 109.273 C2 C1 N3 115.756
C2 C1 H11 109.070 C2 C1 H12 109.070
C2 N4 H5 109.273 C2 N4 H6 109.273
N3 C1 H11 108.027 N3 C1 H12 108.027
N4 C2 H9 108.027 N4 C2 H10 108.027
H5 N4 H6 105.860 H7 N3 H8 105.860
H9 C2 H10 106.499 H11 C1 H12 106.499
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability