Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G**
Geometric Data calculated at QCISD/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -207.401003 |
| Energy at 298.15K | -207.404366 |
| HF Energy | -206.772300 |
| Nuclear repulsion energy | 101.680810 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3900 |
3671 |
33.27 |
|
|
|
| 2 |
A |
3196 |
3009 |
7.81 |
|
|
|
| 3 |
A |
3109 |
2927 |
27.32 |
|
|
|
| 4 |
A |
2344 |
2207 |
2.41 |
|
|
|
| 5 |
A |
1551 |
1460 |
0.77 |
|
|
|
| 6 |
A |
1452 |
1366 |
52.34 |
|
|
|
| 7 |
A |
1417 |
1334 |
2.85 |
|
|
|
| 8 |
A |
1250 |
1177 |
19.69 |
|
|
|
| 9 |
A |
1119 |
1054 |
91.53 |
|
|
|
| 10 |
A |
1013 |
953 |
26.98 |
|
|
|
| 11 |
A |
915 |
861 |
22.08 |
|
|
|
| 12 |
A |
577 |
543 |
3.08 |
|
|
|
| 13 |
A |
393 |
370 |
84.11 |
|
|
|
| 14 |
A |
308 |
290 |
77.55 |
|
|
|
| 15 |
A |
210 |
198 |
8.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11376.0 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 10709.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.579 |
0.594 |
0.037 |
| C2 |
0.827 |
0.116 |
-0.006 |
| O3 |
-1.509 |
-0.460 |
-0.114 |
| H4 |
-0.722 |
1.152 |
0.967 |
| H5 |
-0.742 |
1.277 |
-0.794 |
| H6 |
-1.428 |
-1.043 |
0.648 |
| N7 |
1.925 |
-0.280 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4856 | 1.4130 | 1.0938 | 1.0885 | 1.9423 | 2.6531 |
C2 | 1.4856 | | 2.4077 | 2.1020 | 2.1049 | 2.6181 | 1.1677 | O3 | 1.4130 | 2.4077 | | 2.0941 | 2.0165 | 0.9634 | 3.4400 | H4 | 1.0938 | 2.1020 | 2.0941 | | 1.7660 | 2.3274 | 3.1657 | H5 | 1.0885 | 2.1049 | 2.0165 | 1.7660 | | 2.8167 | 3.1855 | H6 | 1.9423 | 2.6181 | 0.9634 | 2.3274 | 2.8167 | | 3.5022 | N7 | 2.6531 | 1.1677 | 3.4400 | 3.1657 | 3.1855 | 3.5022 | |
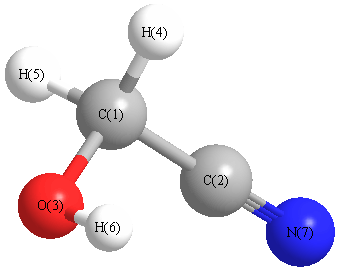 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.370 |
|
C1 |
O3 |
H6 |
108.147 |
| C2 |
C1 |
O3 |
112.302 |
|
C2 |
C1 |
H4 |
108.198 |
| C2 |
C1 |
H5 |
108.739 |
|
O3 |
C1 |
H4 |
112.688 |
| O3 |
C1 |
H5 |
106.717 |
|
H4 |
C1 |
H5 |
108.048 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
