Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -207.739477 |
| Energy at 298.15K | -207.742861 |
| HF Energy | -207.739477 |
| Nuclear repulsion energy | 102.298754 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3876 |
3699 |
40.65 |
65.71 |
0.27 |
0.43 |
| 2 |
A |
3132 |
2988 |
4.30 |
82.65 |
0.42 |
0.59 |
| 3 |
A |
3039 |
2900 |
25.78 |
112.22 |
0.15 |
0.27 |
| 4 |
A |
2395 |
2285 |
0.42 |
53.49 |
0.27 |
0.43 |
| 5 |
A |
1492 |
1423 |
3.62 |
15.48 |
0.66 |
0.80 |
| 6 |
A |
1420 |
1355 |
47.99 |
8.00 |
0.74 |
0.85 |
| 7 |
A |
1384 |
1321 |
1.33 |
7.01 |
0.73 |
0.84 |
| 8 |
A |
1229 |
1173 |
15.57 |
5.06 |
0.60 |
0.75 |
| 9 |
A |
1123 |
1071 |
108.11 |
5.13 |
0.31 |
0.48 |
| 10 |
A |
991 |
946 |
22.15 |
0.63 |
0.42 |
0.59 |
| 11 |
A |
914 |
872 |
16.19 |
2.04 |
0.17 |
0.29 |
| 12 |
A |
592 |
565 |
2.33 |
1.72 |
0.32 |
0.49 |
| 13 |
A |
402 |
383 |
62.97 |
1.29 |
0.72 |
0.84 |
| 14 |
A |
314 |
299 |
96.66 |
5.11 |
0.75 |
0.86 |
| 15 |
A |
215 |
205 |
8.87 |
4.56 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11257.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 10743.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.573 |
0.585 |
0.038 |
| C2 |
0.822 |
0.118 |
-0.007 |
| O3 |
-1.503 |
-0.453 |
-0.112 |
| H4 |
-0.710 |
1.155 |
0.969 |
| H5 |
-0.734 |
1.279 |
-0.793 |
| H6 |
-1.400 |
-1.062 |
0.628 |
| N7 |
1.910 |
-0.280 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4724 | 1.4014 | 1.1001 | 1.0939 | 1.9344 | 2.6307 |
C2 | 1.4724 | | 2.3963 | 2.0914 | 2.0943 | 2.5943 | 1.1587 | O3 | 1.4014 | 2.3963 | | 2.0935 | 2.0138 | 0.9631 | 3.4189 | H4 | 1.1001 | 2.0914 | 2.0935 | | 1.7660 | 2.3462 | 3.1451 | H5 | 1.0939 | 2.0943 | 2.0138 | 1.7660 | | 2.8176 | 3.1665 | H6 | 1.9344 | 2.5943 | 0.9631 | 2.3462 | 2.8176 | | 3.4612 | N7 | 2.6307 | 1.1587 | 3.4189 | 3.1451 | 3.1665 | 3.4612 | |
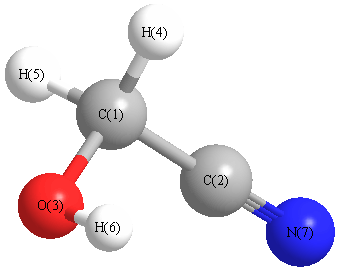 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
177.918 |
|
C1 |
O3 |
H6 |
108.361 |
| C2 |
C1 |
O3 |
112.970 |
|
C2 |
C1 |
H4 |
107.902 |
| C2 |
C1 |
H5 |
108.485 |
|
O3 |
C1 |
H4 |
113.077 |
| O3 |
C1 |
H5 |
106.966 |
|
H4 |
C1 |
H5 |
107.203 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.039 |
|
|
|
| 2 |
C |
0.320 |
|
|
|
| 3 |
O |
-0.518 |
|
|
|
| 4 |
H |
0.167 |
|
|
|
| 5 |
H |
0.194 |
|
|
|
| 6 |
H |
0.335 |
|
|
|
| 7 |
N |
-0.461 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.418 |
1.111 |
1.367 |
2.992 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.231 |
1.221 |
-1.974 |
| y |
1.221 |
-20.273 |
-1.782 |
| z |
-1.974 |
-1.782 |
-21.462 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.363 |
1.221 |
-1.974 |
| y |
1.221 |
6.073 |
-1.782 |
| z |
-1.974 |
-1.782 |
4.290 |
|
| Polar |
|---|
| 3z2-r2 | 8.580 |
|---|
| x2-y2 | -10.958 |
|---|
| xy | 1.221 |
|---|
| xz | -1.974 |
|---|
| yz | -1.782 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.299 |
-0.474 |
-0.039 |
| y |
-0.474 |
3.514 |
-0.137 |
| z |
-0.039 |
-0.137 |
3.007 |
<r2> (average value of r
2) Å
2
| <r2> |
78.331 |
| (<r2>)1/2 |
8.850 |
