Jump to
S1C2
Vibrational Frequencies calculated at CCD/6-31G**
Geometric Data calculated at CCD/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -207.393665 |
| Energy at 298.15K | -207.397042 |
| HF Energy | -206.773029 |
| Nuclear repulsion energy | 101.847209 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3923 |
3678 |
35.38 |
|
|
|
| 2 |
A |
3206 |
3006 |
7.37 |
|
|
|
| 3 |
A |
3121 |
2926 |
25.69 |
|
|
|
| 4 |
A |
2381 |
2233 |
2.79 |
|
|
|
| 5 |
A |
1555 |
1458 |
0.87 |
|
|
|
| 6 |
A |
1457 |
1366 |
52.42 |
|
|
|
| 7 |
A |
1423 |
1334 |
3.43 |
|
|
|
| 8 |
A |
1256 |
1177 |
19.10 |
|
|
|
| 9 |
A |
1131 |
1061 |
91.64 |
|
|
|
| 10 |
A |
1017 |
953 |
26.10 |
|
|
|
| 11 |
A |
924 |
866 |
22.26 |
|
|
|
| 12 |
A |
583 |
546 |
2.84 |
|
|
|
| 13 |
A |
393 |
369 |
82.13 |
|
|
|
| 14 |
A |
310 |
290 |
80.64 |
|
|
|
| 15 |
A |
212 |
199 |
8.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11446.0 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 10731.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.578 |
0.592 |
0.037 |
| C2 |
0.826 |
0.115 |
-0.005 |
| O3 |
-1.507 |
-0.459 |
-0.114 |
| H4 |
-0.721 |
1.150 |
0.966 |
| H5 |
-0.739 |
1.276 |
-0.793 |
| H6 |
-1.427 |
-1.044 |
0.645 |
| N7 |
1.922 |
-0.279 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4836 | 1.4111 | 1.0932 | 1.0880 | 1.9415 | 2.6474 |
C2 | 1.4836 | | 2.4052 | 2.1004 | 2.1028 | 2.6159 | 1.1640 | O3 | 1.4111 | 2.4052 | | 2.0913 | 2.0149 | 0.9623 | 3.4348 | H4 | 1.0932 | 2.1004 | 2.0913 | | 1.7644 | 2.3273 | 3.1606 | H5 | 1.0880 | 2.1028 | 2.0149 | 1.7644 | | 2.8154 | 3.1794 | H6 | 1.9415 | 2.6159 | 0.9623 | 2.3273 | 2.8154 | | 3.4974 | N7 | 2.6474 | 1.1640 | 3.4348 | 3.1606 | 3.1794 | 3.4974 | |
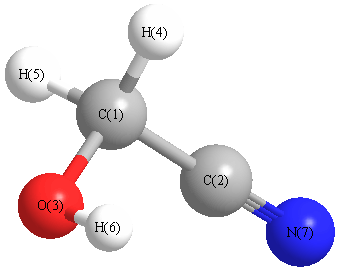 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.428 |
|
C1 |
O3 |
H6 |
108.279 |
| C2 |
C1 |
O3 |
112.351 |
|
C2 |
C1 |
H4 |
108.244 |
| C2 |
C1 |
H5 |
108.734 |
|
O3 |
C1 |
H4 |
112.630 |
| O3 |
C1 |
H5 |
106.746 |
|
H4 |
C1 |
H5 |
107.983 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
