return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-192.568072
Energy at 298.15K-192.574336
HF Energy-191.928954
Nuclear repulsion energy116.479976
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3905 3676 14.15      
2 A 3309 3115 19.05      
3 A 3239 3049 6.98      
4 A 3215 3027 11.85      
5 A 3091 2910 58.23      
6 A 3048 2869 59.33      
7 A 1758 1655 0.43      
8 A 1578 1486 1.69      
9 A 1523 1434 13.68      
10 A 1468 1382 6.91      
11 A 1341 1263 0.60      
12 A 1304 1227 29.32      
13 A 1270 1196 58.77      
14 A 1192 1122 22.11      
15 A 1107 1042 72.05      
16 A 1036 975 15.88      
17 A 998 939 6.58      
18 A 948 892 28.41      
19 A 942 887 4.62      
20 A 657 618 3.81      
21 A 452 426 4.11      
22 A 337 317 10.07      
23 A 270 254 130.12      
24 A 111 105 1.30      

Unscaled Zero Point Vibrational Energy (zpe) 19049.3 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 17933.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.92520 0.14362 0.13770

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.588 0.458 0.283
C2 -0.666 -0.364 0.255
C3 -1.836 0.077 -0.217
O4 1.625 -0.308 -0.328
H5 0.424 1.406 -0.243
H6 0.847 0.697 1.323
H7 -0.575 -1.366 0.661
H8 -2.725 -0.539 -0.197
H9 -1.940 1.070 -0.637
H10 2.451 0.171 -0.210

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.49972.50451.42761.09661.09822.19603.49332.75931.9487
C21.49971.33692.36472.13762.13521.08472.11582.11593.1966
C32.50451.33693.48492.62223.15582.10781.08151.08334.2887
O41.42762.36473.48492.09552.08422.63404.35873.83530.9624
H51.09662.13762.62222.09551.77053.08213.70162.42022.3742
H61.09822.13523.15582.08421.77052.59234.07503.42832.2802
H72.19601.08472.10782.63403.08212.59232.45863.07933.5041
H83.49332.11581.08154.35873.70164.07502.45861.84325.2250
H92.75932.11591.08333.83532.42023.42833.07931.84324.5029
H101.94873.19664.28870.96242.37422.28023.50415.22504.5029

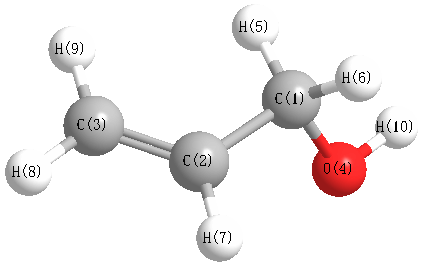
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.898 C1 C2 H7 115.431
C1 O4 H10 107.659 C2 C1 O4 107.743
C2 C1 H5 109.867 C2 C1 H6 109.581
C2 C3 H8 121.705 C2 C3 H9 121.559
C3 C2 H7 120.667 O4 C1 H5 111.558
O4 C1 H6 110.542 H5 C1 H6 107.548
H8 C3 H9 116.735
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability