Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -613.113775 |
| Energy at 298.15K | -613.116974 |
| HF Energy | -613.113775 |
| Nuclear repulsion energy | 142.775407 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3182 |
3037 |
0.21 |
|
|
|
| 2 |
A |
3108 |
2966 |
8.63 |
|
|
|
| 3 |
A |
2968 |
2832 |
63.17 |
|
|
|
| 4 |
A |
1873 |
1788 |
145.97 |
|
|
|
| 5 |
A |
1445 |
1379 |
15.69 |
|
|
|
| 6 |
A |
1413 |
1348 |
6.79 |
|
|
|
| 7 |
A |
1278 |
1220 |
21.53 |
|
|
|
| 8 |
A |
1200 |
1145 |
1.60 |
|
|
|
| 9 |
A |
1054 |
1005 |
41.44 |
|
|
|
| 10 |
A |
1043 |
995 |
3.82 |
|
|
|
| 11 |
A |
822 |
784 |
41.45 |
|
|
|
| 12 |
A |
693 |
661 |
6.39 |
|
|
|
| 13 |
A |
464 |
443 |
9.03 |
|
|
|
| 14 |
A |
271 |
259 |
12.84 |
|
|
|
| 15 |
A |
37 |
35 |
18.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10424.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 9948.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.027 |
0.641 |
0.079 |
| C2 |
1.161 |
-0.365 |
0.094 |
| Cl3 |
-1.564 |
-0.150 |
-0.043 |
| O4 |
2.299 |
-0.031 |
-0.110 |
| H5 |
0.050 |
1.219 |
1.008 |
| H6 |
0.155 |
1.327 |
-0.760 |
| H7 |
0.873 |
-1.412 |
0.323 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5161 | 1.7808 | 2.3771 | 1.0941 | 1.0914 | 2.2341 |
C2 | 1.5161 | | 2.7373 | 1.2031 | 2.1392 | 2.1459 | 1.1101 | Cl3 | 1.7808 | 2.7373 | | 3.8655 | 2.3631 | 2.3768 | 2.7694 | O4 | 2.3771 | 1.2031 | 3.8655 | | 2.8055 | 2.6200 | 2.0314 | H5 | 1.0941 | 2.1392 | 2.3631 | 2.8055 | | 1.7743 | 2.8404 | H6 | 1.0914 | 2.1459 | 2.3768 | 2.6200 | 1.7743 | | 3.0319 | H7 | 2.2341 | 1.1101 | 2.7694 | 2.0314 | 2.8404 | 3.0319 | |
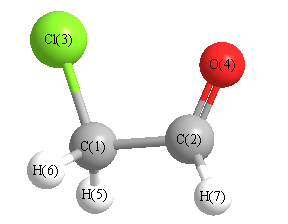 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
121.479 |
|
C1 |
C2 |
H7 |
115.715 |
| C2 |
C1 |
Cl3 |
112.008 |
|
C2 |
C1 |
H5 |
109.010 |
| C2 |
C1 |
H6 |
109.690 |
|
Cl3 |
C1 |
H5 |
108.181 |
| Cl3 |
C1 |
H6 |
109.331 |
|
O4 |
C2 |
H7 |
122.797 |
| H5 |
C1 |
H6 |
108.546 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.430 |
|
|
|
| 2 |
C |
0.292 |
|
|
|
| 3 |
Cl |
-0.051 |
|
|
|
| 4 |
O |
-0.373 |
|
|
|
| 5 |
H |
0.216 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.608 |
0.535 |
0.540 |
0.973 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.846 |
-0.453 |
0.677 |
| y |
-0.453 |
-27.983 |
0.024 |
| z |
0.677 |
0.024 |
-28.998 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.355 |
-0.453 |
0.677 |
| y |
-0.453 |
5.939 |
0.024 |
| z |
0.677 |
0.024 |
4.417 |
|
| Polar |
|---|
| 3z2-r2 | 8.833 |
|---|
| x2-y2 | -10.863 |
|---|
| xy | -0.453 |
|---|
| xz | 0.677 |
|---|
| yz | 0.024 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.657 |
0.559 |
-0.095 |
| y |
0.559 |
4.371 |
-0.121 |
| z |
-0.095 |
-0.121 |
3.221 |
<r2> (average value of r
2) Å
2
| <r2> |
123.519 |
| (<r2>)1/2 |
11.114 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -613.112728 |
| Energy at 298.15K | -613.116190 |
| HF Energy | -613.112728 |
| Nuclear repulsion energy | 147.052421 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3084 |
2943 |
15.78 |
|
|
|
| 2 |
A' |
2918 |
2785 |
97.25 |
|
|
|
| 3 |
A' |
1891 |
1805 |
136.22 |
|
|
|
| 4 |
A' |
1443 |
1377 |
26.98 |
|
|
|
| 5 |
A' |
1416 |
1351 |
12.24 |
|
|
|
| 6 |
A' |
1320 |
1260 |
21.63 |
|
|
|
| 7 |
A' |
961 |
917 |
5.07 |
|
|
|
| 8 |
A' |
788 |
752 |
8.81 |
|
|
|
| 9 |
A' |
648 |
619 |
44.38 |
|
|
|
| 10 |
A' |
213 |
203 |
2.63 |
|
|
|
| 11 |
A" |
3139 |
2996 |
0.54 |
|
|
|
| 12 |
A" |
1214 |
1158 |
1.89 |
|
|
|
| 13 |
A" |
1044 |
996 |
1.86 |
|
|
|
| 14 |
A" |
691 |
660 |
2.72 |
|
|
|
| 15 |
A" |
176 |
168 |
0.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10472.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 9994.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.917 |
0.000 |
| C2 |
1.362 |
0.266 |
0.000 |
| Cl3 |
-1.343 |
-0.231 |
0.000 |
| O4 |
1.580 |
-0.914 |
0.000 |
| H5 |
-0.080 |
1.563 |
0.882 |
| H6 |
-0.080 |
1.563 |
-0.882 |
| H7 |
2.184 |
1.017 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5097 | 1.7673 | 2.4193 | 1.0956 | 1.0956 | 2.1863 |
C2 | 1.5097 | | 2.7502 | 1.2002 | 2.1302 | 2.1302 | 1.1138 | Cl3 | 1.7673 | 2.7502 | | 3.0024 | 2.3646 | 2.3646 | 3.7416 | O4 | 2.4193 | 1.2002 | 3.0024 | | 3.1095 | 3.1095 | 2.0236 | H5 | 1.0956 | 2.1302 | 2.3646 | 3.1095 | | 1.7639 | 2.4900 | H6 | 1.0956 | 2.1302 | 2.3646 | 3.1095 | 1.7639 | | 2.4900 | H7 | 2.1863 | 1.1138 | 3.7416 | 2.0236 | 2.4900 | 2.4900 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.070 |
|
C1 |
C2 |
H7 |
112.007 |
| C2 |
C1 |
Cl3 |
113.894 |
|
C2 |
C1 |
H5 |
108.654 |
| C2 |
C1 |
H6 |
108.654 |
|
Cl3 |
C1 |
H5 |
109.106 |
| Cl3 |
C1 |
H6 |
109.106 |
|
O4 |
C2 |
H7 |
121.923 |
| H5 |
C1 |
H6 |
107.217 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.438 |
|
|
|
| 2 |
C |
0.295 |
|
|
|
| 3 |
Cl |
-0.019 |
|
|
|
| 4 |
O |
-0.351 |
|
|
|
| 5 |
H |
0.203 |
|
|
|
| 6 |
H |
0.203 |
|
|
|
| 7 |
H |
0.107 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.662 |
3.437 |
0.000 |
3.501 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.049 |
3.556 |
0.000 |
| y |
3.556 |
-29.208 |
0.000 |
| z |
0.000 |
0.000 |
-29.088 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.901 |
3.556 |
0.000 |
| y |
3.556 |
1.361 |
0.000 |
| z |
0.000 |
0.000 |
1.540 |
|
| Polar |
|---|
| 3z2-r2 | 3.080 |
|---|
| x2-y2 | -2.842 |
|---|
| xy | 3.556 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.847 |
0.627 |
0.000 |
| y |
0.627 |
5.348 |
0.000 |
| z |
0.000 |
0.000 |
3.165 |
<r2> (average value of r
2) Å
2
| <r2> |
105.915 |
| (<r2>)1/2 |
10.292 |
