Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.543270 |
| Energy at 298.15K | -208.549296 |
| HF Energy | -207.893845 |
| Nuclear repulsion energy | 117.109415 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3881 |
3626 |
77.15 |
|
|
|
| 2 |
A |
3254 |
3040 |
9.21 |
|
|
|
| 3 |
A |
3211 |
3001 |
9.96 |
|
|
|
| 4 |
A |
3132 |
2926 |
14.89 |
|
|
|
| 5 |
A |
1724 |
1610 |
0.73 |
|
|
|
| 6 |
A |
1546 |
1444 |
5.91 |
|
|
|
| 7 |
A |
1481 |
1384 |
1.58 |
|
|
|
| 8 |
A |
1435 |
1341 |
22.55 |
|
|
|
| 9 |
A |
1303 |
1217 |
62.60 |
|
|
|
| 10 |
A |
1185 |
1108 |
4.97 |
|
|
|
| 11 |
A |
1032 |
964 |
101.64 |
|
|
|
| 12 |
A |
930 |
869 |
16.30 |
|
|
|
| 13 |
A |
570 |
533 |
15.78 |
|
|
|
| 14 |
A |
326 |
305 |
1.85 |
|
|
|
| 15 |
A |
3215 |
3004 |
13.60 |
|
|
|
| 16 |
A |
1540 |
1439 |
7.40 |
|
|
|
| 17 |
A |
1107 |
1035 |
0.71 |
|
|
|
| 18 |
A |
923 |
862 |
10.48 |
|
|
|
| 19 |
A |
399 |
373 |
144.19 |
|
|
|
| 20 |
A |
286 |
267 |
3.65 |
|
|
|
| 21 |
A |
202 |
189 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16340.5 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15268.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.300 |
1.298 |
0.000 |
| C2 |
0.000 |
0.570 |
0.000 |
| N3 |
-0.003 |
-0.715 |
0.000 |
| O4 |
1.324 |
-1.203 |
0.000 |
| H5 |
1.180 |
-2.157 |
0.000 |
| H6 |
-2.121 |
0.587 |
0.000 |
| H7 |
-1.384 |
1.936 |
0.878 |
| H8 |
-1.384 |
1.936 |
-0.878 |
| H9 |
0.940 |
1.115 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4898 | 2.3942 | 3.6241 | 4.2527 | 1.0860 | 1.0890 | 1.0890 | 2.2472 |
C2 | 1.4898 | | 1.2845 | 2.2119 | 2.9710 | 2.1206 | 2.1340 | 2.1340 | 1.0869 | N3 | 2.3942 | 1.2845 | | 1.4136 | 1.8655 | 2.4855 | 3.1155 | 3.1155 | 2.0588 | O4 | 3.6241 | 2.2119 | 1.4136 | | 0.9650 | 3.8813 | 4.2372 | 4.2372 | 2.3492 | H5 | 4.2527 | 2.9710 | 1.8655 | 0.9650 | | 4.2922 | 4.9092 | 4.9092 | 3.2808 | H6 | 1.0860 | 2.1206 | 2.4855 | 3.8813 | 4.2922 | | 1.7706 | 1.7706 | 3.1059 | H7 | 1.0890 | 2.1340 | 3.1155 | 4.2372 | 4.9092 | 1.7706 | | 1.7564 | 2.6165 | H8 | 1.0890 | 2.1340 | 3.1155 | 4.2372 | 4.9092 | 1.7706 | 1.7564 | | 2.6165 | H9 | 2.2472 | 1.0869 | 2.0588 | 2.3492 | 3.2808 | 3.1059 | 2.6165 | 2.6165 | |
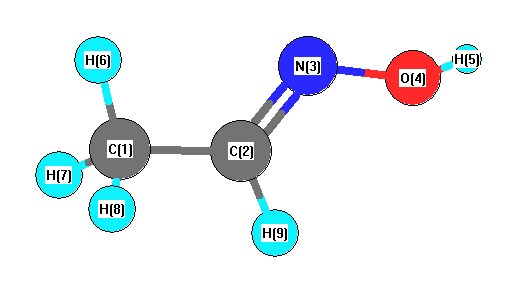 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.121 |
|
C1 |
C2 |
H9 |
120.611 |
| C2 |
C1 |
H6 |
109.835 |
|
C2 |
C1 |
H7 |
110.730 |
| C2 |
C1 |
H8 |
110.730 |
|
C2 |
N3 |
O4 |
110.036 |
| N3 |
C2 |
H9 |
120.268 |
|
N3 |
O4 |
H5 |
101.635 |
| H6 |
C1 |
H7 |
108.993 |
|
H6 |
C1 |
H8 |
108.993 |
| H7 |
C1 |
H8 |
107.500 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.542684 |
| Energy at 298.15K | |
| HF Energy | -207.892560 |
| Nuclear repulsion energy | 119.673784 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3883 |
3628 |
78.98 |
|
|
|
| 2 |
A' |
3283 |
3068 |
4.17 |
|
|
|
| 3 |
A' |
3259 |
3045 |
9.38 |
|
|
|
| 4 |
A' |
3139 |
2933 |
10.95 |
|
|
|
| 5 |
A' |
1733 |
1620 |
2.09 |
|
|
|
| 6 |
A' |
1541 |
1440 |
9.45 |
|
|
|
| 7 |
A' |
1456 |
1361 |
12.86 |
|
|
|
| 8 |
A' |
1399 |
1307 |
15.85 |
|
|
|
| 9 |
A' |
1356 |
1267 |
63.72 |
|
|
|
| 10 |
A' |
1180 |
1103 |
9.60 |
|
|
|
| 11 |
A' |
957 |
894 |
77.61 |
|
|
|
| 12 |
A' |
934 |
873 |
33.21 |
|
|
|
| 13 |
A' |
678 |
633 |
13.58 |
|
|
|
| 14 |
A' |
321 |
299 |
1.68 |
|
|
|
| 15 |
A" |
3217 |
3006 |
11.25 |
|
|
|
| 16 |
A" |
1550 |
1448 |
8.92 |
|
|
|
| 17 |
A" |
1095 |
1024 |
0.55 |
|
|
|
| 18 |
A" |
878 |
821 |
11.79 |
|
|
|
| 19 |
A" |
495 |
462 |
38.32 |
|
|
|
| 20 |
A" |
380 |
355 |
96.96 |
|
|
|
| 21 |
A" |
81i |
75i |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16326.7 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15255.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.443 |
0.485 |
0.000 |
| C2 |
0.000 |
0.867 |
0.000 |
| N3 |
1.026 |
0.090 |
0.000 |
| O4 |
0.642 |
-1.270 |
0.000 |
| H5 |
1.502 |
-1.706 |
0.000 |
| H6 |
-1.560 |
-0.592 |
0.000 |
| H7 |
-1.937 |
0.899 |
0.878 |
| H8 |
-1.937 |
0.899 |
-0.878 |
| H9 |
0.270 |
1.916 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4923 | 2.5000 | 2.7252 | 3.6710 | 1.0832 | 1.0889 | 1.0889 | 2.2318 |
C2 | 1.4923 | | 1.2873 | 2.2320 | 2.9802 | 2.1356 | 2.1268 | 2.1268 | 1.0829 | N3 | 2.5000 | 1.2873 | | 1.4137 | 1.8585 | 2.6740 | 3.1945 | 3.1945 | 1.9762 | O4 | 2.7252 | 2.2320 | 1.4137 | | 0.9648 | 2.3036 | 3.4823 | 3.4823 | 3.2080 | H5 | 3.6710 | 2.9802 | 1.8585 | 0.9648 | | 3.2587 | 4.4032 | 4.4032 | 3.8262 | H6 | 1.0832 | 2.1356 | 2.6740 | 2.3036 | 3.2587 | | 1.7703 | 1.7703 | 3.1043 | H7 | 1.0889 | 2.1268 | 3.1945 | 3.4823 | 4.4032 | 1.7703 | | 1.7551 | 2.5841 | H8 | 1.0889 | 2.1268 | 3.1945 | 3.4823 | 4.4032 | 1.7703 | 1.7551 | | 2.5841 | H9 | 2.2318 | 1.0829 | 1.9762 | 3.2080 | 3.8262 | 3.1043 | 2.5841 | 2.5841 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.013 |
|
C1 |
C2 |
H9 |
119.294 |
| C2 |
C1 |
H6 |
111.042 |
|
C2 |
C1 |
H7 |
109.985 |
| C2 |
C1 |
H8 |
109.985 |
|
C2 |
N3 |
O4 |
111.376 |
| N3 |
C2 |
H9 |
112.693 |
|
N3 |
O4 |
H5 |
101.088 |
| H6 |
C1 |
H7 |
109.180 |
|
H6 |
C1 |
H8 |
109.180 |
| H7 |
C1 |
H8 |
107.388 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
