Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.889884 |
| Energy at 298.15K | |
| HF Energy | -208.889884 |
| Nuclear repulsion energy | 120.180774 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3830 |
3651 |
77.81 |
|
|
|
| 2 |
A' |
3213 |
3063 |
8.92 |
|
|
|
| 3 |
A' |
3197 |
3048 |
6.31 |
|
|
|
| 4 |
A' |
3075 |
2931 |
11.32 |
|
|
|
| 5 |
A' |
1787 |
1703 |
6.85 |
|
|
|
| 6 |
A' |
1507 |
1436 |
19.15 |
|
|
|
| 7 |
A' |
1433 |
1366 |
31.05 |
|
|
|
| 8 |
A' |
1414 |
1348 |
34.78 |
|
|
|
| 9 |
A' |
1366 |
1302 |
35.91 |
|
|
|
| 10 |
A' |
1164 |
1110 |
9.16 |
|
|
|
| 11 |
A' |
992 |
945 |
144.35 |
|
|
|
| 12 |
A' |
940 |
896 |
3.74 |
|
|
|
| 13 |
A' |
688 |
655 |
12.02 |
|
|
|
| 14 |
A' |
312 |
298 |
1.43 |
|
|
|
| 15 |
A" |
3135 |
2987 |
11.09 |
|
|
|
| 16 |
A" |
1512 |
1441 |
11.79 |
|
|
|
| 17 |
A" |
1073 |
1023 |
0.10 |
|
|
|
| 18 |
A" |
871 |
830 |
14.15 |
|
|
|
| 19 |
A" |
512 |
488 |
39.72 |
|
|
|
| 20 |
A" |
396 |
377 |
94.64 |
|
|
|
| 21 |
A" |
17i |
16i |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16199.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 15440.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.444 |
0.484 |
0.000 |
| C2 |
0.000 |
0.858 |
0.000 |
| N3 |
1.014 |
0.084 |
0.000 |
| O4 |
0.656 |
-1.259 |
0.000 |
| H5 |
1.517 |
-1.696 |
0.000 |
| H6 |
-1.573 |
-0.599 |
0.000 |
| H7 |
-1.945 |
0.903 |
0.881 |
| H8 |
-1.945 |
0.903 |
-0.881 |
| H9 |
0.270 |
1.914 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4921 | 2.4904 | 2.7290 | 3.6774 | 1.0903 | 1.0967 | 1.0967 | 2.2321 |
C2 | 1.4921 | | 1.2755 | 2.2162 | 2.9711 | 2.1440 | 2.1358 | 2.1358 | 1.0892 | N3 | 2.4904 | 1.2755 | | 1.3897 | 1.8503 | 2.6752 | 3.1940 | 3.1940 | 1.9749 | O4 | 2.7290 | 2.2162 | 1.3897 | | 0.9659 | 2.3242 | 3.4949 | 3.4949 | 3.1956 | H5 | 3.6774 | 2.9711 | 1.8503 | 0.9659 | | 3.2788 | 4.4180 | 4.4180 | 3.8193 | H6 | 1.0903 | 2.1440 | 2.6752 | 2.3242 | 3.2788 | | 1.7806 | 1.7806 | 3.1156 | H7 | 1.0967 | 2.1358 | 3.1940 | 3.4949 | 4.4180 | 1.7806 | | 1.7623 | 2.5890 | H8 | 1.0967 | 2.1358 | 3.1940 | 3.4949 | 4.4180 | 1.7806 | 1.7623 | | 2.5890 | H9 | 2.2321 | 1.0892 | 1.9749 | 3.1956 | 3.8193 | 3.1156 | 2.5890 | 2.5890 | |
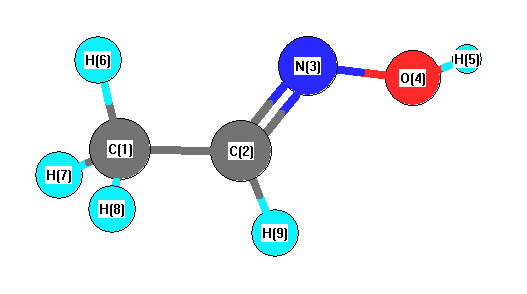 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.104 |
|
C1 |
C2 |
H9 |
118.872 |
| C2 |
C1 |
H6 |
111.291 |
|
C2 |
C1 |
H7 |
110.244 |
| C2 |
C1 |
H8 |
110.244 |
|
C2 |
N3 |
O4 |
112.442 |
| N3 |
C2 |
H9 |
113.025 |
|
N3 |
O4 |
H5 |
102.026 |
| H6 |
C1 |
H7 |
109.012 |
|
H6 |
C1 |
H8 |
109.012 |
| H7 |
C1 |
H8 |
106.923 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.590 |
|
|
|
| 2 |
C |
0.041 |
|
|
|
| 3 |
N |
-0.154 |
|
|
|
| 4 |
O |
-0.527 |
|
|
|
| 5 |
H |
0.434 |
|
|
|
| 6 |
H |
0.224 |
|
|
|
| 7 |
H |
0.195 |
|
|
|
| 8 |
H |
0.195 |
|
|
|
| 9 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.566 |
0.103 |
0.000 |
0.575 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.412 |
-3.937 |
0.000 |
| y |
-3.937 |
-20.708 |
0.000 |
| z |
0.000 |
0.000 |
-24.814 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.349 |
-3.937 |
0.000 |
| y |
-3.937 |
2.905 |
0.000 |
| z |
0.000 |
0.000 |
-3.254 |
|
| Polar |
|---|
| 3z2-r2 | -6.508 |
|---|
| x2-y2 | -1.704 |
|---|
| xy | -3.937 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.509 |
-1.253 |
0.000 |
| y |
-1.253 |
5.706 |
0.000 |
| z |
0.000 |
0.000 |
2.959 |
<r2> (average value of r
2) Å
2
| <r2> |
78.340 |
| (<r2>)1/2 |
8.851 |
