Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G**
Geometric Data calculated at B2PLYP/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.927976 |
| Energy at 298.15K | |
| HF Energy | -208.718107 |
| Nuclear repulsion energy | 119.575031 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3856 |
3856 |
72.40 |
|
|
|
| 2 |
A' |
3224 |
3224 |
7.43 |
|
|
|
| 3 |
A' |
3205 |
3205 |
8.37 |
|
|
|
| 4 |
A' |
3088 |
3088 |
12.77 |
|
|
|
| 5 |
A' |
1741 |
1741 |
5.35 |
|
|
|
| 6 |
A' |
1516 |
1516 |
11.25 |
|
|
|
| 7 |
A' |
1437 |
1437 |
12.68 |
|
|
|
| 8 |
A' |
1390 |
1390 |
21.59 |
|
|
|
| 9 |
A' |
1354 |
1354 |
59.09 |
|
|
|
| 10 |
A' |
1164 |
1164 |
10.53 |
|
|
|
| 11 |
A' |
944 |
944 |
100.16 |
|
|
|
| 12 |
A' |
923 |
923 |
31.42 |
|
|
|
| 13 |
A' |
679 |
679 |
13.24 |
|
|
|
| 14 |
A' |
316 |
316 |
1.44 |
|
|
|
| 15 |
A" |
3147 |
3147 |
14.20 |
|
|
|
| 16 |
A" |
1525 |
1525 |
8.95 |
|
|
|
| 17 |
A" |
1082 |
1082 |
0.33 |
|
|
|
| 18 |
A" |
872 |
872 |
11.30 |
|
|
|
| 19 |
A" |
498 |
498 |
35.61 |
|
|
|
| 20 |
A" |
380 |
380 |
94.70 |
|
|
|
| 21 |
A" |
47i |
47i |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16145.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16145.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.448 |
0.485 |
0.000 |
| C2 |
0.000 |
0.865 |
0.000 |
| N3 |
1.022 |
0.091 |
0.000 |
| O4 |
0.651 |
-1.271 |
0.000 |
| H5 |
1.513 |
-1.705 |
0.000 |
| H6 |
-1.570 |
-0.594 |
0.000 |
| H7 |
-1.945 |
0.903 |
0.879 |
| H8 |
-1.945 |
0.903 |
-0.879 |
| H9 |
0.267 |
1.918 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4967 | 2.5013 | 2.7371 | 3.6828 | 1.0864 | 1.0928 | 1.0928 | 2.2340 |
C2 | 1.4967 | | 1.2821 | 2.2332 | 2.9820 | 2.1436 | 2.1345 | 2.1345 | 1.0862 | N3 | 2.5013 | 1.2821 | | 1.4121 | 1.8619 | 2.6818 | 3.1992 | 3.1992 | 1.9768 | O4 | 2.7371 | 2.2332 | 1.4121 | | 0.9647 | 2.3226 | 3.4983 | 3.4983 | 3.2122 | H5 | 3.6828 | 2.9820 | 1.8619 | 0.9647 | | 3.2774 | 4.4190 | 4.4190 | 3.8311 | H6 | 1.0864 | 2.1436 | 2.6818 | 2.3226 | 3.2774 | | 1.7759 | 1.7759 | 3.1119 | H7 | 1.0928 | 2.1345 | 3.1992 | 3.4983 | 4.4190 | 1.7759 | | 1.7586 | 2.5871 | H8 | 1.0928 | 2.1345 | 3.1992 | 3.4983 | 4.4190 | 1.7759 | 1.7586 | | 2.5871 | H9 | 2.2340 | 1.0862 | 1.9768 | 3.2122 | 3.8311 | 3.1119 | 2.5871 | 2.5871 | |
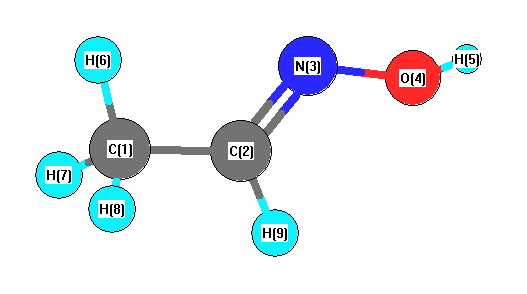 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.193 |
|
C1 |
C2 |
H9 |
118.895 |
| C2 |
C1 |
H6 |
111.174 |
|
C2 |
C1 |
H7 |
110.058 |
| C2 |
C1 |
H8 |
110.058 |
|
C2 |
N3 |
O4 |
111.882 |
| N3 |
C2 |
H9 |
112.912 |
|
N3 |
O4 |
H5 |
101.474 |
| H6 |
C1 |
H7 |
109.155 |
|
H6 |
C1 |
H8 |
109.155 |
| H7 |
C1 |
H8 |
107.146 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.378 |
|
|
|
| 2 |
C |
0.103 |
|
|
|
| 3 |
N |
-0.156 |
|
|
|
| 4 |
O |
-0.444 |
|
|
|
| 5 |
H |
0.339 |
|
|
|
| 6 |
H |
0.157 |
|
|
|
| 7 |
H |
0.129 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.593 |
0.198 |
0.000 |
0.625 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.761 |
-3.816 |
0.000 |
| y |
-3.816 |
-21.014 |
0.000 |
| z |
0.000 |
0.000 |
-24.905 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.198 |
-3.816 |
0.000 |
| y |
-3.816 |
2.819 |
0.000 |
| z |
0.000 |
0.000 |
-3.017 |
|
| Polar |
|---|
| 3z2-r2 | -6.033 |
|---|
| x2-y2 | -1.747 |
|---|
| xy | -3.816 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.479 |
-1.236 |
0.000 |
| y |
-1.236 |
5.778 |
0.000 |
| z |
0.000 |
0.000 |
2.977 |
<r2> (average value of r
2) Å
2
| <r2> |
78.976 |
| (<r2>)1/2 |
8.887 |
