Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.918588 |
| Energy at 298.15K | -208.924546 |
| HF Energy | -208.918588 |
| Nuclear repulsion energy | 120.247593 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3885 |
3707 |
81.72 |
|
|
|
| 2 |
A' |
3208 |
3061 |
6.62 |
|
|
|
| 3 |
A' |
3187 |
3041 |
8.25 |
|
|
|
| 4 |
A' |
3069 |
2929 |
11.45 |
|
|
|
| 5 |
A' |
1779 |
1698 |
7.22 |
|
|
|
| 6 |
A' |
1488 |
1420 |
16.82 |
|
|
|
| 7 |
A' |
1415 |
1350 |
24.07 |
|
|
|
| 8 |
A' |
1389 |
1326 |
37.05 |
|
|
|
| 9 |
A' |
1353 |
1291 |
41.57 |
|
|
|
| 10 |
A' |
1157 |
1104 |
9.71 |
|
|
|
| 11 |
A' |
986 |
941 |
144.99 |
|
|
|
| 12 |
A' |
935 |
893 |
5.09 |
|
|
|
| 13 |
A' |
687 |
655 |
12.38 |
|
|
|
| 14 |
A' |
315 |
300 |
1.43 |
|
|
|
| 15 |
A" |
3132 |
2989 |
11.20 |
|
|
|
| 16 |
A" |
1495 |
1427 |
10.95 |
|
|
|
| 17 |
A" |
1065 |
1016 |
0.16 |
|
|
|
| 18 |
A" |
869 |
829 |
13.52 |
|
|
|
| 19 |
A" |
511 |
487 |
36.68 |
|
|
|
| 20 |
A" |
391 |
373 |
94.48 |
|
|
|
| 21 |
A" |
28 |
27 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16172.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15433.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.443 |
0.483 |
0.000 |
| C2 |
0.000 |
0.859 |
0.000 |
| N3 |
1.014 |
0.085 |
0.000 |
| O4 |
0.655 |
-1.258 |
0.000 |
| H5 |
1.512 |
-1.696 |
0.000 |
| H6 |
-1.568 |
-0.600 |
0.000 |
| H7 |
-1.944 |
0.901 |
0.880 |
| H8 |
-1.944 |
0.901 |
-0.880 |
| H9 |
0.269 |
1.914 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4912 | 2.4883 | 2.7256 | 3.6708 | 1.0894 | 1.0956 | 1.0956 | 2.2318 |
C2 | 1.4912 | | 1.2755 | 2.2163 | 2.9688 | 2.1413 | 2.1340 | 2.1340 | 1.0890 | N3 | 2.4883 | 1.2755 | | 1.3902 | 1.8490 | 2.6702 | 3.1914 | 3.1914 | 1.9751 | O4 | 2.7256 | 2.2163 | 1.3902 | | 0.9625 | 2.3176 | 3.4909 | 3.4909 | 3.1959 | H5 | 3.6708 | 2.9688 | 1.8490 | 0.9625 | | 3.2686 | 4.4110 | 4.4110 | 3.8180 | H6 | 1.0894 | 2.1413 | 2.6702 | 2.3176 | 3.2686 | | 1.7797 | 1.7797 | 3.1135 | H7 | 1.0956 | 2.1340 | 3.1914 | 3.4909 | 4.4110 | 1.7797 | | 1.7602 | 2.5882 | H8 | 1.0956 | 2.1340 | 3.1914 | 3.4909 | 4.4110 | 1.7797 | 1.7602 | | 2.5882 | H9 | 2.2318 | 1.0890 | 1.9751 | 3.1959 | 3.8180 | 3.1135 | 2.5882 | 2.5882 | |
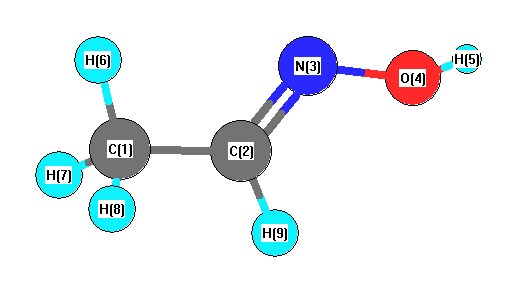 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.988 |
|
C1 |
C2 |
H9 |
118.944 |
| C2 |
C1 |
H6 |
111.204 |
|
C2 |
C1 |
H7 |
110.237 |
| C2 |
C1 |
H8 |
110.237 |
|
C2 |
N3 |
O4 |
112.416 |
| N3 |
C2 |
H9 |
113.068 |
|
N3 |
O4 |
H5 |
102.076 |
| H6 |
C1 |
H7 |
109.080 |
|
H6 |
C1 |
H8 |
109.080 |
| H7 |
C1 |
H8 |
106.893 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.448 |
|
|
|
| 2 |
C |
0.082 |
|
|
|
| 3 |
N |
-0.164 |
|
|
|
| 4 |
O |
-0.439 |
|
|
|
| 5 |
H |
0.347 |
|
|
|
| 6 |
H |
0.177 |
|
|
|
| 7 |
H |
0.153 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
| 9 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.591 |
0.092 |
0.000 |
0.598 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.435 |
-3.898 |
0.000 |
| y |
-3.898 |
-20.699 |
0.000 |
| z |
0.000 |
0.000 |
-24.768 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.298 |
-3.898 |
0.000 |
| y |
-3.898 |
2.902 |
0.000 |
| z |
0.000 |
0.000 |
-3.201 |
|
| Polar |
|---|
| 3z2-r2 | -6.402 |
|---|
| x2-y2 | -1.736 |
|---|
| xy | -3.898 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.576 |
-1.254 |
0.000 |
| y |
-1.254 |
5.774 |
0.000 |
| z |
0.000 |
0.000 |
3.013 |
<r2> (average value of r
2) Å
2
| <r2> |
78.229 |
| (<r2>)1/2 |
8.845 |
