Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
Geometric Data calculated at B2PLYP=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.932761 |
| Energy at 298.15K | |
| HF Energy | -208.718145 |
| Nuclear repulsion energy | 119.609934 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3857 |
3857 |
72.57 |
|
|
|
| 2 |
A' |
3225 |
3225 |
7.28 |
|
|
|
| 3 |
A' |
3206 |
3206 |
8.47 |
|
|
|
| 4 |
A' |
3089 |
3089 |
12.72 |
|
|
|
| 5 |
A' |
1742 |
1742 |
5.35 |
|
|
|
| 6 |
A' |
1516 |
1516 |
11.29 |
|
|
|
| 7 |
A' |
1438 |
1438 |
12.78 |
|
|
|
| 8 |
A' |
1391 |
1391 |
21.42 |
|
|
|
| 9 |
A' |
1354 |
1354 |
59.31 |
|
|
|
| 10 |
A' |
1165 |
1165 |
10.54 |
|
|
|
| 11 |
A' |
944 |
944 |
100.62 |
|
|
|
| 12 |
A' |
923 |
923 |
31.14 |
|
|
|
| 13 |
A' |
679 |
679 |
13.23 |
|
|
|
| 14 |
A' |
316 |
316 |
1.44 |
|
|
|
| 15 |
A" |
3149 |
3149 |
14.11 |
|
|
|
| 16 |
A" |
1526 |
1526 |
8.99 |
|
|
|
| 17 |
A" |
1083 |
1083 |
0.32 |
|
|
|
| 18 |
A" |
872 |
872 |
11.29 |
|
|
|
| 19 |
A" |
499 |
499 |
35.64 |
|
|
|
| 20 |
A" |
380 |
380 |
94.75 |
|
|
|
| 21 |
A" |
46i |
46i |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16152.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16152.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.447 |
0.485 |
0.000 |
| C2 |
0.000 |
0.865 |
0.000 |
| N3 |
1.022 |
0.091 |
0.000 |
| O4 |
0.651 |
-1.271 |
0.000 |
| H5 |
1.513 |
-1.704 |
0.000 |
| H6 |
-1.570 |
-0.594 |
0.000 |
| H7 |
-1.944 |
0.903 |
0.879 |
| H8 |
-1.944 |
0.903 |
-0.879 |
| H9 |
0.266 |
1.917 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4962 | 2.5005 | 2.7364 | 3.6819 | 1.0860 | 1.0925 | 1.0925 | 2.2333 |
C2 | 1.4962 | | 1.2817 | 2.2327 | 2.9814 | 2.1429 | 2.1339 | 2.1339 | 1.0859 | N3 | 2.5005 | 1.2817 | | 1.4117 | 1.8615 | 2.6809 | 3.1981 | 3.1981 | 1.9762 | O4 | 2.7364 | 2.2327 | 1.4117 | | 0.9645 | 2.3221 | 3.4974 | 3.4974 | 3.2114 | H5 | 3.6819 | 2.9814 | 1.8615 | 0.9645 | | 3.2767 | 4.4180 | 4.4180 | 3.8302 | H6 | 1.0860 | 2.1429 | 2.6809 | 2.3221 | 3.2767 | | 1.7753 | 1.7753 | 3.1110 | H7 | 1.0925 | 2.1339 | 3.1981 | 3.4974 | 4.4180 | 1.7753 | | 1.7580 | 2.5863 | H8 | 1.0925 | 2.1339 | 3.1981 | 3.4974 | 4.4180 | 1.7753 | 1.7580 | | 2.5863 | H9 | 2.2333 | 1.0859 | 1.9762 | 3.2114 | 3.8302 | 3.1110 | 2.5863 | 2.5863 | |
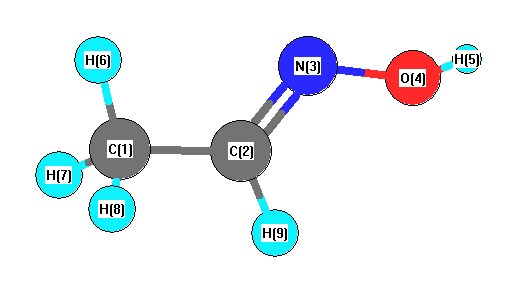 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.190 |
|
C1 |
C2 |
H9 |
118.894 |
| C2 |
C1 |
H6 |
111.178 |
|
C2 |
C1 |
H7 |
110.061 |
| C2 |
C1 |
H8 |
110.061 |
|
C2 |
N3 |
O4 |
111.895 |
| N3 |
C2 |
H9 |
112.915 |
|
N3 |
O4 |
H5 |
101.488 |
| H6 |
C1 |
H7 |
109.152 |
|
H6 |
C1 |
H8 |
109.152 |
| H7 |
C1 |
H8 |
107.142 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
