Jump to
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.555016 |
| Energy at 298.15K | -208.561044 |
| HF Energy | -207.894800 |
| Nuclear repulsion energy | 117.170973 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3923 |
3679 |
63.83 |
|
|
|
| 2 |
A |
3232 |
3031 |
15.34 |
|
|
|
| 3 |
A |
3208 |
3008 |
7.81 |
|
|
|
| 4 |
A |
3120 |
2925 |
17.33 |
|
|
|
| 5 |
A |
1789 |
1677 |
1.11 |
|
|
|
| 6 |
A |
1549 |
1452 |
5.39 |
|
|
|
| 7 |
A |
1495 |
1402 |
5.62 |
|
|
|
| 8 |
A |
1453 |
1362 |
23.09 |
|
|
|
| 9 |
A |
1328 |
1245 |
63.59 |
|
|
|
| 10 |
A |
1189 |
1114 |
4.47 |
|
|
|
| 11 |
A |
1055 |
990 |
97.98 |
|
|
|
| 12 |
A |
943 |
884 |
7.66 |
|
|
|
| 13 |
A |
575 |
539 |
15.02 |
|
|
|
| 14 |
A |
330 |
310 |
1.55 |
|
|
|
| 15 |
A |
3192 |
2993 |
18.49 |
|
|
|
| 16 |
A |
1541 |
1445 |
6.58 |
|
|
|
| 17 |
A |
1116 |
1047 |
1.28 |
|
|
|
| 18 |
A |
930 |
872 |
10.00 |
|
|
|
| 19 |
A |
388 |
364 |
144.61 |
|
|
|
| 20 |
A |
288 |
270 |
3.76 |
|
|
|
| 21 |
A |
194 |
182 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16419.3 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 15394.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.301 |
1.303 |
0.000 |
| C2 |
0.000 |
0.563 |
0.000 |
| N3 |
-0.001 |
-0.715 |
0.000 |
| O4 |
1.321 |
-1.203 |
0.000 |
| H5 |
1.190 |
-2.156 |
0.000 |
| H6 |
-2.129 |
0.598 |
0.000 |
| H7 |
-1.379 |
1.943 |
0.880 |
| H8 |
-1.379 |
1.943 |
-0.880 |
| H9 |
0.939 |
1.111 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4967 | 2.4007 | 3.6270 | 4.2625 | 1.0876 | 1.0908 | 1.0908 | 2.2482 |
C2 | 1.4967 | | 1.2782 | 2.2054 | 2.9679 | 2.1296 | 2.1403 | 2.1403 | 1.0875 | N3 | 2.4007 | 1.2782 | | 1.4092 | 1.8691 | 2.5011 | 3.1208 | 3.1208 | 2.0542 | O4 | 3.6270 | 2.2054 | 1.4092 | | 0.9622 | 3.8924 | 4.2382 | 4.2382 | 2.3455 | H5 | 4.2625 | 2.9679 | 1.8691 | 0.9622 | | 4.3131 | 4.9168 | 4.9168 | 3.2769 | H6 | 1.0876 | 2.1296 | 2.5011 | 3.8924 | 4.3131 | | 1.7732 | 1.7732 | 3.1111 | H7 | 1.0908 | 2.1403 | 3.1208 | 4.2382 | 4.9168 | 1.7732 | | 1.7601 | 2.6156 | H8 | 1.0908 | 2.1403 | 3.1208 | 4.2382 | 4.9168 | 1.7732 | 1.7601 | | 2.6156 | H9 | 2.2482 | 1.0875 | 2.0542 | 2.3455 | 3.2769 | 3.1111 | 2.6156 | 2.6156 | |
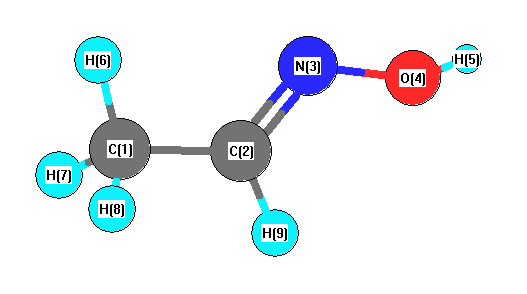 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.595 |
|
C1 |
C2 |
H9 |
120.088 |
| C2 |
C1 |
H6 |
109.979 |
|
C2 |
C1 |
H7 |
110.640 |
| C2 |
C1 |
H8 |
110.640 |
|
C2 |
N3 |
O4 |
110.207 |
| N3 |
C2 |
H9 |
120.317 |
|
N3 |
O4 |
H5 |
102.391 |
| H6 |
C1 |
H7 |
108.979 |
|
H6 |
C1 |
H8 |
108.979 |
| H7 |
C1 |
H8 |
107.564 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.554188 |
| Energy at 298.15K | |
| HF Energy | -207.893544 |
| Nuclear repulsion energy | 119.715304 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3928 |
3682 |
65.95 |
|
|
|
| 2 |
A' |
3266 |
3062 |
7.87 |
|
|
|
| 3 |
A' |
3245 |
3043 |
10.46 |
|
|
|
| 4 |
A' |
3126 |
2931 |
13.76 |
|
|
|
| 5 |
A' |
1800 |
1687 |
3.02 |
|
|
|
| 6 |
A' |
1545 |
1448 |
9.20 |
|
|
|
| 7 |
A' |
1468 |
1377 |
11.33 |
|
|
|
| 8 |
A' |
1420 |
1332 |
31.57 |
|
|
|
| 9 |
A' |
1384 |
1298 |
52.30 |
|
|
|
| 10 |
A' |
1185 |
1111 |
9.85 |
|
|
|
| 11 |
A' |
978 |
917 |
97.14 |
|
|
|
| 12 |
A' |
944 |
885 |
1.65 |
|
|
|
| 13 |
A' |
684 |
641 |
13.27 |
|
|
|
| 14 |
A' |
324 |
304 |
1.69 |
|
|
|
| 15 |
A" |
3193 |
2994 |
16.24 |
|
|
|
| 16 |
A" |
1551 |
1454 |
8.02 |
|
|
|
| 17 |
A" |
1106 |
1037 |
0.91 |
|
|
|
| 18 |
A" |
888 |
833 |
9.11 |
|
|
|
| 19 |
A" |
491 |
461 |
32.27 |
|
|
|
| 20 |
A" |
377 |
354 |
105.34 |
|
|
|
| 21 |
A" |
71i |
66i |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16416.3 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 15391.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.449 |
0.479 |
0.000 |
| C2 |
0.000 |
0.864 |
0.000 |
| N3 |
1.024 |
0.094 |
0.000 |
| O4 |
0.652 |
-1.265 |
0.000 |
| H5 |
1.508 |
-1.704 |
0.000 |
| H6 |
-1.571 |
-0.599 |
0.000 |
| H7 |
-1.943 |
0.895 |
0.879 |
| H8 |
-1.943 |
0.895 |
-0.879 |
| H9 |
0.258 |
1.917 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4994 | 2.5027 | 2.7311 | 3.6761 | 1.0844 | 1.0908 | 1.0908 | 2.2319 |
C2 | 1.4994 | | 1.2806 | 2.2266 | 2.9783 | 2.1462 | 2.1326 | 2.1326 | 1.0842 | N3 | 2.5027 | 1.2806 | | 1.4092 | 1.8628 | 2.6855 | 3.1958 | 3.1958 | 1.9768 | O4 | 2.7311 | 2.2266 | 1.4092 | | 0.9619 | 2.3211 | 3.4891 | 3.4891 | 3.2063 | H5 | 3.6761 | 2.9783 | 1.8628 | 0.9619 | | 3.2717 | 4.4089 | 4.4089 | 3.8309 | H6 | 1.0844 | 2.1462 | 2.6855 | 2.3211 | 3.2717 | | 1.7727 | 1.7727 | 3.1099 | H7 | 1.0908 | 2.1326 | 3.1958 | 3.4891 | 4.4089 | 1.7727 | | 1.7587 | 2.5806 | H8 | 1.0908 | 2.1326 | 3.1958 | 3.4891 | 4.4089 | 1.7727 | 1.7587 | | 2.5806 | H9 | 2.2319 | 1.0842 | 1.9768 | 3.2063 | 3.8309 | 3.1099 | 2.5806 | 2.5806 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.202 |
|
C1 |
C2 |
H9 |
118.630 |
| C2 |
C1 |
H6 |
111.312 |
|
C2 |
C1 |
H7 |
109.833 |
| C2 |
C1 |
H8 |
109.833 |
|
C2 |
N3 |
O4 |
111.655 |
| N3 |
C2 |
H9 |
113.168 |
|
N3 |
O4 |
H5 |
101.895 |
| H6 |
C1 |
H7 |
109.164 |
|
H6 |
C1 |
H8 |
109.164 |
| H7 |
C1 |
H8 |
107.449 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
