Jump to
S1C2
Vibrational Frequencies calculated at MP3=FULL/6-31G**
Geometric Data calculated at MP3=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -208.566141 |
| Energy at 298.15K | |
| HF Energy | -207.894033 |
| Nuclear repulsion energy | 119.990164 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3963 |
3963 |
70.63 |
|
|
|
| 2 |
A' |
3287 |
3287 |
7.43 |
|
|
|
| 3 |
A' |
3268 |
3268 |
9.78 |
|
|
|
| 4 |
A' |
3148 |
3148 |
12.37 |
|
|
|
| 5 |
A' |
1812 |
1812 |
3.76 |
|
|
|
| 6 |
A' |
1550 |
1550 |
10.18 |
|
|
|
| 7 |
A' |
1472 |
1472 |
12.72 |
|
|
|
| 8 |
A' |
1426 |
1426 |
32.94 |
|
|
|
| 9 |
A' |
1391 |
1391 |
51.94 |
|
|
|
| 10 |
A' |
1190 |
1190 |
10.14 |
|
|
|
| 11 |
A' |
986 |
986 |
105.34 |
|
|
|
| 12 |
A' |
951 |
951 |
0.29 |
|
|
|
| 13 |
A' |
689 |
689 |
13.35 |
|
|
|
| 14 |
A' |
327 |
327 |
1.68 |
|
|
|
| 15 |
A" |
3216 |
3216 |
14.70 |
|
|
|
| 16 |
A" |
1556 |
1556 |
8.53 |
|
|
|
| 17 |
A" |
1109 |
1109 |
0.86 |
|
|
|
| 18 |
A" |
893 |
893 |
9.51 |
|
|
|
| 19 |
A" |
495 |
495 |
32.14 |
|
|
|
| 20 |
A" |
380 |
380 |
106.53 |
|
|
|
| 21 |
A" |
69i |
69i |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16518.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16518.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.446 |
0.476 |
0.000 |
| C2 |
0.000 |
0.863 |
0.000 |
| N3 |
1.021 |
0.094 |
0.000 |
| O4 |
0.651 |
-1.262 |
0.000 |
| H5 |
1.504 |
-1.701 |
0.000 |
| H6 |
-1.564 |
-0.599 |
0.000 |
| H7 |
-1.939 |
0.891 |
0.877 |
| H8 |
-1.939 |
0.891 |
-0.877 |
| H9 |
0.257 |
1.914 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4967 | 2.4963 | 2.7237 | 3.6666 | 1.0822 | 1.0883 | 1.0883 | 2.2283 |
C2 | 1.4967 | | 1.2779 | 2.2222 | 2.9727 | 2.1410 | 2.1284 | 2.1284 | 1.0817 | N3 | 2.4963 | 1.2779 | | 1.4056 | 1.8594 | 2.6764 | 3.1883 | 3.1883 | 1.9732 | O4 | 2.7237 | 2.2222 | 1.4056 | | 0.9594 | 2.3120 | 3.4801 | 3.4801 | 3.1998 | H5 | 3.6666 | 2.9727 | 1.8594 | 0.9594 | | 3.2598 | 4.3979 | 4.3979 | 3.8240 | H6 | 1.0822 | 2.1410 | 2.6764 | 2.3120 | 3.2598 | | 1.7694 | 1.7694 | 3.1034 | H7 | 1.0883 | 2.1284 | 3.1883 | 3.4801 | 4.3979 | 1.7694 | | 1.7546 | 2.5764 | H8 | 1.0883 | 2.1284 | 3.1883 | 3.4801 | 4.3979 | 1.7694 | 1.7546 | | 2.5764 | H9 | 2.2283 | 1.0817 | 1.9732 | 3.1998 | 3.8240 | 3.1034 | 2.5764 | 2.5764 | |
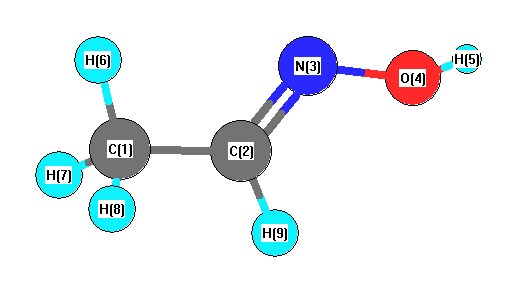 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.066 |
|
C1 |
C2 |
H9 |
118.706 |
| C2 |
C1 |
H6 |
111.227 |
|
C2 |
C1 |
H7 |
109.837 |
| C2 |
C1 |
H8 |
109.837 |
|
C2 |
N3 |
O4 |
111.712 |
| N3 |
C2 |
H9 |
113.228 |
|
N3 |
O4 |
H5 |
102.008 |
| H6 |
C1 |
H7 |
109.211 |
|
H6 |
C1 |
H8 |
109.211 |
| H7 |
C1 |
H8 |
107.435 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
