Jump to
S1C2
Vibrational Frequencies calculated at TPSSh/6-31G**
Geometric Data calculated at TPSSh/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -209.146870 |
| Energy at 298.15K | -209.152748 |
| HF Energy | -209.146870 |
| Nuclear repulsion energy | 119.430722 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3794 |
3637 |
59.79 |
|
|
|
| 2 |
A' |
3182 |
3051 |
13.45 |
|
|
|
| 3 |
A' |
3166 |
3035 |
7.21 |
|
|
|
| 4 |
A' |
3045 |
2919 |
16.54 |
|
|
|
| 5 |
A' |
1731 |
1660 |
7.02 |
|
|
|
| 6 |
A' |
1503 |
1441 |
11.26 |
|
|
|
| 7 |
A' |
1419 |
1361 |
12.20 |
|
|
|
| 8 |
A' |
1376 |
1319 |
25.19 |
|
|
|
| 9 |
A' |
1343 |
1288 |
55.31 |
|
|
|
| 10 |
A' |
1149 |
1102 |
10.24 |
|
|
|
| 11 |
A' |
932 |
893 |
125.26 |
|
|
|
| 12 |
A' |
916 |
878 |
18.04 |
|
|
|
| 13 |
A' |
671 |
643 |
12.76 |
|
|
|
| 14 |
A' |
305 |
292 |
1.30 |
|
|
|
| 15 |
A" |
3098 |
2970 |
17.88 |
|
|
|
| 16 |
A" |
1512 |
1450 |
8.42 |
|
|
|
| 17 |
A" |
1063 |
1019 |
0.48 |
|
|
|
| 18 |
A" |
860 |
824 |
13.78 |
|
|
|
| 19 |
A" |
503 |
482 |
42.53 |
|
|
|
| 20 |
A" |
389 |
373 |
82.32 |
|
|
|
| 21 |
A" |
12 |
12 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15984.0 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 15323.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.450 |
0.485 |
0.000 |
| C2 |
0.000 |
0.866 |
0.000 |
| N3 |
1.023 |
0.092 |
0.000 |
| O4 |
0.652 |
-1.271 |
0.000 |
| H5 |
1.519 |
-1.704 |
0.000 |
| H6 |
-1.570 |
-0.598 |
0.000 |
| H7 |
-1.950 |
0.903 |
0.881 |
| H8 |
-1.950 |
0.903 |
-0.881 |
| H9 |
0.268 |
1.921 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4989 | 2.5040 | 2.7393 | 3.6887 | 1.0895 | 1.0962 | 1.0962 | 2.2391 |
C2 | 1.4989 | | 1.2829 | 2.2343 | 2.9853 | 2.1465 | 2.1400 | 2.1400 | 1.0889 | N3 | 2.5040 | 1.2829 | | 1.4125 | 1.8632 | 2.6836 | 3.2052 | 3.2052 | 1.9788 | O4 | 2.7393 | 2.2343 | 1.4125 | | 0.9686 | 2.3226 | 3.5039 | 3.5039 | 3.2152 | H5 | 3.6887 | 2.9853 | 1.8632 | 0.9686 | | 3.2814 | 4.4281 | 4.4281 | 3.8349 | H6 | 1.0895 | 2.1465 | 2.6836 | 2.3226 | 3.2814 | | 1.7816 | 1.7816 | 3.1184 | H7 | 1.0962 | 2.1400 | 3.2052 | 3.5039 | 4.4281 | 1.7816 | | 1.7628 | 2.5946 | H8 | 1.0962 | 2.1400 | 3.2052 | 3.5039 | 4.4281 | 1.7816 | 1.7628 | | 2.5946 | H9 | 2.2391 | 1.0889 | 1.9788 | 3.2152 | 3.8349 | 3.1184 | 2.5946 | 2.5946 | |
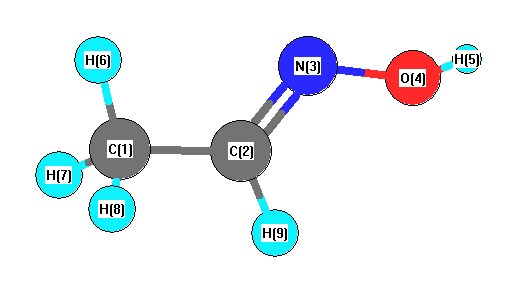 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.188 |
|
C1 |
C2 |
H9 |
118.978 |
| C2 |
C1 |
H6 |
111.062 |
|
C2 |
C1 |
H7 |
110.133 |
| C2 |
C1 |
H8 |
110.133 |
|
C2 |
N3 |
O4 |
111.887 |
| N3 |
C2 |
H9 |
112.834 |
|
N3 |
O4 |
H5 |
101.342 |
| H6 |
C1 |
H7 |
109.196 |
|
H6 |
C1 |
H8 |
109.196 |
| H7 |
C1 |
H8 |
107.027 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.417 |
|
|
|
| 2 |
C |
0.093 |
|
|
|
| 3 |
N |
-0.176 |
|
|
|
| 4 |
O |
-0.435 |
|
|
|
| 5 |
H |
0.344 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.143 |
|
|
|
| 8 |
H |
0.143 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.608 |
0.146 |
0.000 |
0.625 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.688 |
-1.284 |
0.000 |
| y |
-1.284 |
5.928 |
0.000 |
| z |
0.000 |
0.000 |
3.031 |
<r2> (average value of r
2) Å
2
| <r2> |
78.983 |
| (<r2>)1/2 |
8.887 |
