return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10O (cyclohexanone)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-308.999138
Energy at 298.15K-309.011435
HF Energy-307.919387
Nuclear repulsion energy309.383985
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3184 2998 18.48      
2 A' 3155 2970 48.01      
3 A' 3147 2962 41.75      
4 A' 3099 2918 44.57      
5 A' 3093 2912 7.90      
6 A' 3089 2908 19.68      
7 A' 1823 1717 147.49      
8 A' 1563 1471 0.51      
9 A' 1548 1458 7.46      
10 A' 1529 1440 5.93      
11 A' 1426 1343 5.70      
12 A' 1394 1312 0.55      
13 A' 1310 1233 0.47      
14 A' 1283 1207 0.55      
15 A' 1168 1099 0.51      
16 A' 1058 996 1.75      
17 A' 1030 970 0.57      
18 A' 895 842 2.02      
19 A' 868 818 0.09      
20 A' 773 728 1.81      
21 A' 667 628 1.29      
22 A' 486 457 0.48      
23 A' 409 385 0.96      
24 A' 312 294 0.25      
25 A' 94 89 3.09      
26 A" 3183 2996 17.57      
27 A" 3150 2965 35.14      
28 A" 3097 2916 17.72      
29 A" 3088 2907 4.76      
30 A" 1548 1457 3.37      
31 A" 1522 1433 5.17      
32 A" 1432 1348 0.99      
33 A" 1410 1327 11.06      
34 A" 1394 1312 10.73      
35 A" 1324 1246 0.02      
36 A" 1276 1201 26.32      
37 A" 1184 1115 14.05      
38 A" 1114 1049 1.30      
39 A" 1093 1029 4.50      
40 A" 942 887 4.44      
41 A" 930 876 1.88      
42 A" 787 741 2.86      
43 A" 496 467 11.54      
44 A" 418 393 2.25      
45 A" 189 178 0.73      

Unscaled Zero Point Vibrational Energy (zpe) 33990.4 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 31998.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.13995 0.08314 0.05828

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.419 -1.074 0.000
C2 -0.165 -0.504 1.284
C3 -0.165 -0.504 -1.284
C4 -0.165 1.034 1.263
C5 -0.165 1.034 -1.263
C6 -0.841 1.576 0.000
O7 1.296 -1.926 0.000
H8 0.400 -0.903 2.126
H9 -1.198 -0.861 1.367
H10 0.400 -0.903 -2.126
H11 -1.198 -0.861 -1.367
H12 0.870 1.391 1.299
H13 -0.661 1.414 2.159
H14 0.870 1.391 -1.299
H15 -0.661 1.414 -2.159
H16 -0.812 2.668 0.000
H17 -1.897 1.286 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.52061.52062.52512.52512.93371.22292.13262.12752.13262.12752.82233.46592.82233.46593.93923.3061
C21.52062.56721.53792.97482.53612.40871.08971.09623.47882.86722.15922.16573.36603.97173.48302.8024
C31.52062.56722.97481.53792.53612.40873.47882.86721.08971.09623.36603.97172.15922.16573.48302.8024
C42.52511.53792.97482.52581.53173.53352.19462.16093.94393.40231.09481.09222.78553.47822.16482.1588
C52.52512.97481.53792.52581.53173.53353.94393.40232.19462.16092.78553.47821.09481.09222.16482.1588
C62.93372.53612.53611.53171.53174.10213.49372.81743.49372.81742.15532.17232.15532.17231.09261.0957
O71.22292.40872.40873.53353.53354.10212.52313.03632.52313.03633.58754.43173.58754.43175.05464.5288
H82.13261.08973.47882.19463.94393.49372.52311.76914.25143.84112.48402.54874.14884.98514.32973.8196
H92.12751.09622.86722.16093.40232.81743.03631.76913.84112.73423.05822.46824.05634.23053.80482.6400
H102.13263.47881.08973.94392.19463.49372.52314.25143.84111.76914.14884.98512.48402.54874.32973.8196
H112.12752.86721.09623.40232.16092.81743.03633.84112.73421.76914.05634.23053.05822.46823.80482.6400
H122.82232.15923.36601.09482.78552.15533.58752.48403.05824.14884.05631.75542.59733.78092.47923.0582
H133.46592.16573.97171.09223.47822.17234.43172.54872.46824.98514.23051.75543.78094.31752.50142.4912
H142.82233.36602.15922.78551.09482.15533.58754.14884.05632.48403.05822.59733.78091.75542.47923.0582
H153.46593.97172.16573.47821.09222.17234.43174.98514.23052.54872.46823.78094.31751.75542.50142.4912
H163.93923.48303.48302.16482.16481.09265.05464.32973.80484.32973.80482.47922.50142.47922.50141.7572
H173.30612.80242.80242.15882.15881.09574.52883.81962.64003.81962.64003.05822.49123.05822.49121.7572

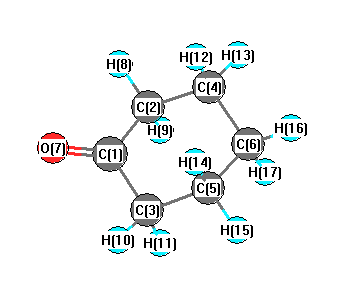
picture of cyclohexanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 111.294 C1 C2 H8 108.437
C1 C2 H9 107.676 C1 C3 C5 111.294
C1 C3 H10 108.437 C1 C3 H11 107.676
C2 C1 C3 115.159 C2 C1 O7 122.420
C2 C4 C6 111.417 C2 C4 H12 109.031
C2 C4 H13 109.692 C3 C1 O7 122.420
C3 C5 C6 111.417 C3 C5 H14 109.031
C3 C5 H15 109.692 C4 C2 H8 112.141
C4 C2 H9 109.088 C4 C6 C5 111.079
C4 C6 H16 110.034 C4 C6 H17 109.378
C5 C3 H10 112.141 C5 C3 H11 109.088
C5 C6 H16 110.034 C5 C6 H17 109.378
C6 C4 H12 109.160 C6 C4 H13 110.648
C6 C5 H14 109.160 C6 C5 H15 110.648
H8 C2 H9 108.054 H10 C3 H11 108.054
H12 C4 H13 106.768 H14 C5 H15 106.768
H16 C6 H17 106.838
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability