return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12 (Pentane)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-197.115466
Energy at 298.15K-197.128459
HF Energy-196.351483
Nuclear repulsion energy185.526811
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3223 3011 39.28      
2 A1 3126 2921 23.39      
3 A1 3122 2917 63.36      
4 A1 3102 2899 7.99      
5 A1 1572 1469 4.19      
6 A1 1558 1455 0.01      
7 A1 1544 1443 0.01      
8 A1 1469 1373 2.52      
9 A1 1411 1319 0.11      
10 A1 1204 1125 0.73      
11 A1 1094 1023 0.61      
12 A1 906 847 0.88      
13 A1 409 382 0.00      
14 A1 182 170 0.01      
15 A2 3221 3010 0.00      
16 A2 3168 2960 0.00      
17 A2 1563 1461 0.00      
18 A2 1372 1282 0.00      
19 A2 1312 1226 0.00      
20 A2 1022 955 0.00      
21 A2 786 734 0.00      
22 A2 268 250 0.00      
23 A2 111 103 0.00      
24 B1 3222 3011 86.43      
25 B1 3181 2972 53.20      
26 B1 3149 2942 0.98      
27 B1 1562 1460 11.12      
28 B1 1369 1279 0.80      
29 B1 1255 1173 0.03      
30 B1 892 834 0.96      
31 B1 749 699 3.09      
32 B1 258 241 0.00      
33 B1 118 111 0.00      
34 B2 3223 3011 24.33      
35 B2 3126 2920 37.46      
36 B2 3116 2912 0.00      
37 B2 1568 1465 1.28      
38 B2 1547 1446 0.47      
39 B2 1465 1369 1.02      
40 B2 1456 1361 1.11      
41 B2 1325 1238 3.55      
42 B2 1124 1051 0.38      
43 B2 1080 1009 1.82      
44 B2 964 901 2.28      
45 B2 406 379 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 36449.3 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 34058.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.57267 0.06523 0.06189

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.313
C2 0.000 1.274 -0.523
C3 0.000 -1.274 -0.523
C4 0.000 2.539 0.325
C5 0.000 -2.539 0.325
H6 0.875 0.000 0.969
H7 -0.875 0.000 0.969
H8 0.874 1.272 -1.178
H9 -0.874 1.272 -1.178
H10 -0.874 -1.272 -1.178
H11 0.874 -1.272 -1.178
H12 0.000 3.435 -0.293
H13 0.000 -3.435 -0.293
H14 -0.880 2.575 0.966
H15 0.880 2.575 0.966
H16 0.880 -2.575 0.966
H17 -0.880 -2.575 0.966

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.52371.52372.53912.53911.09351.09352.14552.14552.14552.14553.48793.48792.79812.79812.79812.7981
C21.52372.54781.52313.90612.14812.14811.09211.09212.77002.77002.17324.71442.16422.16424.21944.2194
C31.52372.54783.90611.52312.14812.14812.77002.77001.09211.09214.71442.17324.21944.21942.16422.1642
C42.53911.52313.90615.07812.76172.76172.15142.15144.18864.18861.08856.00581.08901.08905.22835.2283
C52.53913.90611.52315.07812.76172.76174.18864.18862.15142.15146.00581.08855.22835.22831.08901.0890
H61.09352.14812.14812.76172.76171.74982.49503.04703.04702.49503.76253.76253.11562.57472.57473.1156
H71.09352.14812.14812.76172.76171.74983.04702.49502.49503.04703.76253.76252.57473.11563.11562.5747
H82.14551.09212.77002.15144.18862.49503.04701.74833.08632.54342.49524.86813.06102.50884.40354.7399
H92.14551.09212.77002.15144.18863.04702.49501.74832.54343.08632.49524.86812.50883.06104.73994.4035
H102.14552.77001.09214.18862.15143.04702.49503.08632.54341.74834.86812.49524.40354.73993.06102.5088
H112.14552.77001.09214.18862.15142.49503.04702.54343.08631.74834.86812.49524.73994.40352.50883.0610
H123.48792.17324.71441.08856.00583.76253.76252.49522.49524.86814.86816.86971.76061.76066.20286.2028
H133.48794.71442.17326.00581.08853.76253.76254.86814.86812.49522.49526.86976.20286.20281.76061.7606
H142.79812.16424.21941.08905.22833.11562.57473.06102.50884.40354.73991.76066.20281.75925.44155.1494
H152.79812.16424.21941.08905.22832.57473.11562.50883.06104.73994.40351.76066.20281.75925.14945.4415
H162.79814.21942.16425.22831.08902.57473.11564.40354.73993.06102.50886.20281.76065.44155.14941.7592
H172.79814.21942.16425.22831.08903.11562.57474.73994.40352.50883.06106.20281.76065.14945.44151.7592

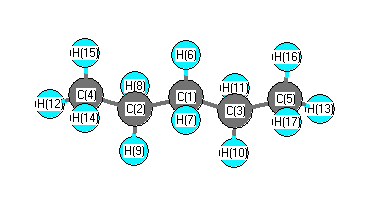
picture of Pentane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.888 C1 C2 H8 109.097
C1 C2 H9 109.097 C1 C3 C5 112.888
C1 C3 H10 109.097 C1 C3 H11 109.097
C2 C1 C3 113.452 C2 C1 H6 109.217
C2 C1 H7 109.217 C2 C4 H12 111.549
C2 C4 H14 110.794 C2 C4 H15 110.794
C3 C1 H6 109.217 C3 C1 H7 109.217
C3 C5 H13 111.549 C3 C5 H16 110.794
C3 C5 H17 110.794 C4 C2 H8 109.601
C4 C2 H9 109.601 C5 C3 H10 109.601
C5 C3 H11 109.601 H6 C1 H7 106.270
H8 C2 H9 106.343 H10 C3 H11 106.343
H12 C4 H14 107.907 H12 C4 H15 107.907
H13 C5 H16 107.907 H13 C5 H17 107.907
H14 C4 H15 107.739 H16 C5 H17 107.739
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability