Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -196.310495 |
| Energy at 298.15K | -196.320764 |
| HF Energy | -196.310495 |
| Nuclear repulsion energy | 173.602274 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3260 |
3111 |
16.83 |
|
|
|
| 2 |
A |
3171 |
3026 |
4.15 |
|
|
|
| 3 |
A |
3153 |
3009 |
29.79 |
|
|
|
| 4 |
A |
3153 |
3008 |
30.50 |
|
|
|
| 5 |
A |
3141 |
2997 |
34.63 |
|
|
|
| 6 |
A |
3101 |
2959 |
27.16 |
|
|
|
| 7 |
A |
3091 |
2950 |
22.47 |
|
|
|
| 8 |
A |
3062 |
2922 |
22.14 |
|
|
|
| 9 |
A |
3053 |
2914 |
29.54 |
|
|
|
| 10 |
A |
3041 |
2902 |
25.61 |
|
|
|
| 11 |
A |
1747 |
1667 |
8.27 |
|
|
|
| 12 |
A |
1517 |
1448 |
3.21 |
|
|
|
| 13 |
A |
1509 |
1440 |
9.51 |
|
|
|
| 14 |
A |
1499 |
1430 |
0.45 |
|
|
|
| 15 |
A |
1487 |
1420 |
6.80 |
|
|
|
| 16 |
A |
1459 |
1393 |
1.14 |
|
|
|
| 17 |
A |
1425 |
1360 |
7.24 |
|
|
|
| 18 |
A |
1380 |
1317 |
2.34 |
|
|
|
| 19 |
A |
1362 |
1300 |
1.18 |
|
|
|
| 20 |
A |
1327 |
1266 |
1.25 |
|
|
|
| 21 |
A |
1293 |
1234 |
0.18 |
|
|
|
| 22 |
A |
1264 |
1206 |
2.07 |
|
|
|
| 23 |
A |
1199 |
1145 |
0.43 |
|
|
|
| 24 |
A |
1109 |
1058 |
2.28 |
|
|
|
| 25 |
A |
1088 |
1038 |
5.43 |
|
|
|
| 26 |
A |
1041 |
994 |
10.17 |
|
|
|
| 27 |
A |
1018 |
971 |
1.81 |
|
|
|
| 28 |
A |
947 |
904 |
4.43 |
|
|
|
| 29 |
A |
944 |
901 |
39.22 |
|
|
|
| 30 |
A |
889 |
849 |
0.50 |
|
|
|
| 31 |
A |
879 |
839 |
2.12 |
|
|
|
| 32 |
A |
778 |
743 |
4.55 |
|
|
|
| 33 |
A |
650 |
620 |
7.95 |
|
|
|
| 34 |
A |
435 |
415 |
0.42 |
|
|
|
| 35 |
A |
391 |
373 |
1.82 |
|
|
|
| 36 |
A |
301 |
287 |
0.20 |
|
|
|
| 37 |
A |
225 |
214 |
0.15 |
|
|
|
| 38 |
A |
125 |
119 |
0.02 |
|
|
|
| 39 |
A |
91 |
87 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30302.7 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 28917.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.090 |
0.570 |
0.118 |
| C2 |
-1.179 |
-0.253 |
-0.397 |
| C3 |
-0.065 |
-0.901 |
0.365 |
| C4 |
1.325 |
-0.517 |
-0.159 |
| C5 |
1.649 |
0.960 |
0.022 |
| H6 |
-2.875 |
1.009 |
-0.491 |
| H7 |
-2.086 |
0.830 |
1.174 |
| H8 |
-1.220 |
-0.483 |
-1.463 |
| H9 |
-0.175 |
-1.992 |
0.302 |
| H10 |
-0.145 |
-0.638 |
1.428 |
| H11 |
2.078 |
-1.127 |
0.355 |
| H12 |
1.393 |
-0.783 |
-1.223 |
| H13 |
2.639 |
1.203 |
-0.374 |
| H14 |
0.914 |
1.591 |
-0.488 |
| H15 |
1.636 |
1.236 |
1.083 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3310 | 2.5150 | 3.5946 | 3.7603 | 1.0859 | 1.0877 | 2.0900 | 3.2041 | 2.6377 | 4.5061 | 3.9696 | 4.7968 | 3.2305 | 3.9064 |
C2 | 1.3310 | | 1.4975 | 2.5294 | 3.1055 | 2.1155 | 2.1126 | 1.0918 | 2.1267 | 2.1325 | 3.4549 | 2.7529 | 4.0867 | 2.7910 | 3.5120 | C3 | 2.5150 | 1.4975 | | 1.5342 | 2.5526 | 3.5036 | 2.7812 | 2.2024 | 1.0990 | 1.0976 | 2.1542 | 2.1585 | 3.5049 | 2.8095 | 2.8242 | C4 | 3.5946 | 2.5294 | 1.5342 | | 1.5228 | 4.4808 | 3.9021 | 2.8597 | 2.1540 | 2.1664 | 1.0967 | 1.0983 | 2.1753 | 2.1723 | 2.1709 | C5 | 3.7603 | 3.1055 | 2.5526 | 1.5228 | | 4.5529 | 3.9103 | 3.5381 | 3.4815 | 2.7830 | 2.1566 | 2.1569 | 1.0942 | 1.0943 | 1.0957 | H6 | 1.0859 | 2.1155 | 3.5036 | 4.4808 | 4.5529 | | 1.8510 | 2.4316 | 4.1139 | 3.7208 | 5.4593 | 4.6861 | 5.5188 | 3.8336 | 4.7831 | H7 | 1.0877 | 2.1126 | 2.7812 | 3.9021 | 3.9103 | 1.8510 | | 3.0712 | 3.5184 | 2.4467 | 4.6728 | 4.5218 | 4.9861 | 3.5131 | 3.7453 | H8 | 2.0900 | 1.0918 | 2.2024 | 2.8597 | 3.5381 | 2.4316 | 3.0712 | | 2.5462 | 3.0882 | 3.8199 | 2.6406 | 4.3500 | 3.1315 | 4.1947 | H9 | 3.2041 | 2.1267 | 1.0990 | 2.1540 | 3.4815 | 4.1139 | 3.5184 | 2.5462 | | 1.7617 | 2.4135 | 2.4990 | 4.3114 | 3.8273 | 3.7836 | H10 | 2.6377 | 2.1325 | 1.0976 | 2.1664 | 2.7830 | 3.7208 | 2.4467 | 3.0882 | 1.7617 | | 2.5157 | 3.0675 | 3.7929 | 3.1238 | 2.6086 | H11 | 4.5061 | 3.4549 | 2.1542 | 1.0967 | 2.1566 | 5.4593 | 4.6728 | 3.8199 | 2.4135 | 2.5157 | | 1.7538 | 2.5056 | 3.0742 | 2.5122 | H12 | 3.9696 | 2.7529 | 2.1585 | 1.0983 | 2.1569 | 4.6861 | 4.5218 | 2.6406 | 2.4990 | 3.0675 | 1.7538 | | 2.4935 | 2.5301 | 3.0740 | H13 | 4.7968 | 4.0867 | 3.5049 | 2.1753 | 1.0942 | 5.5188 | 4.9861 | 4.3500 | 4.3114 | 3.7929 | 2.5056 | 2.4935 | | 1.7715 | 1.7686 | H14 | 3.2305 | 2.7910 | 2.8095 | 2.1723 | 1.0943 | 3.8336 | 3.5131 | 3.1315 | 3.8273 | 3.1238 | 3.0742 | 2.5301 | 1.7715 | | 1.7645 | H15 | 3.9064 | 3.5120 | 2.8242 | 2.1709 | 1.0957 | 4.7831 | 3.7453 | 4.1947 | 3.7836 | 2.6086 | 2.5122 | 3.0740 | 1.7686 | 1.7645 | |
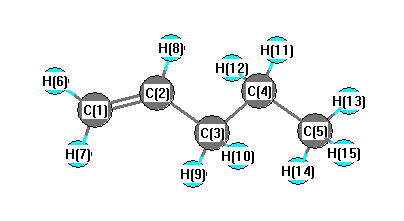 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.437 |
|
C1 |
C2 |
H8 |
118.895 |
| C2 |
C1 |
H6 |
121.834 |
|
C2 |
C1 |
H7 |
121.395 |
| C2 |
C3 |
C4 |
113.088 |
|
C2 |
C3 |
H9 |
109.013 |
| C2 |
C3 |
H10 |
109.554 |
|
C3 |
C2 |
H8 |
115.665 |
| C3 |
C4 |
C5 |
113.232 |
|
C3 |
C4 |
H11 |
108.792 |
| C3 |
C4 |
H12 |
109.037 |
|
C4 |
C3 |
H9 |
108.642 |
| C4 |
C3 |
H10 |
109.690 |
|
C4 |
C5 |
H13 |
111.398 |
| C4 |
C5 |
H14 |
111.144 |
|
C4 |
C5 |
H15 |
110.947 |
| C5 |
C4 |
H11 |
109.761 |
|
C5 |
C4 |
H12 |
109.692 |
| H6 |
C1 |
H7 |
116.770 |
|
H9 |
C3 |
H10 |
106.644 |
| H11 |
C4 |
H12 |
106.071 |
|
H13 |
C5 |
H14 |
108.091 |
| H13 |
C5 |
H15 |
107.732 |
|
H14 |
C5 |
H15 |
107.356 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.295 |
|
|
|
| 2 |
C |
-0.050 |
|
|
|
| 3 |
C |
-0.273 |
|
|
|
| 4 |
C |
-0.243 |
|
|
|
| 5 |
C |
-0.385 |
|
|
|
| 6 |
H |
0.122 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
| 8 |
H |
0.113 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.123 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.122 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
H |
0.138 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.252 |
-0.208 |
-0.050 |
0.330 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.362 |
-1.189 |
0.357 |
| y |
-1.189 |
-33.437 |
0.427 |
| z |
0.357 |
0.427 |
-31.758 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.765 |
-1.189 |
0.357 |
| y |
-1.189 |
-0.877 |
0.427 |
| z |
0.357 |
0.427 |
1.642 |
|
| Polar |
|---|
| 3z2-r2 | 3.283 |
|---|
| x2-y2 | 0.074 |
|---|
| xy | -1.189 |
|---|
| xz | 0.357 |
|---|
| yz | 0.427 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.126 |
-1.398 |
-0.194 |
| y |
-1.398 |
7.341 |
0.459 |
| z |
-0.194 |
0.459 |
7.277 |
<r2> (average value of r
2) Å
2
| <r2> |
151.917 |
| (<r2>)1/2 |
12.325 |
