return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2CH2CH2CH3 (1-Butanamine)

using model chemistry: CCD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/6-31G**
 hartrees
Energy at 0K-213.159745
Energy at 298.15K-213.172520
Nuclear repulsion energy185.629821
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3562 3340 1.08      
2 A' 3198 2999 37.90      
3 A' 3112 2918 65.82      
4 A' 3112 2918 31.37      
5 A' 3103 2909 11.34      
6 A' 3085 2893 16.37      
7 A' 1716 1609 22.24      
8 A' 1573 1475 1.98      
9 A' 1563 1466 0.27      
10 A' 1552 1455 0.48      
11 A' 1547 1451 0.23      
12 A' 1477 1385 1.65      
13 A' 1469 1377 6.88      
14 A' 1429 1340 3.92      
15 A' 1329 1246 9.49      
16 A' 1186 1112 7.39      
17 A' 1133 1062 9.04      
18 A' 1109 1040 4.81      
19 A' 1061 995 30.10      
20 A' 949 890 65.06      
21 A' 914 857 107.69      
22 A' 440 412 5.20      
23 A' 401 376 0.42      
24 A' 186 174 2.45      
25 A" 3653 3425 0.44      
26 A" 3194 2995 56.32      
27 A" 3165 2967 66.99      
28 A" 3147 2951 6.34      
29 A" 3124 2929 3.21      
30 A" 1562 1465 4.87      
31 A" 1432 1343 0.98      
32 A" 1372 1286 0.29      
33 A" 1352 1268 0.07      
34 A" 1278 1198 0.00      
35 A" 1076 1009 0.70      
36 A" 948 888 0.12      
37 A" 807 756 0.52      
38 A" 749 702 1.67      
39 A" 305 286 53.43      
40 A" 258 242 0.21      
41 A" 126 118 0.04      
42 A" 110 103 3.18      

Unscaled Zero Point Vibrational Energy (zpe) 33931.0 cm-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 31813.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G**
ABC
0.59822 0.06488 0.06176

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 2.536 0.422 0.000
C2 1.337 -0.420 0.000
C3 0.000 0.325 0.000
C4 -1.212 -0.607 0.000
C5 -2.541 0.145 0.000
H6 2.511 1.035 0.808
H7 2.511 1.035 -0.808
H8 1.388 -1.073 0.875
H9 1.388 -1.073 -0.875
H10 -0.044 0.980 -0.877
H11 -0.044 0.980 0.877
H12 -1.161 -1.261 0.875
H13 -1.161 -1.261 -0.875
H14 -3.387 -0.544 0.000
H15 -2.628 0.782 0.881
H16 -2.628 0.782 -0.881

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46522.53783.88625.08501.01481.01482.07822.07822.78152.78154.15544.15546.00095.25065.2506
C21.46521.53032.55523.91912.03692.03691.09311.09312.15312.15312.77722.77724.72514.23534.2353
C32.53781.53031.52862.54792.73192.73192.15582.15581.09551.09552.15192.15193.49642.80892.8089
C43.88622.55521.52861.52764.14824.14822.78222.78222.15672.15671.09391.09392.17602.17052.1705
C55.08503.91912.54791.52765.19385.19384.20594.20592.77582.77582.15592.15591.09021.09111.0911
H61.01482.03692.73194.14825.19381.61672.38972.92223.06142.55664.33164.64676.15885.14555.4153
H71.01482.03692.73194.14825.19381.61672.92222.38972.55663.06144.64674.33166.15885.41535.1455
H82.07821.09312.15582.78224.20592.38972.92221.74963.05562.50352.55613.09774.88304.42354.7594
H92.07821.09312.15582.78224.20592.92222.38971.74962.50353.05563.09772.55614.88304.75944.4235
H102.78152.15311.09552.15672.77583.06142.55663.05562.50351.75413.05652.50443.77703.13152.5912
H112.78152.15311.09552.15672.77582.55663.06142.50353.05561.75412.50443.05653.77702.59123.1315
H124.15542.77722.15191.09392.15594.33164.64672.55613.09773.05652.50441.75022.49682.51513.0677
H134.15542.77722.15191.09392.15594.64674.33163.09772.55612.50443.05651.75022.49683.06772.5151
H146.00094.72513.49642.17601.09026.15886.15884.88304.88303.77703.77702.49682.49681.76381.7638
H155.25064.23532.80892.17051.09115.14555.41534.42354.75943.13152.59122.51513.06771.76381.7628
H165.25064.23532.80892.17051.09115.41535.14554.75944.42352.59123.13153.06772.51511.76381.7628

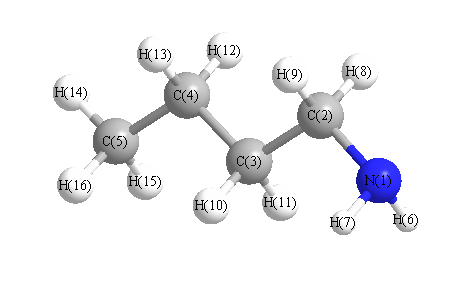
picture of 1-Butanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 115.799 N1 C2 H8 107.762
N1 C2 H9 107.762 C2 N1 H6 109.090
C2 N1 H7 109.090 C2 C3 C4 113.299
C2 C3 H10 109.034 C2 C3 H11 109.034
C3 C2 H8 109.389 C3 C2 H9 109.389
C3 C4 C5 112.959 C3 C4 H12 109.156
C3 C4 H13 109.156 C4 C3 H10 109.437
C4 C3 H11 109.437 C4 C5 H14 111.361
C4 C5 H15 110.861 C4 C5 H16 110.861
C5 C4 H12 109.542 C5 C4 H13 109.542
H6 N1 H7 105.614 H8 C2 H9 106.316
H10 C3 H11 106.364 H12 C4 H13 106.263
H14 C5 H15 107.923 H14 C5 H16 107.923
H15 C5 H16 107.764
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability