return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: CCD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/6-31G**
 hartrees
Energy at 0K-213.153929
Energy at 298.15K-213.166739
HF Energy-212.332492
Nuclear repulsion energy188.611899
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3575 3352 1.02      
2 A' 3218 3017 47.03      
3 A' 3195 2996 66.15      
4 A' 3137 2941 53.82      
5 A' 3116 2922 12.31      
6 A' 3030 2841 143.58      
7 A' 1592 1493 0.10      
8 A' 1562 1465 2.10      
9 A' 1554 1457 8.70      
10 A' 1491 1398 2.27      
11 A' 1450 1360 0.14      
12 A' 1362 1277 4.66      
13 A' 1281 1201 1.90      
14 A' 1208 1132 13.00      
15 A' 1101 1032 5.08      
16 A' 935 877 9.76      
17 A' 852 799 5.13      
18 A' 811 760 70.08      
19 A' 436 409 0.35      
20 A' 269 253 1.29      
21 A' 192 180 0.78      
22 A' 115 107 1.43      
23 A" 3217 3017 9.01      
24 A" 3195 2996 17.51      
25 A" 3133 2938 0.62      
26 A" 3116 2921 29.80      
27 A" 3025 2837 7.51      
28 A" 1581 1483 4.38      
29 A" 1564 1466 3.57      
30 A" 1559 1461 7.51      
31 A" 1532 1436 30.40      
32 A" 1473 1381 8.26      
33 A" 1409 1321 28.94      
34 A" 1327 1245 2.63      
35 A" 1205 1129 34.86      
36 A" 1150 1079 8.03      
37 A" 1102 1033 3.20      
38 A" 976 915 0.42      
39 A" 829 778 0.57      
40 A" 437 410 0.54      
41 A" 269 253 0.66      
42 A" 110 103 1.16      

Unscaled Zero Point Vibrational Energy (zpe) 33845.8 cm-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 31733.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G**
ABC
0.59294 0.07021 0.06618

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.288 0.000
C2 0.017 0.519 1.217
C3 0.017 0.519 -1.217
C4 0.017 -0.371 2.452
C5 0.017 -0.371 -2.452
H6 -0.817 -0.869 0.000
H7 -0.831 1.221 1.261
H8 0.924 1.130 1.206
H9 -0.831 1.221 -1.261
H10 0.924 1.130 -1.206
H11 0.058 0.226 3.363
H12 -0.888 -0.980 2.492
H13 0.876 -1.041 2.431
H14 0.058 0.226 -3.363
H15 -0.888 -0.980 -2.492
H16 0.876 -1.041 -2.431

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46051.46052.45342.45341.01602.14122.07072.14122.07073.40222.74042.68603.40222.74042.6860
C21.46052.43421.52233.77552.02541.10131.09302.71122.65862.16602.16622.15514.58954.10204.0597
C31.46052.43423.77551.52232.02542.71122.65861.10131.09304.58954.10204.05972.16602.16622.1551
C42.45341.52233.77554.90402.63732.16142.15084.12754.05681.09011.09161.08885.84565.06335.0032
C52.45343.77551.52234.90402.63734.12754.05682.16142.15085.84565.06335.00321.09011.09161.0888
H61.01602.02542.02542.63732.63732.44002.91162.44002.91163.64312.49592.96733.64312.49592.9673
H72.14121.10132.71122.16144.12752.44001.75752.52143.02862.48932.52243.06524.81194.35104.6536
H82.07071.09302.65862.15084.05682.91161.75753.02862.41282.49323.06372.49294.73744.62754.2364
H92.14122.71121.10134.12752.16142.44002.52143.02861.75754.81194.35104.65362.48932.52243.0652
H102.07072.65861.09304.05682.15082.91163.02862.41281.75754.73744.62754.23642.49323.06372.4929
H113.40222.16604.58951.09015.84563.64312.48932.49324.81194.73741.76281.77256.72566.05265.9872
H122.74042.16624.10201.09165.06332.49592.52243.06374.35104.62751.76281.76566.05264.98485.2304
H132.68602.15514.05971.08885.00322.96733.06522.49294.65364.23641.77251.76565.98725.23044.8628
H143.40224.58952.16605.84561.09013.64314.81194.73742.48932.49326.72566.05265.98721.76281.7725
H152.74044.10202.16625.06331.09162.49594.35104.62752.52243.06376.05264.98485.23041.76281.7656
H162.68604.05972.15515.00321.08882.96734.65364.23643.06522.49295.98725.23044.86281.77251.7656

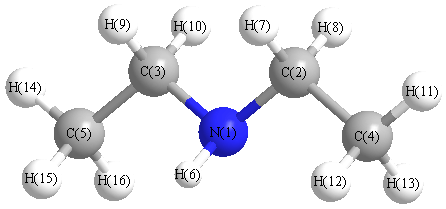
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.657 N1 C2 H7 112.648
N1 C2 H8 107.498 N1 C3 C5 110.657
N1 C3 H9 112.648 N1 C3 H10 107.498
C2 N1 C3 112.888 C2 N1 H6 108.403
C2 C4 H11 110.933 C2 C4 H12 110.864
C2 C4 H13 110.141 C3 N1 H6 108.403
C3 C5 H14 110.933 C3 C5 H15 110.864
C3 C5 H16 110.141 C4 C2 H7 109.899
C4 C2 H8 109.554 C5 C3 H8 150.904
C5 C3 H10 109.554 H7 C2 H8 106.436
H9 C3 H10 106.436 H11 C4 H12 107.796
H11 C4 H13 108.878 H12 C4 H13 108.141
H14 C5 H15 107.796 H14 C5 H16 108.878
H15 C5 H16 108.141
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability