return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: MP3=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3=FULL/6-31G**
 hartrees
Energy at 0K-213.172980
Energy at 298.15K-213.185808
HF Energy-212.332707
Nuclear repulsion energy188.930985
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3601 3601 0.85      
2 A' 3240 3240 44.34      
3 A' 3218 3218 63.64      
4 A' 3157 3157 54.86      
5 A' 3138 3138 11.81      
6 A' 3049 3049 144.23      
7 A' 1598 1598 0.17      
8 A' 1568 1568 2.21      
9 A' 1560 1560 9.42      
10 A' 1496 1496 2.48      
11 A' 1454 1454 0.17      
12 A' 1368 1368 4.71      
13 A' 1284 1284 1.90      
14 A' 1212 1212 13.48      
15 A' 1106 1106 5.24      
16 A' 938 938 9.62      
17 A' 855 855 4.80      
18 A' 813 813 71.29      
19 A' 438 438 0.36      
20 A' 271 271 1.29      
21 A' 192 192 0.79      
22 A' 115 115 1.41      
23 A" 3240 3240 8.94      
24 A" 3218 3218 16.58      
25 A" 3154 3154 0.68      
26 A" 3137 3137 28.06      
27 A" 3045 3045 7.60      
28 A" 1587 1587 4.59      
29 A" 1570 1570 3.26      
30 A" 1564 1564 7.99      
31 A" 1537 1537 31.18      
32 A" 1477 1477 9.17      
33 A" 1413 1413 28.05      
34 A" 1332 1332 2.66      
35 A" 1210 1210 35.51      
36 A" 1155 1155 7.89      
37 A" 1106 1106 3.29      
38 A" 980 980 0.35      
39 A" 832 832 0.57      
40 A" 438 438 0.55      
41 A" 271 271 0.67      
42 A" 111 111 1.18      

Unscaled Zero Point Vibrational Energy (zpe) 34023.0 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 34023.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3=FULL/6-31G**
ABC
0.59521 0.07042 0.06638

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.288 0.000
C2 0.017 0.518 1.215
C3 0.017 0.518 -1.215
C4 0.017 -0.370 2.449
C5 0.017 -0.370 -2.449
H6 -0.814 -0.869 0.000
H7 -0.829 1.218 1.258
H8 0.922 1.128 1.204
H9 -0.829 1.218 -1.258
H10 0.922 1.128 -1.204
H11 0.058 0.226 3.357
H12 -0.886 -0.978 2.488
H13 0.874 -1.038 2.428
H14 0.058 0.226 -3.357
H15 -0.886 -0.978 -2.488
H16 0.874 -1.038 -2.428

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.45831.45832.44992.44991.01362.13732.06722.13732.06723.39662.73542.68143.39662.73542.6814
C21.45832.42981.52023.76962.02251.09901.09072.70572.65352.16242.16222.15114.58154.09484.0529
C31.45832.42983.76961.52022.02252.70572.65351.09901.09074.58154.09484.05292.16242.16222.1511
C42.44991.52023.76964.89702.63332.15792.14714.12024.05011.08761.08921.08645.83635.05544.9959
C52.44993.76961.52024.89702.63334.12024.05012.15792.14715.83635.05544.99591.08761.08921.0864
H61.01362.02252.02252.63332.63332.43692.90652.43692.90653.63712.49182.96153.63712.49182.9615
H72.13731.09902.70572.15794.12022.43691.75382.51583.02232.48522.51783.05934.80294.34284.6449
H82.06721.09072.65352.14714.05012.90651.75383.02232.40822.48903.05772.48824.72914.61894.2290
H92.13732.70571.09904.12022.15792.43692.51583.02231.75384.80294.34284.64492.48522.51783.0593
H102.06722.65351.09074.05012.14712.90653.02232.40821.75384.72914.61894.22902.48903.05772.4882
H113.39662.16244.58151.08765.83633.63712.48522.48904.80294.72911.75901.76896.71426.04225.9775
H122.73542.16224.09481.08925.05542.49182.51783.05774.34284.61891.75901.76176.04224.97675.2219
H132.68142.15114.05291.08644.99592.96153.05932.48824.64494.22901.76891.76175.97755.22194.8556
H143.39664.58152.16245.83631.08763.63714.80294.72912.48522.48906.71426.04225.97751.75901.7689
H152.73544.09482.16225.05541.08922.49184.34284.61892.51783.05776.04224.97675.22191.75901.7617
H162.68144.05292.15114.99591.08642.96154.64494.22903.05932.48825.97755.22194.85561.76891.7617

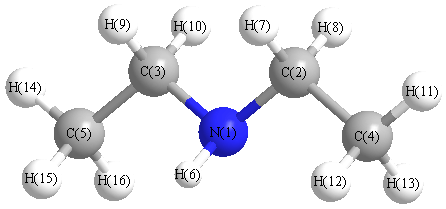
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.659 N1 C2 H7 112.636
N1 C2 H8 107.506 N1 C3 C5 110.659
N1 C3 H9 112.636 N1 C3 H10 107.506
C2 N1 C3 112.838 C2 N1 H6 108.467
C2 C4 H11 110.944 C2 C4 H12 110.832
C2 C4 H13 110.115 C3 N1 H6 108.467
C3 C5 H14 110.944 C3 C5 H15 110.832
C3 C5 H16 110.115 C4 C2 H7 109.905
C4 C2 H8 109.548 C5 C3 H8 150.925
C5 C3 H10 109.548 H7 C2 H8 106.438
H9 C3 H10 106.438 H11 C4 H12 107.819
H11 C4 H13 108.905 H12 C4 H13 108.139
H14 C5 H15 107.819 H14 C5 H16 108.905
H15 C5 H16 108.139
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability